return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (Ethylcyclobutane)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-233.419521
Energy at 298.15K-233.433201
HF Energy-232.870769
Nuclear repulsion energy246.591879
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3191 3036 49.62      
2 A 3171 3017 11.10      
3 A 3165 3011 13.98      
4 A 3157 3004 34.17      
5 A 3152 2998 31.01      
6 A 3126 2974 2.24      
7 A 3123 2971 56.23      
8 A 3110 2958 13.92      
9 A 3107 2956 34.68      
10 A 3102 2951 5.59      
11 A 3081 2931 17.84      
12 A 3070 2920 17.66      
13 A 1606 1528 4.63      
14 A 1602 1524 6.25      
15 A 1593 1515 2.01      
16 A 1583 1506 1.36      
17 A 1575 1498 2.32      
18 A 1564 1488 3.06      
19 A 1490 1418 5.56      
20 A 1431 1361 0.25      
21 A 1401 1333 3.05      
22 A 1366 1300 0.02      
23 A 1322 1258 1.06      
24 A 1317 1253 3.21      
25 A 1313 1249 0.07      
26 A 1291 1228 0.56      
27 A 1284 1221 1.12      
28 A 1223 1163 0.32      
29 A 1217 1157 1.00      
30 A 1176 1119 1.58      
31 A 1088 1035 0.36      
32 A 1065 1013 2.48      
33 A 1033 983 2.01      
34 A 984 936 0.75      
35 A 931 886 1.39      
36 A 912 867 0.96      
37 A 904 860 1.68      
38 A 899 856 0.49      
39 A 840 799 0.44      
40 A 785 746 0.04      
41 A 775 738 3.20      
42 A 683 650 3.61      
43 A 435 413 0.24      
44 A 371 353 0.19      
45 A 259 246 0.08      
46 A 227 216 0.07      
47 A 175 167 0.02      
48 A 126 120 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 37698.7 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 35862.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.23011 0.07734 0.06887

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.412 0.260 -0.056
H2 2.769 0.061 0.961
H3 3.216 0.017 -0.760
H4 2.191 1.330 -0.141
C5 1.144 -0.580 -0.351
H6 1.385 -1.649 -0.263
H7 0.815 -0.403 -1.384
C8 -1.781 0.342 -0.581
H9 -2.821 0.663 -0.498
H10 -1.491 0.287 -1.634
C11 -0.707 1.135 0.266
H12 -1.162 1.579 1.157
H13 -0.100 1.879 -0.255
C14 -0.014 -0.247 0.608
H15 0.305 -0.381 1.647
C16 -1.360 -0.969 0.195
H17 -1.265 -1.892 -0.383
H18 -2.004 -1.133 1.065

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 C11 H12 H13 C14 H15 C16 H17 H18
C11.09591.09571.09531.54992.17752.18084.22685.26774.21023.25563.99832.99582.56642.78443.97604.27304.7651
H21.09591.77821.77652.18482.51713.08754.81235.80894.99313.70394.21853.60812.82222.59574.32444.67884.9217
H31.09571.77821.77662.19552.52532.51705.01116.07784.79564.20655.02823.83693.51823.79844.77854.88545.6490
H41.09531.77651.77662.18823.08772.53844.11665.06924.10792.93313.60422.35932.81263.11114.24394.73065.0122
C51.54992.18482.19552.18821.09881.09833.07534.15803.05612.59743.50012.75781.53992.17622.59252.74263.4960
H62.17752.51712.52533.08771.09881.76993.75314.80523.72783.52154.34923.82772.16292.53392.86502.66333.6762
H72.18083.08752.51702.53841.09831.76992.81733.89122.41982.72083.78032.70572.16323.07362.74682.74593.8047
C84.22684.81235.01114.11663.07533.75312.81731.09191.09311.58072.22082.30072.21013.13671.58082.30142.2219
H95.26775.80896.07785.06924.15804.80523.89121.09191.78892.29692.51592.99023.15193.93262.29752.99402.5175
H104.21024.99314.79564.10793.05613.72782.41981.09311.78892.22293.09242.52372.73753.79942.22252.52233.0924
C113.25563.70394.20652.93312.59743.52152.72081.58072.29692.22291.09451.09291.58362.28652.20393.14512.7322
H123.99834.21855.02823.60423.50014.34923.78032.22082.51593.09241.09451.79232.22562.49622.73033.79822.8412
H132.99583.60813.83692.35932.75783.82772.70572.30072.99022.52371.09291.79232.29712.98183.14713.94883.8006
C142.56642.82223.51822.81261.53992.16292.16322.21013.15192.73751.58362.22562.29711.09481.58252.29162.2260
H152.78442.59573.79843.11112.17622.53393.07363.13673.93263.79942.28652.49622.98181.09482.28652.97832.4978
C163.97604.32444.77854.24392.59252.86502.74681.58082.29752.22252.20392.73033.14711.58252.28651.09341.0945
H174.27304.67884.88544.73062.74262.66332.74592.30142.99402.52233.14513.79823.94882.29162.97831.09341.7944
H184.76514.92175.64905.01223.49603.67623.80472.22192.51753.09242.73222.84123.80062.22602.49781.09451.7944

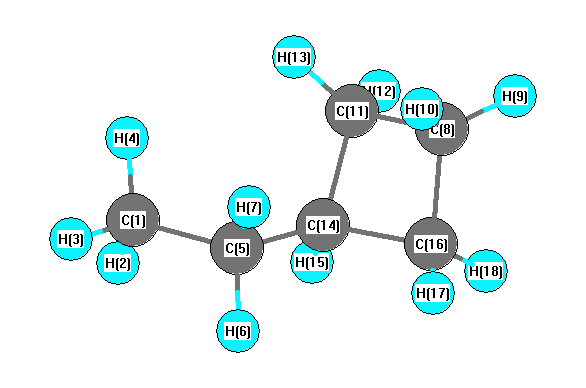
picture of Ethylcyclobutane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 109.412 C1 C5 H7 109.689
C1 C5 C14 112.316 H2 C1 H3 108.462
H2 C1 H4 108.345 H2 C1 C5 110.146
H3 C1 H4 108.361 H3 C1 C5 110.998
H4 C1 C5 110.452 C5 C14 C11 112.514
C5 C14 H15 110.227 C5 C14 C16 112.248
H6 C5 H7 107.327 H6 C5 C14 108.959
H7 C5 C14 109.008 C8 C11 H12 110.922
C8 C11 H13 117.586 C8 C11 C14 88.606
C8 C16 C14 88.638 C8 C16 H17 117.610
C8 C16 H18 110.997 H9 C8 H10 109.917
H9 C8 C11 117.337 H9 C8 C16 117.377
H10 C8 C11 111.173 H10 C8 C16 111.132
C11 C8 C16 88.393 C11 C14 H15 116.029
C11 C14 C16 88.232 H12 C11 H13 110.044
H12 C11 C14 111.099 H13 C11 C14 117.054
C14 C16 H17 116.638 C14 C16 H18 111.200
H15 C14 C16 116.112 H17 C16 H18 110.205
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability