return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (1,3-Dioxolane, 2-methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-304.734335
Energy at 298.15K-304.744735
HF Energy-304.136743
Nuclear repulsion energy257.922459
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3193 3038 28.44      
2 A 3191 3035 21.71      
3 A 3189 3033 13.55      
4 A 3181 3026 13.28      
5 A 3108 2957 9.55      
6 A 3105 2953 54.81      
7 A 3097 2947 42.61      
8 A 3093 2942 24.72      
9 A 1605 1527 0.32      
10 A 1598 1520 2.49      
11 A 1593 1515 2.13      
12 A 1587 1510 3.63      
13 A 1491 1418 12.49      
14 A 1456 1385 38.90      
15 A 1415 1346 1.29      
16 A 1409 1341 17.53      
17 A 1341 1276 8.28      
18 A 1263 1201 5.84      
19 A 1229 1169 0.20      
20 A 1181 1124 19.21      
21 A 1162 1106 9.30      
22 A 1143 1088 31.03      
23 A 1123 1068 112.13      
24 A 1041 991 14.32      
25 A 991 943 33.61      
26 A 932 887 12.49      
27 A 919 874 3.90      
28 A 838 797 27.20      
29 A 814 774 9.97      
30 A 682 649 2.54      
31 A 622 592 5.46      
32 A 459 437 16.04      
33 A 346 330 2.52      
34 A 251 239 0.83      
35 A 233 222 0.66      
36 A 52 49 12.28      

Unscaled Zero Point Vibrational Energy (zpe) 26965.7 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 25652.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.20518 0.11474 0.08694

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.986 0.006 0.394
H2 2.551 -0.928 0.345
H3 2.621 0.839 0.082
H4 1.650 0.168 1.421
C5 0.761 -0.079 -0.504
H6 1.008 -0.207 -1.562
O7 -0.051 1.157 -0.404
O8 -0.069 -1.201 -0.021
C9 -1.462 -0.688 0.016
H10 -2.001 -1.265 0.767
H11 -1.943 -0.775 -0.964
C12 -1.252 0.791 0.389
H13 -1.064 0.893 1.463
H14 -2.063 1.448 0.075

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 O7 O8 C9 H10 H11 C12 H13 H14
C11.09301.09281.09281.52072.19692.47102.41943.53664.20094.22983.33113.35054.3098
H21.09301.78781.78092.15472.55593.41652.65934.03274.58354.68314.17314.19855.1969
H31.09281.78781.78522.15512.52942.73383.37804.35905.12414.95293.88513.93514.7237
H41.09281.78091.78522.13483.07452.68362.62903.51983.97614.41493.14232.80894.1521
C51.52072.15472.15512.13481.09411.48181.47742.36243.26372.83032.36772.85363.2628
H62.19692.55592.52943.07451.09412.07912.12692.97003.94963.06473.14823.82813.8545
O72.47103.41652.73382.68361.48182.07912.38912.35983.32272.76181.48522.14072.0896
O82.41942.65933.37802.62901.47742.12692.38911.48472.08742.14092.35302.75263.3177
C93.53664.03274.35903.51982.36242.97002.35981.48471.09041.09541.53922.17942.2197
H104.20094.58355.12413.97613.26373.94963.32272.08741.09041.80022.22052.45302.8006
H114.22984.68314.95294.41492.83033.06472.76182.14091.09541.80022.18203.07342.4570
C123.33114.17313.88513.14232.36773.14821.48522.35301.53922.22052.18201.09521.0905
H133.35054.19853.93512.80892.85363.82812.14072.75262.17942.45303.07341.09521.7985
H144.30985.19694.72374.15213.26283.85452.08963.31772.21972.80062.45701.09051.7985

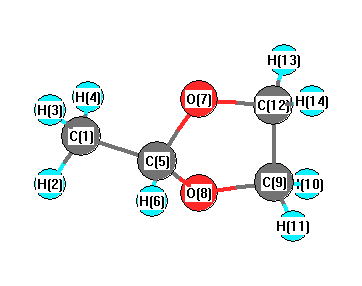
picture of 1,3-Dioxolane, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 113.310 C1 C5 O7 110.763
C1 C5 O8 107.593 H2 C1 H3 109.752
H2 C1 H4 109.126 H2 C1 C5 109.968
H3 C1 H4 109.529 H3 C1 C5 110.014
H4 C1 C5 108.428 C5 O7 C12 105.883
C5 O8 C9 105.798 H6 C5 O7 106.660
H6 C5 O8 110.721 O7 C5 O8 107.679
O7 C12 C9 102.549 O7 C12 H13 111.217
O7 C12 H14 107.449 O8 C9 H10 107.326
O8 C9 H11 111.262 O8 C9 C12 102.166
C9 C12 H13 110.509 C9 C12 H14 114.064
H10 C9 H11 110.895 H10 C9 C12 114.135
H11 C9 C12 110.698 H13 C12 H14 110.742
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability