return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (1,3-Dioxane)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-304.732752
Energy at 298.15K-304.743952
Nuclear repulsion energy259.120885
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3197 3041 28.66      
2 A' 3174 3020 20.79      
3 A' 3163 3009 12.42      
4 A' 3107 2955 13.22      
5 A' 3038 2890 79.63      
6 A' 2994 2848 70.96      
7 A' 1600 1522 8.83      
8 A' 1583 1506 1.95      
9 A' 1560 1484 4.40      
10 A' 1441 1371 16.70      
11 A' 1342 1277 2.54      
12 A' 1215 1156 6.11      
13 A' 1151 1095 70.39      
14 A' 1104 1050 49.47      
15 A' 992 944 46.39      
16 A' 884 841 2.68      
17 A' 821 781 9.31      
18 A' 621 591 6.73      
19 A' 479 456 1.53      
20 A' 419 399 13.81      
21 A' 271 258 2.86      
22 A" 3165 3011 31.75      
23 A" 3032 2884 15.89      
24 A" 1586 1509 0.54      
25 A" 1483 1411 6.62      
26 A" 1430 1360 0.81      
27 A" 1421 1351 6.35      
28 A" 1340 1275 0.05      
29 A" 1292 1229 12.36      
30 A" 1247 1186 5.80      
31 A" 1049 998 5.89      
32 A" 1014 965 76.64      
33 A" 946 900 20.54      
34 A" 858 816 12.52      
35 A" 447 425 9.66      
36 A" 251 239 2.44      

Unscaled Zero Point Vibrational Energy (zpe) 27357.6 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 26025.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.15953 0.15299 0.08793

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.651 -1.231 0.000
O2 0.013 -0.786 1.217
O3 0.013 -0.786 -1.217
C4 0.013 0.695 1.256
C5 0.013 0.695 -1.256
C6 0.723 1.248 0.000
H7 -0.598 -2.317 0.000
H8 -1.696 -0.876 0.000
H9 0.533 0.967 2.176
H10 -1.021 1.072 1.292
H11 0.533 0.967 -2.176
H12 -1.021 1.072 -1.292
H13 1.758 0.895 0.000
H14 0.708 2.344 0.000

Atom - Atom Distances (Å)
  C1 O2 O3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.45591.45592.39332.39332.83371.08791.10353.31162.66643.31162.66643.21273.8239
O21.45592.43341.48182.88222.47382.04912.10012.06462.12813.85413.28912.71103.4289
O31.45592.43342.88221.48182.47382.04912.10013.85413.28912.06462.12812.71103.4289
C42.39331.48182.88222.51151.54453.32062.63991.09151.10183.48182.77592.15872.1855
C52.39332.88221.48182.51151.54453.32062.63993.48182.77591.09151.10182.15872.1855
C62.83372.47382.47381.54451.54453.80183.21902.20262.17792.20262.17791.09341.0959
H71.08792.04912.04913.32063.32063.80181.81174.09873.65194.09873.65193.98364.8404
H81.10352.10012.10012.63992.63993.21901.81173.61972.43333.61972.43333.88164.0184
H93.31162.06463.85411.09153.48182.20264.09873.61971.79164.35273.80262.49822.5813
H102.66642.12813.28911.10182.77592.17793.65192.43331.79163.80262.58463.07012.5056
H113.31163.85412.06463.48181.09152.20264.09873.61974.35273.80261.79162.49822.5813
H122.66643.28912.12812.77591.10182.17793.65192.43333.80262.58461.79163.07012.5056
H133.21272.71102.71102.15872.15871.09343.98363.88162.49823.07012.49823.07011.7888
H143.82393.42893.42892.18552.18551.09594.84044.01842.58132.50562.58132.50561.7888

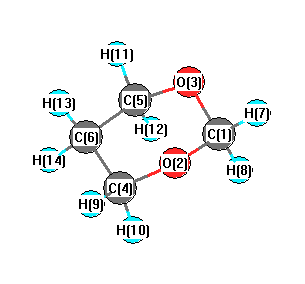
picture of 1,3-Dioxane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 C4 109.110 C1 O3 C5 109.110
O2 C1 O3 113.383 O2 C1 H7 106.431
O2 C1 H8 109.516 O2 C4 C6 109.639
O2 C4 H9 105.698 O2 C4 H10 110.036
O3 C1 H7 106.431 O3 C1 H8 109.516
O3 C5 C6 109.639 O3 C5 H11 105.698
O3 C5 H12 110.036 C4 C6 C5 108.793
C4 C6 H13 108.631 C4 C6 H14 110.577
C5 C6 H13 108.631 C5 C6 H14 110.577
C6 C4 H9 112.214 C6 C4 H10 109.640
C6 C5 H11 112.214 C6 C5 H12 109.640
H7 C1 H8 111.529 H9 C4 H10 109.543
H11 C5 H12 109.543 H13 C6 H14 109.582
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability