return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (2-Butene, 2-methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-194.522071
Energy at 298.15K-194.531890
HF Energy-194.063972
Nuclear repulsion energy173.030050
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3182 3027 32.18      
2 A 3166 3012 23.09      
3 A 3149 2996 23.91      
4 A 3142 2989 3.96      
5 A 3077 2927 13.12      
6 A 3072 2923 28.78      
7 A 3065 2915 18.82      
8 A 1730 1645 0.11      
9 A 1603 1525 10.78      
10 A 1596 1518 2.04      
11 A 1590 1513 0.85      
12 A 1501 1428 3.37      
13 A 1490 1417 5.86      
14 A 1476 1404 2.84      
15 A 1437 1367 2.88      
16 A 1250 1189 4.31      
17 A 1160 1103 8.44      
18 A 1101 1048 5.33      
19 A 997 948 1.06      
20 A 969 921 1.79      
21 A 754 717 1.65      
22 A 533 507 2.07      
23 A 390 371 0.30      
24 A 309 294 0.16      
25 A 3128 2975 36.10      
26 A 3127 2974 26.38      
27 A 3123 2971 0.33      
28 A 1596 1519 19.12      
29 A 1591 1513 0.37      
30 A 1578 1501 1.14      
31 A 1167 1111 0.71      
32 A 1125 1070 0.14      
33 A 1078 1026 1.93      
34 A 849 808 16.30      
35 A 439 418 3.44      
36 A 260 247 1.89      
37 A 184 175 0.97      
38 A 138 131 0.43      
39 A 97 92 0.42      

Unscaled Zero Point Vibrational Energy (zpe) 30608.4 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 29117.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.26431 0.11515 0.08400

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.525 0.481 0.000
H2 -1.962 -0.521 0.000
H3 -1.888 1.017 0.886
H4 -1.888 1.017 -0.886
C5 0.286 -2.112 0.000
H6 0.665 -2.638 -0.886
H7 0.665 -2.638 0.886
H8 -0.804 -2.187 0.000
C9 0.756 -0.666 0.000
H10 1.840 -0.543 0.000
C11 0.000 0.450 0.000
C12 0.644 1.835 0.000
H13 1.737 1.757 0.000
H14 0.334 2.402 0.887
H15 0.334 2.402 -0.887

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 C11 C12 H13 H14 H15
C11.09291.09771.09773.16313.91223.91222.76422.55263.51761.52512.55653.50232.81622.8162
H21.09291.77661.77662.75413.48763.48762.02912.72113.80202.18863.51244.34343.82073.8207
H31.09771.77661.77293.91224.79734.45803.49713.25684.13772.16142.80453.80432.61803.1621
H41.09771.77661.77293.91224.45804.79733.49713.25684.13772.16142.80453.80433.16212.6180
C53.16312.75413.91223.91221.09721.09721.09331.52082.20862.57813.96304.13174.60074.6007
H63.91223.48764.79734.45801.09721.77151.77362.16372.56013.28024.55924.60895.35255.0505
H73.91223.48764.45804.79731.09721.77151.77362.16372.56013.28024.55924.60895.05055.3525
H82.76422.02913.49713.49711.09331.77361.77362.17953.11442.75754.27504.69194.81104.8110
C92.55262.72113.25683.25681.52082.16372.16372.17951.09161.34752.50272.61333.22103.2210
H103.51763.80204.13774.13772.20862.56012.56013.11441.09162.09122.66172.30183.42483.4248
C111.52512.18862.16142.16142.57813.28023.28022.75751.34752.09121.52712.17352.16992.1699
C122.55653.51242.80452.80453.96304.55924.55924.27502.50272.66171.52711.09581.09761.0976
H133.50234.34343.80433.80434.13174.60894.60894.69192.61332.30182.17351.09581.78111.7811
H142.81623.82072.61803.16214.60075.35255.05054.81103.22103.42482.16991.09761.78111.7739
H152.81623.82073.16212.61804.60075.05055.35254.81103.22103.42482.16991.09761.78111.7739

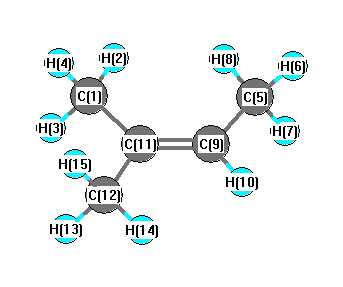
picture of 2-Butene, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C11 C9 125.286 C1 C11 C12 113.774
H2 C1 H3 108.387 H2 C1 H4 108.387
H2 C1 C11 112.383 H3 C1 H4 107.711
H3 C1 C11 109.921 H4 C1 C11 109.921
C5 C9 H10 114.431 C5 C9 C11 127.903
H6 C5 H7 107.654 H6 C5 H8 108.128
H6 C5 C9 110.427 H7 C5 H8 108.128
H7 C5 C9 110.427 H8 C5 C9 111.935
C9 C11 C12 120.940 H10 C9 C11 117.666
C11 C12 H13 110.853 C11 C12 H14 110.457
C11 C12 H15 110.457 H13 C12 H14 108.589
H13 C12 H15 108.589 H14 C12 H15 107.809
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability