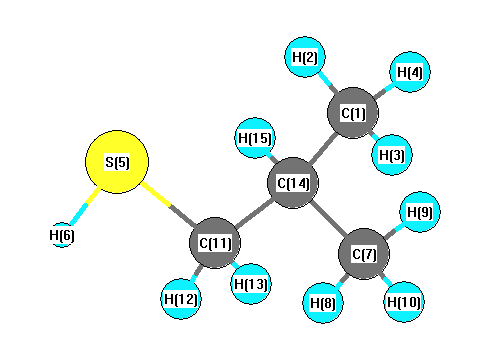
Vibrational Frequencies calculated at B3PW91/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3132 |
3005 |
27.02 |
|
|
|
| 2 |
A |
3130 |
3003 |
46.98 |
|
|
|
| 3 |
A |
3126 |
2999 |
26.99 |
|
|
|
| 4 |
A |
3121 |
2994 |
5.08 |
|
|
|
| 5 |
A |
3118 |
2991 |
4.00 |
|
|
|
| 6 |
A |
3070 |
2945 |
21.10 |
|
|
|
| 7 |
A |
3061 |
2936 |
4.90 |
|
|
|
| 8 |
A |
3050 |
2926 |
12.87 |
|
|
|
| 9 |
A |
3046 |
2923 |
18.49 |
|
|
|
| 10 |
A |
2704 |
2595 |
16.91 |
|
|
|
| 11 |
A |
1567 |
1503 |
22.22 |
|
|
|
| 12 |
A |
1563 |
1500 |
5.56 |
|
|
|
| 13 |
A |
1552 |
1489 |
6.67 |
|
|
|
| 14 |
A |
1545 |
1482 |
0.09 |
|
|
|
| 15 |
A |
1524 |
1462 |
6.15 |
|
|
|
| 16 |
A |
1465 |
1405 |
14.39 |
|
|
|
| 17 |
A |
1446 |
1388 |
16.05 |
|
|
|
| 18 |
A |
1396 |
1340 |
2.42 |
|
|
|
| 19 |
A |
1381 |
1324 |
0.79 |
|
|
|
| 20 |
A |
1315 |
1262 |
18.50 |
|
|
|
| 21 |
A |
1276 |
1224 |
2.72 |
|
|
|
| 22 |
A |
1217 |
1168 |
9.01 |
|
|
|
| 23 |
A |
1164 |
1117 |
6.34 |
|
|
|
| 24 |
A |
1116 |
1071 |
5.25 |
|
|
|
| 25 |
A |
994 |
953 |
0.89 |
|
|
|
| 26 |
A |
993 |
952 |
0.60 |
|
|
|
| 27 |
A |
968 |
929 |
3.98 |
|
|
|
| 28 |
A |
921 |
883 |
2.06 |
|
|
|
| 29 |
A |
898 |
861 |
4.92 |
|
|
|
| 30 |
A |
801 |
768 |
3.11 |
|
|
|
| 31 |
A |
737 |
707 |
1.97 |
|
|
|
| 32 |
A |
430 |
412 |
0.22 |
|
|
|
| 33 |
A |
402 |
386 |
0.99 |
|
|
|
| 34 |
A |
341 |
327 |
0.21 |
|
|
|
| 35 |
A |
262 |
251 |
0.11 |
|
|
|
| 36 |
A |
235 |
226 |
0.13 |
|
|
|
| 37 |
A |
212 |
204 |
2.14 |
|
|
|
| 38 |
A |
156 |
150 |
10.97 |
|
|
|
| 39 |
A |
72 |
69 |
16.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29251.5 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 28063.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.592 |
|
|
|
| 2 |
H |
0.234 |
|
|
|
| 3 |
H |
0.206 |
|
|
|
| 4 |
H |
0.210 |
|
|
|
| 5 |
S |
-0.084 |
|
|
|
| 6 |
H |
0.134 |
|
|
|
| 7 |
C |
-0.583 |
|
|
|
| 8 |
H |
0.209 |
|
|
|
| 9 |
H |
0.215 |
|
|
|
| 10 |
H |
0.208 |
|
|
|
| 11 |
C |
-0.553 |
|
|
|
| 12 |
H |
0.239 |
|
|
|
| 13 |
H |
0.236 |
|
|
|
| 14 |
C |
-0.314 |
|
|
|
| 15 |
H |
0.235 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.109 |
-1.183 |
0.693 |
1.763 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.569 |
2.566 |
-1.834 |
| y |
2.566 |
-39.988 |
-1.099 |
| z |
-1.834 |
-1.099 |
-41.395 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.122 |
2.566 |
-1.834 |
| y |
2.566 |
0.994 |
-1.099 |
| z |
-1.834 |
-1.099 |
-1.116 |
|
| Polar |
|---|
| 3z2-r2 | -2.231 |
|---|
| x2-y2 | -0.581 |
|---|
| xy | 2.566 |
|---|
| xz | -1.834 |
|---|
| yz | -1.099 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
201.369 |
| (<r2>)1/2 |
14.190 |