return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12S (Propane, 2-(ethylthio)-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-591.501493
Energy at 298.15K-591.514440
HF Energy-590.913534
Nuclear repulsion energy312.625426
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3173 3018 25.83      
2 A 3164 3010 16.57      
3 A 3154 3000 42.89      
4 A 3107 2955 14.43      
5 A 3091 2940 18.02      
6 A 3090 2940 2.61      
7 A 3082 2932 16.80      
8 A 1607 1529 6.76      
9 A 1605 1526 6.53      
10 A 1600 1522 12.31      
11 A 1585 1508 3.71      
12 A 1499 1426 6.61      
13 A 1495 1422 3.04      
14 A 1374 1307 16.35      
15 A 1363 1297 12.63      
16 A 1239 1179 7.55      
17 A 1152 1096 22.21      
18 A 1101 1047 1.24      
19 A 982 935 2.27      
20 A 892 848 1.01      
21 A 705 671 0.49      
22 A 626 595 1.78      
23 A 476 453 1.98      
24 A 384 365 0.17      
25 A 304 289 0.88      
26 A 259 247 0.11      
27 A 173 164 0.33      
28 A 3178 3024 19.36      
29 A 3170 3015 13.58      
30 A 3154 3000 4.90      
31 A 3146 2993 0.18      
32 A 3079 2929 11.07      
33 A 1596 1518 3.20      
34 A 1594 1516 5.94      
35 A 1586 1509 0.76      
36 A 1482 1409 9.93      
37 A 1395 1327 0.27      
38 A 1343 1278 0.24      
39 A 1147 1092 1.79      
40 A 1115 1061 0.01      
41 A 1003 954 1.42      
42 A 973 926 0.62      
43 A 840 799 4.84      
44 A 335 319 1.34      
45 A 257 245 0.04      
46 A 248 236 0.03      
47 A 83 79 1.95      
48 A 33 31 0.31      

Unscaled Zero Point Vibrational Energy (zpe) 36517.0 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 34738.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.15354 0.05162 0.05039

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.738 0.732 0.000
C2 1.446 -0.127 0.000
C3 -1.459 -1.052 1.281
C4 -1.459 -1.052 -1.281
S5 0.000 0.996 0.000
C6 -1.413 -0.185 0.000
H7 3.617 0.079 0.000
H8 -2.285 0.483 0.000
H9 -1.452 -0.419 2.173
H10 -1.452 -0.419 -2.173
H11 2.773 1.369 0.889
H12 2.773 1.369 -0.889
H13 1.426 -0.763 -0.890
H14 1.426 -0.763 0.890
H15 -2.372 -1.661 1.281
H16 -2.372 -1.661 -1.281
H17 -0.598 -1.727 1.320
H18 -0.598 -1.727 -1.320

Atom - Atom Distances (Å)
  C1 C2 C3 C4 S5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.55154.73694.73692.75074.25131.09535.02904.85844.85841.09411.09412.17882.17885.78565.78564.34914.3491
C21.55153.30683.30681.83082.85972.18083.78023.63393.63392.18842.18841.09381.09384.30894.30892.91172.9117
C34.73693.30682.56122.82241.54765.35582.16311.09413.51114.89155.33673.62182.92581.09692.78631.09502.8211
C44.73693.30682.56122.82241.54765.35582.16313.51111.09415.33674.89152.92583.62182.78631.09692.82111.0950
S52.75071.83082.82242.82241.84193.73162.34192.97232.97232.93582.93582.43312.43313.78503.78503.08493.0849
C64.25132.85971.54761.54761.84195.03721.09782.18592.18594.55304.55303.03113.03112.17672.17672.18782.1878
H71.09532.18085.35585.35583.73165.03725.91575.53765.53761.77971.77972.50992.50996.36626.36624.77124.7712
H85.02903.78022.16312.16312.34191.09785.91572.49592.49595.21145.21144.01434.01432.49892.49893.07783.0778
H94.85843.63391.09413.51112.97232.18595.53762.49594.34594.76435.51594.21683.17001.78423.78351.78013.8261
H104.85843.63393.51111.09412.97232.18595.53762.49594.34595.51594.76433.17004.21683.78351.78423.82611.7801
H111.09412.18844.89155.33672.93584.55301.77975.21144.76435.51591.77813.08582.52165.98346.35264.59705.0818
H121.09412.18845.33674.89152.93584.55301.77975.21145.51594.76431.77812.52163.08586.35265.98345.08184.5970
H132.17881.09383.62182.92582.43313.03112.50994.01434.21683.17003.08582.52161.77924.46523.92193.14732.2825
H142.17881.09382.92583.62182.43313.03112.50994.01433.17004.21682.52163.08581.77923.92194.46522.28253.1473
H155.78564.30891.09692.78633.78502.17676.36622.49891.78423.78355.98346.35264.46523.92192.56141.77563.1485
H165.78564.30892.78631.09693.78502.17676.36622.49893.78351.78426.35265.98343.92194.46522.56143.14851.7756
H174.34912.91171.09502.82113.08492.18784.77123.07781.78013.82614.59705.08183.14732.28251.77563.14852.6393
H184.34912.91172.82111.09503.08492.18784.77123.07783.82611.78015.08184.59702.28253.14733.14851.77562.6393

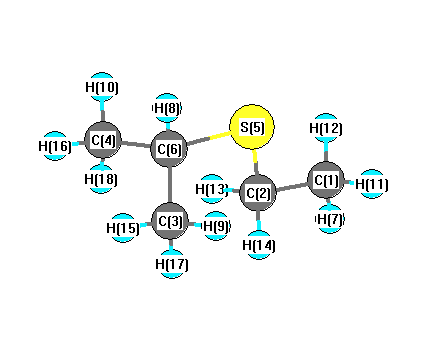
picture of Propane, 2-(ethylthio)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 S5 108.548 C1 C2 H13 109.691
C1 C2 H14 109.691 C2 C1 H7 109.763
C2 C1 H11 110.421 C2 C1 H12 110.421
C2 S5 C6 102.274 C3 C6 C4 111.687
C3 C6 S5 112.469 C3 C6 H8 108.510
C4 C6 S5 112.469 C4 C6 H8 108.510
S5 C2 H13 110.028 S5 C2 H14 110.028
S5 C6 H8 102.668 C6 C3 H9 110.507
C6 C3 H15 109.621 C6 C3 H17 110.596
C6 C4 H10 110.507 C6 C4 H16 109.621
C6 C4 H18 110.596 H7 C1 H11 108.752
H7 C1 H12 108.752 H9 C3 H15 109.045
H9 C3 H17 108.816 H10 C4 H16 109.045
H10 C4 H18 108.816 H11 C1 H12 108.688
H13 C2 H14 108.845 H15 C3 H17 108.206
H16 C4 H18 108.206
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability