return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (2,4-Hexadiene, (E,Z)-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-232.239246
Energy at 298.15K-232.248663
HF Energy-231.699202
Nuclear repulsion energy209.657436
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3184 3029 52.24      
2 A' 3171 3017 2.34      
3 A' 3162 3008 23.81      
4 A' 3156 3002 8.15      
5 A' 3141 2988 2.39      
6 A' 3139 2986 4.79      
7 A' 3077 2927 17.54      
8 A' 3069 2920 21.50      
9 A' 1703 1620 0.95      
10 A' 1660 1579 0.63      
11 A' 1596 1518 7.02      
12 A' 1589 1512 9.03      
13 A' 1493 1421 7.65      
14 A' 1492 1419 0.57      
15 A' 1488 1415 5.86      
16 A' 1391 1324 0.65      
17 A' 1376 1309 0.92      
18 A' 1333 1268 0.84      
19 A' 1196 1138 2.41      
20 A' 1125 1070 1.04      
21 A' 1053 1001 2.79      
22 A' 960 914 12.90      
23 A' 908 864 4.62      
24 A' 617 587 4.57      
25 A' 474 451 0.07      
26 A' 334 317 1.30      
27 A' 157 149 0.32      
28 A" 3130 2977 22.28      
29 A" 3129 2977 16.32      
30 A" 1589 1512 6.93      
31 A" 1584 1507 5.64      
32 A" 1127 1072 0.26      
33 A" 1125 1070 0.15      
34 A" 1018 969 26.55      
35 A" 983 935 17.16      
36 A" 830 790 12.04      
37 A" 724 689 21.05      
38 A" 437 416 5.11      
39 A" 197 187 0.80      
40 A" 190 181 0.82      
41 A" 105 100 0.06      
42 A" 89 85 0.64      

Unscaled Zero Point Vibrational Energy (zpe) 31649.6 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 30108.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.36092 0.04838 0.04336

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.316 2.983 0.000
H2 1.385 2.740 0.000
H3 0.100 3.592 0.886
H4 0.100 3.592 -0.886
C5 1.049 -2.506 0.000
H6 1.750 -1.667 0.000
H7 1.249 -3.122 -0.886
H8 1.249 -3.122 0.886
C9 -0.397 -2.032 0.000
H10 -1.141 -2.827 0.000
C11 -0.832 -0.752 0.000
H12 -1.910 -0.577 0.000
C13 0.000 0.463 0.000
H14 1.084 0.344 0.000
C15 -0.514 1.712 0.000
H16 -1.598 1.843 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 C11 H12 C13 H14 C15 H16
C11.09621.09711.09715.53784.86646.23866.23865.06555.99033.90724.19902.53982.74811.51842.2279
H21.09621.77861.77865.25744.42305.93005.93005.09436.11424.13664.67612.66572.41522.16013.1152
H31.09711.77861.77216.23535.58357.03846.81175.71566.59854.53054.71293.25423.50792.16782.5939
H41.09711.77861.77216.23535.58356.81177.03845.71566.59854.53054.71293.25423.50792.16782.5939
C55.53785.25746.23536.23531.09351.09711.09711.52092.21322.57203.53183.14882.85064.49815.0908
H64.86644.42305.58355.58351.09351.77471.77472.17753.11532.73983.81902.75702.11914.06764.8507
H76.23865.93007.03846.81171.09711.77471.77162.16362.56653.27624.15213.89793.58095.22065.7909
H86.23865.93006.81177.03841.09711.77471.77162.16362.56653.27624.15213.89793.58095.22065.7909
C95.06555.09435.71565.71561.52092.17752.16362.16361.08941.35242.09932.52632.79973.74574.0567
H105.99036.11426.59856.59852.21323.11532.56652.56651.08942.09852.37783.48253.87424.58214.6923
C113.90724.13664.53054.53052.57202.73983.27623.27621.35242.09851.09191.47232.20722.48382.7051
H124.19904.67614.71294.71293.53183.81904.15214.15212.09932.37781.09192.17493.13242.68102.4401
C132.53982.66573.25423.25423.14882.75703.89793.89792.52633.48251.47232.17491.09031.35052.1111
H142.74812.41523.50793.50792.85062.11913.58093.58092.79973.87422.20723.13241.09032.10323.0719
C151.51842.16012.16782.16784.49814.06765.22065.22063.74574.58212.48382.68101.35052.10321.0915
H162.22793.11522.59392.59395.09084.85075.79095.79094.05674.69232.70512.44012.11113.07191.0915

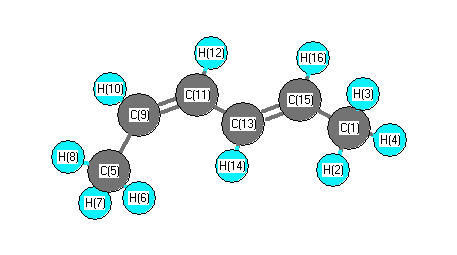
picture of 2,4-Hexadiene, (E,Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C15 C13 124.470 C1 C15 H16 116.258
H2 C1 H3 108.382 H2 C1 H4 108.382
H2 C1 C15 110.371 H3 C1 H4 107.739
H3 C1 C15 110.933 H4 C1 C15 110.933
C5 C9 H10 114.958 C5 C9 C11 126.946
H6 C5 H7 108.220 H6 C5 H8 108.220
H6 C5 C9 111.749 H7 C5 H8 107.686
H7 C5 C9 110.418 H8 C5 C9 110.418
C9 C11 H12 117.977 C9 C11 C13 126.801
H10 C9 C11 118.096 C11 C13 H14 118.163
C11 C13 C15 123.208 H12 C11 C13 115.222
C13 C15 H16 119.272 H14 C13 C15 118.629
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability