return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (2,4-Hexadiene, (E,E)-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-232.241240
Energy at 298.15K-232.250634
HF Energy-231.701710
Nuclear repulsion energy206.596153
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3160 3006 0.00      
2 Ag 3142 2989 0.00      
3 Ag 3130 2978 0.00      
4 Ag 3071 2921 0.00      
5 Ag 1708 1625 0.00      
6 Ag 1598 1520 0.00      
7 Ag 1493 1420 0.00      
8 Ag 1419 1350 0.00      
9 Ag 1373 1306 0.00      
10 Ag 1208 1149 0.00      
11 Ag 1085 1032 0.00      
12 Ag 970 923 0.00      
13 Ag 441 420 0.00      
14 Ag 342 326 0.00      
15 Au 3131 2978 40.56      
16 Au 1584 1507 12.97      
17 Au 1126 1071 0.28      
18 Au 1031 980 79.03      
19 Au 758 721 3.24      
20 Au 218 207 1.95      
21 Au 156 149 0.32      
22 Au 96 91 1.30      
23 Bg 3131 2978 0.00      
24 Bg 1584 1507 0.00      
25 Bg 1127 1073 0.00      
26 Bg 984 936 0.00      
27 Bg 845 804 0.00      
28 Bg 340 324 0.00      
29 Bg 179 171 0.00      
30 Bu 3160 3006 74.18      
31 Bu 3142 2989 10.19      
32 Bu 3137 2984 12.59      
33 Bu 3071 2921 45.14      
34 Bu 1671 1590 2.43      
35 Bu 1592 1514 14.22      
36 Bu 1492 1420 7.14      
37 Bu 1392 1324 2.22      
38 Bu 1336 1270 0.90      
39 Bu 1123 1068 9.46      
40 Bu 943 897 11.53      
41 Bu 562 535 0.04      
42 Bu 144 137 0.73      

Unscaled Zero Point Vibrational Energy (zpe) 31596.3 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 30057.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.76094 0.03964 0.03822

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.125 -3.011 0.000
C2 -1.125 3.011 0.000
C3 0.000 0.736 0.000
C4 0.000 -0.736 0.000
C5 -1.117 1.493 0.000
C6 1.117 -1.493 0.000
H7 0.100 -3.398 0.000
H8 -0.100 3.398 0.000
H9 1.642 -3.400 0.886
H10 1.642 -3.400 -0.886
H11 -1.642 3.400 0.886
H12 -1.642 3.400 -0.886
H13 0.977 1.226 0.000
H14 -0.977 -1.226 0.000
H15 -2.093 1.004 0.000
H16 2.093 -1.004 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C16.42963.91232.53895.03161.51851.09616.52511.09691.09697.03947.03944.23962.75855.14602.2284
C26.42962.53893.91231.51855.03166.52511.09617.03947.03941.09691.09692.75854.23962.22845.1460
C33.91232.53891.47101.34962.49274.13452.66414.53724.53723.25303.25301.09322.19112.11012.7216
C42.53893.91231.47102.49271.34962.66414.13453.25303.25304.53724.53722.19111.09322.72162.1101
C55.03161.51851.34962.49273.72925.03972.15965.68695.68692.16772.16772.11132.72211.09144.0669
C61.51855.03162.49271.34963.72922.15965.03972.16772.16775.68695.68692.72212.11134.06691.0914
H71.09616.52514.13452.66415.03972.15966.79841.77861.77867.07337.07334.70592.42444.91773.1150
H86.52511.09612.66414.13452.15965.03976.79847.07337.07331.77861.77862.42444.70593.11504.9177
H91.09697.03944.53723.25305.68692.16771.77867.07331.77227.55197.75714.75673.51765.84232.5944
H101.09697.03944.53723.25305.68692.16771.77867.07331.77227.75717.55194.75673.51765.84232.5944
H117.03941.09693.25304.53722.16775.68697.07331.77867.55197.75711.77223.51764.75672.59445.8423
H127.03941.09693.25304.53722.16775.68697.07331.77867.75717.55191.77223.51764.75672.59445.8423
H134.23962.75851.09322.19112.11132.72214.70592.42444.75674.75673.51763.51763.13493.07812.4933
H142.75854.23962.19111.09322.72212.11132.42444.70593.51763.51764.75674.75673.13492.49333.0781
H155.14602.22842.11012.72161.09144.06694.91773.11505.84235.84232.59442.59443.07812.49334.6428
H162.22845.14602.72162.11014.06691.09143.11504.91772.59442.59445.84235.84232.49333.07814.6428

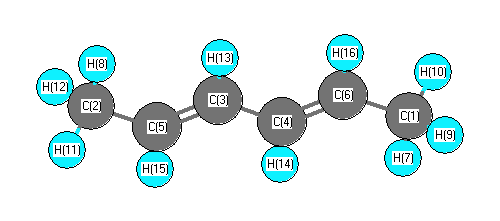
picture of 2,4-Hexadiene, (E,E)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C6 C4 124.444 C1 C6 H16 116.297
C2 C5 C3 124.444 C2 C5 H15 116.297
C3 C4 C6 124.138 C3 C4 H14 116.634
C3 C5 H15 119.259 C4 C3 C5 124.138
C4 C3 H13 116.634 C4 C6 H16 119.259
C5 C2 H8 110.331 C5 C2 H11 110.923
C5 C2 H12 110.923 C5 C3 H13 119.228
C6 C1 H7 110.331 C6 C1 H9 110.923
C6 C1 H10 110.923 C6 C4 H14 119.228
H7 C1 H9 108.399 H7 C1 H10 108.399
H8 C2 H11 108.399 H8 C2 H12 108.399
H9 C1 H10 107.767 H11 C2 H12 107.767
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability