return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CCH (phenylacetylene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-305.370370
Energy at 298.15K-305.375000
HF Energy-304.666103
Nuclear repulsion energy296.344891
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3482 3313 55.94      
2 A 3220 3064 10.76      
3 A 3199 3043 5.42      
4 A 3186 3031 1.26      
5 A 2129 2026 0.85      
6 A 1605 1526 3.41      
7 A 1535 1460 18.61      
8 A 1248 1187 0.02      
9 A 1214 1155 2.36      
10 A 1060 1008 2.53      
11 A 1020 971 0.59      
12 A 775 737 2.34      
13 A 483 460 0.02      
14 A 920 875 0.00      
15 A 823 783 0.00      
16 A 410 390 0.00      
17 A 928 883 0.19      
18 A 870 828 0.65      
19 A 723 688 117.80      
20 A 661 629 1.54      
21 A 583 555 13.72      
22 A 472 449 24.11      
23 A 297 282 0.27      
24 A 126 119 1.14      
25 A 3208 3052 18.76      
26 A 3190 3035 0.05      
27 A 1577 1500 3.18      
28 A 1491 1418 5.61      
29 A 1390 1323 0.21      
30 A 1302 1239 0.06      
31 A 1241 1180 0.00      
32 A 1104 1050 2.48      
33 A 676 643 40.36      
34 A 658 626 3.04      
35 A 543 516 4.30      
36 A 161 153 1.21      

Unscaled Zero Point Vibrational Energy (zpe) 23754.1 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 22597.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.18697 0.05010 0.03951

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.594
C2 0.000 0.000 2.038
C3 0.000 1.221 -0.120
C4 0.000 -1.221 -0.120
C5 0.000 1.217 -1.524
C6 0.000 -1.217 -1.524
C7 0.000 0.000 -2.230
C8 0.000 0.000 3.263
H9 0.000 0.000 4.328
H10 0.000 2.161 0.428
H11 0.000 -2.161 0.428
H12 0.000 2.160 -2.064
H13 0.000 -2.160 -2.064
H14 0.000 0.000 -3.317

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.44371.41481.41482.44272.44272.82422.66903.73372.16702.16703.42523.42523.9109
C21.44372.47942.47943.76383.76384.26791.22542.29002.69412.69414.63604.63605.3546
C31.41482.47942.44271.40392.81362.43813.59684.61231.08773.42622.15903.90043.4222
C41.41482.47942.44272.81361.40392.43813.59684.61233.42621.08773.90042.15903.4222
C52.44273.76381.40392.81362.43401.40714.93925.97682.16843.90131.08683.41992.1670
C62.44273.76382.81361.40392.43401.40714.93925.97683.90132.16843.41991.08682.1670
C72.82424.26792.43812.43811.40711.40715.49326.55783.42593.42592.16622.16621.0867
C82.66901.22543.59683.59684.93924.93925.49321.06463.56423.56425.74865.74866.5799
H93.73372.29004.61234.61235.97685.97686.55781.06464.45794.45796.74716.74717.6446
H102.16702.69411.08773.42622.16843.90133.42593.56424.45794.32132.49284.98814.3239
H112.16702.69413.42621.08773.90132.16843.42593.56424.45794.32134.98812.49284.3239
H123.42524.63602.15903.90041.08683.41992.16625.74866.74712.49284.98814.31982.4968
H133.42524.63603.90042.15903.41991.08682.16625.74866.74714.98812.49284.31982.4968
H143.91095.35463.42223.42222.16702.16701.08676.57997.64464.32394.32392.49682.4968

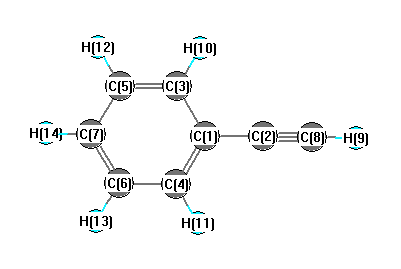
picture of phenylacetylene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C8 180.000 C1 C3 C5 120.135
C1 C3 H10 119.409 C1 C4 C6 120.135
C1 C4 H11 119.409 C2 C1 C3 120.312
C2 C1 C4 120.312 C2 C8 H9 180.000
C3 C1 C4 119.377 C3 C5 C7 120.306
C3 C5 H12 119.646 C4 C6 C7 120.306
C4 C6 H13 119.646 C5 C3 H10 120.457
C5 C7 C6 119.742 C5 C7 H14 120.129
C6 C4 H11 120.457 C6 C7 H14 120.129
C7 C5 H12 120.048 C7 C6 H13 120.048
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability