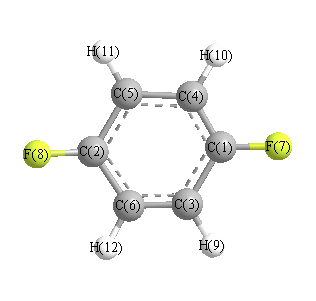
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3251 |
3093 |
0.00 |
219.25 |
0.16 |
0.28 |
| 2 |
Ag |
1643 |
1563 |
0.00 |
7.34 |
0.67 |
0.80 |
| 3 |
Ag |
1307 |
1243 |
0.00 |
13.04 |
0.24 |
0.38 |
| 4 |
Ag |
1216 |
1157 |
0.00 |
6.82 |
0.69 |
0.82 |
| 5 |
Ag |
862 |
820 |
0.00 |
29.89 |
0.11 |
0.20 |
| 6 |
Ag |
466 |
443 |
0.00 |
5.25 |
0.41 |
0.58 |
| 7 |
Au |
967 |
920 |
0.00 |
0.00 |
0.00 |
0.00 |
| 8 |
Au |
453 |
431 |
0.00 |
0.00 |
0.50 |
0.00 |
| 9 |
B1g |
845 |
804 |
0.00 |
6.61 |
0.75 |
0.86 |
| 10 |
B1u |
3234 |
3077 |
2.67 |
0.00 |
0.70 |
0.83 |
| 11 |
B1u |
1571 |
1494 |
173.36 |
0.00 |
0.44 |
0.61 |
| 12 |
B1u |
1288 |
1226 |
84.95 |
0.00 |
0.44 |
0.61 |
| 13 |
B1u |
1048 |
997 |
0.37 |
0.00 |
0.43 |
0.60 |
| 14 |
B1u |
757 |
720 |
31.68 |
0.00 |
0.00 |
0.00 |
| 15 |
B2g |
932 |
886 |
0.00 |
3.04 |
0.75 |
0.86 |
| 16 |
B2g |
710 |
676 |
0.00 |
0.03 |
0.75 |
0.86 |
| 17 |
B2g |
396 |
377 |
0.00 |
3.47 |
0.75 |
0.86 |
| 18 |
B2u |
3249 |
3091 |
0.26 |
0.00 |
0.34 |
0.51 |
| 19 |
B2u |
1439 |
1369 |
0.02 |
0.00 |
0.75 |
0.86 |
| 20 |
B2u |
1317 |
1253 |
0.90 |
0.00 |
0.75 |
0.86 |
| 21 |
B2u |
1137 |
1082 |
13.27 |
0.00 |
0.75 |
0.86 |
| 22 |
B2u |
331 |
315 |
5.05 |
0.00 |
0.00 |
0.00 |
| 23 |
B3g |
3235 |
3078 |
0.00 |
104.08 |
0.75 |
0.86 |
| 24 |
B3g |
1601 |
1523 |
0.00 |
6.95 |
0.75 |
0.86 |
| 25 |
B3g |
1350 |
1284 |
0.00 |
1.37 |
0.75 |
0.86 |
| 26 |
B3g |
671 |
638 |
0.00 |
7.02 |
0.75 |
0.86 |
| 27 |
B3g |
437 |
416 |
0.00 |
0.49 |
0.75 |
0.86 |
| 28 |
B3u |
864 |
822 |
130.82 |
0.00 |
0.00 |
0.00 |
| 29 |
B3u |
530 |
504 |
12.98 |
0.00 |
0.00 |
0.00 |
| 30 |
B3u |
163 |
155 |
2.05 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 18635.2 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 17727.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.