return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4Cl2 (1,3-dichlorobenzene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-1143.848167
Energy at 298.15K-1143.852328
HF Energy-1143.056762
Nuclear repulsion energy448.755236
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3235 3077 2.81      
2 A1 3231 3074 0.25      
3 A1 3207 3051 2.46      
4 A1 1577 1500 27.58      
5 A1 1444 1373 8.63      
6 A1 1152 1096 19.02      
7 A1 1104 1050 13.55      
8 A1 1030 980 0.36      
9 A1 695 661 1.60      
10 A1 400 380 3.31      
11 A1 199 189 0.51      
12 A2 881 838 0.00      
13 A2 535 509 0.00      
14 A2 207 197 0.00      
15 B1 929 883 0.07      
16 B1 849 808 9.99      
17 B1 772 734 79.61      
18 B1 654 622 5.00      
19 B1 445 423 5.10      
20 B1 170 162 0.11      
21 B2 3226 3069 0.15      
22 B2 1579 1502 65.86      
23 B2 1511 1437 44.33      
24 B2 1363 1297 2.00      
25 B2 1306 1242 0.06      
26 B2 1244 1183 0.88      
27 B2 1110 1056 23.63      
28 B2 796 757 66.18      
29 B2 439 418 5.40      
30 B2 371 353 0.44      

Unscaled Zero Point Vibrational Energy (zpe) 17827.9 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 16959.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.09266 0.02813 0.02158

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 2.095
C2 0.000 1.222 1.400
C3 0.000 -1.222 1.400
C4 0.000 1.204 -0.003
C5 0.000 -1.204 -0.003
C6 0.000 0.000 -0.720
Cl7 0.000 2.725 -0.890
Cl8 0.000 -2.725 -0.890
H9 0.000 0.000 3.181
H10 0.000 2.168 1.931
H11 0.000 -2.168 1.931
H12 0.000 0.000 -1.806

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 Cl7 Cl8 H9 H10 H11 H12
C11.40571.40572.41872.41872.81534.04114.04111.08622.17462.17463.9005
C21.40572.44371.40252.80242.44712.73864.56252.16011.08533.43163.4303
C31.40572.44372.80241.40252.44714.56252.73862.16013.43161.08533.4303
C42.41871.40252.80242.40881.40201.76014.02793.40392.16043.88772.1682
C52.41872.80241.40252.40881.40204.02791.76013.40393.88772.16042.1682
C62.81532.44712.44711.40201.40202.72992.72993.90143.42503.42501.0852
Cl74.04112.73864.56251.76014.02792.72995.44934.89842.87475.64772.8745
Cl84.04114.56252.73864.02791.76012.72995.44934.89845.64772.87472.8745
H91.08622.16012.16013.40393.40393.90144.89844.89842.50302.50304.9866
H102.17461.08533.43162.16043.88773.42502.87475.64772.50304.33684.3200
H112.17463.43161.08533.88772.16043.42505.64772.87472.50304.33684.3200
H123.90053.43033.43032.16822.16821.08522.87452.87454.98664.32004.3200

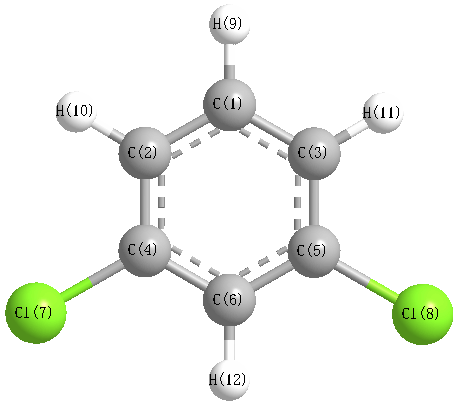
picture of 1,3-dichlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 118.922 C1 C2 H10 121.072
C1 C3 C5 118.922 C1 C3 H11 121.072
C2 C1 C3 120.730 C2 C1 H9 119.635
C2 C4 C6 121.507 C2 C4 Cl7 119.548
C3 C1 H9 119.635 C3 C5 C6 121.507
C3 C5 Cl8 119.548 C4 C2 H10 120.006
C4 C6 C5 118.414 C4 C6 H12 120.793
C5 C3 H11 120.006 C5 C6 H12 120.793
C6 C4 Cl7 118.946 C6 C5 Cl8 118.946
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability