Jump to
S1C2
Energy calculated at CID/3-21G*
| | hartrees |
|---|
| Energy at 0K | -320.487862 |
| Energy at 298.15K | -320.496611 |
| HF Energy | -319.922815 |
| Nuclear repulsion energy | 230.549491 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3208 |
2985 |
25.71 |
|
|
|
| 2 |
A' |
3166 |
2946 |
17.35 |
|
|
|
| 3 |
A' |
3151 |
2932 |
19.70 |
|
|
|
| 4 |
A' |
3134 |
2917 |
13.93 |
|
|
|
| 5 |
A' |
1647 |
1533 |
5.85 |
|
|
|
| 6 |
A' |
1628 |
1515 |
4.65 |
|
|
|
| 7 |
A' |
1618 |
1506 |
2.67 |
|
|
|
| 8 |
A' |
1539 |
1432 |
21.90 |
|
|
|
| 9 |
A' |
1528 |
1422 |
34.33 |
|
|
|
| 10 |
A' |
1475 |
1373 |
62.59 |
|
|
|
| 11 |
A' |
1420 |
1321 |
20.81 |
|
|
|
| 12 |
A' |
1208 |
1124 |
0.77 |
|
|
|
| 13 |
A' |
1086 |
1010 |
67.81 |
|
|
|
| 14 |
A' |
1052 |
979 |
12.48 |
|
|
|
| 15 |
A' |
960 |
893 |
136.43 |
|
|
|
| 16 |
A' |
901 |
839 |
252.70 |
|
|
|
| 17 |
A' |
653 |
608 |
2.00 |
|
|
|
| 18 |
A' |
390 |
363 |
1.35 |
|
|
|
| 19 |
A' |
362 |
337 |
0.25 |
|
|
|
| 20 |
A' |
153 |
142 |
0.21 |
|
|
|
| 21 |
A" |
3221 |
2997 |
56.97 |
|
|
|
| 22 |
A" |
3193 |
2971 |
0.23 |
|
|
|
| 23 |
A" |
3185 |
2963 |
13.37 |
|
|
|
| 24 |
A" |
1628 |
1515 |
9.17 |
|
|
|
| 25 |
A" |
1411 |
1313 |
0.14 |
|
|
|
| 26 |
A" |
1347 |
1254 |
0.12 |
|
|
|
| 27 |
A" |
1250 |
1163 |
0.79 |
|
|
|
| 28 |
A" |
973 |
906 |
1.83 |
|
|
|
| 29 |
A" |
816 |
759 |
2.85 |
|
|
|
| 30 |
A" |
247 |
230 |
0.00 |
|
|
|
| 31 |
A" |
182 |
170 |
1.95 |
|
|
|
| 32 |
A" |
102 |
95 |
1.94 |
|
|
|
| 33 |
A" |
4 |
4 |
0.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23919.0 cm
-1
Scaled (by 0.9305) Zero Point Vibrational Energy (zpe) 22256.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.934 |
2.242 |
0.000 |
| C2 |
-1.523 |
0.753 |
0.000 |
| C3 |
0.000 |
0.602 |
0.000 |
| O4 |
0.261 |
-0.846 |
0.000 |
| N5 |
1.718 |
-1.054 |
0.000 |
| O6 |
1.963 |
-2.237 |
0.000 |
| H7 |
-3.019 |
2.339 |
0.000 |
| H8 |
-1.546 |
2.749 |
0.885 |
| H9 |
-1.546 |
2.749 |
-0.885 |
| H10 |
-1.920 |
0.248 |
-0.880 |
| H11 |
-1.920 |
0.248 |
0.880 |
| H12 |
0.436 |
1.060 |
0.888 |
| H13 |
0.436 |
1.060 |
-0.888 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5440 | 2.5351 | 3.7884 | 4.9194 | 5.9361 | 1.0899 | 1.0914 | 1.0914 | 2.1793 | 2.1793 | 2.7935 | 2.7935 |
C2 | 1.5440 | | 1.5303 | 2.3961 | 3.7110 | 4.5922 | 2.1802 | 2.1833 | 2.1833 | 1.0899 | 1.0899 | 2.1726 | 2.1726 | C3 | 2.5351 | 1.5303 | | 1.4716 | 2.3868 | 3.4513 | 3.4831 | 2.7896 | 2.7896 | 2.1418 | 2.1418 | 1.0897 | 1.0897 | O4 | 3.7884 | 2.3961 | 1.4716 | | 1.4714 | 2.1973 | 4.5722 | 4.1199 | 4.1199 | 2.5944 | 2.5944 | 2.1092 | 2.1092 | N5 | 4.9194 | 3.7110 | 2.3868 | 1.4714 | | 1.2072 | 5.8272 | 5.0896 | 5.0896 | 3.9633 | 3.9633 | 2.6267 | 2.6267 | O6 | 5.9361 | 4.5922 | 3.4513 | 2.1973 | 1.2072 | | 6.7642 | 6.1605 | 6.1605 | 4.6930 | 4.6930 | 3.7392 | 3.7392 | H7 | 1.0899 | 2.1802 | 3.4831 | 4.5722 | 5.8272 | 6.7642 | | 1.7671 | 1.7671 | 2.5207 | 2.5207 | 3.7904 | 3.7904 | H8 | 1.0914 | 2.1833 | 2.7896 | 4.1199 | 5.0896 | 6.1605 | 1.7671 | | 1.7702 | 3.0840 | 2.5289 | 2.6048 | 3.1509 | H9 | 1.0914 | 2.1833 | 2.7896 | 4.1199 | 5.0896 | 6.1605 | 1.7671 | 1.7702 | | 2.5289 | 3.0840 | 3.1509 | 2.6048 | H10 | 2.1793 | 1.0899 | 2.1418 | 2.5944 | 3.9633 | 4.6930 | 2.5207 | 3.0840 | 2.5289 | | 1.7604 | 3.0557 | 2.4923 | H11 | 2.1793 | 1.0899 | 2.1418 | 2.5944 | 3.9633 | 4.6930 | 2.5207 | 2.5289 | 3.0840 | 1.7604 | | 2.4923 | 3.0557 | H12 | 2.7935 | 2.1726 | 1.0897 | 2.1092 | 2.6267 | 3.7392 | 3.7904 | 2.6048 | 3.1509 | 3.0557 | 2.4923 | | 1.7756 | H13 | 2.7935 | 2.1726 | 1.0897 | 2.1092 | 2.6267 | 3.7392 | 3.7904 | 3.1509 | 2.6048 | 2.4923 | 3.0557 | 1.7756 | |
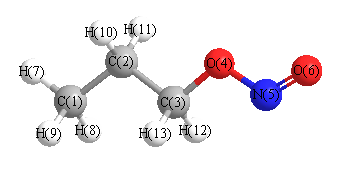 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.099 |
|
C1 |
C2 |
H10 |
110.478 |
| C1 |
C2 |
H11 |
110.478 |
|
C2 |
C1 |
H7 |
110.546 |
| C2 |
C1 |
H8 |
110.706 |
|
C2 |
C1 |
H9 |
110.706 |
| C2 |
C3 |
O4 |
105.899 |
|
C2 |
C3 |
H12 |
110.922 |
| C2 |
C3 |
H13 |
110.922 |
|
C3 |
C2 |
H10 |
108.482 |
| C3 |
C2 |
H11 |
108.482 |
|
C3 |
O4 |
N5 |
108.388 |
| O4 |
C3 |
H12 |
109.976 |
|
O4 |
C3 |
H13 |
109.976 |
| O4 |
N5 |
O6 |
109.846 |
|
H7 |
C1 |
H8 |
108.208 |
| H7 |
C1 |
H9 |
108.208 |
|
H8 |
C1 |
H9 |
108.376 |
| H10 |
C2 |
H11 |
107.722 |
|
H12 |
C3 |
H13 |
109.110 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/3-21G*
| | hartrees |
|---|
| Energy at 0K | -320.489746 |
| Energy at 298.15K | -320.498847 |
| HF Energy | -319.923751 |
| Nuclear repulsion energy | 238.861974 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3228 |
3004 |
24.86 |
|
|
|
| 2 |
A |
3213 |
2990 |
30.96 |
|
|
|
| 3 |
A |
3203 |
2981 |
26.08 |
|
|
|
| 4 |
A |
3188 |
2966 |
6.16 |
|
|
|
| 5 |
A |
3157 |
2938 |
23.77 |
|
|
|
| 6 |
A |
3148 |
2929 |
8.34 |
|
|
|
| 7 |
A |
3140 |
2921 |
18.98 |
|
|
|
| 8 |
A |
1637 |
1523 |
9.33 |
|
|
|
| 9 |
A |
1627 |
1514 |
8.32 |
|
|
|
| 10 |
A |
1614 |
1502 |
1.69 |
|
|
|
| 11 |
A |
1605 |
1494 |
3.56 |
|
|
|
| 12 |
A |
1539 |
1432 |
63.54 |
|
|
|
| 13 |
A |
1522 |
1416 |
12.02 |
|
|
|
| 14 |
A |
1478 |
1375 |
30.57 |
|
|
|
| 15 |
A |
1467 |
1365 |
16.81 |
|
|
|
| 16 |
A |
1389 |
1293 |
0.36 |
|
|
|
| 17 |
A |
1356 |
1262 |
5.61 |
|
|
|
| 18 |
A |
1253 |
1166 |
7.75 |
|
|
|
| 19 |
A |
1177 |
1095 |
12.21 |
|
|
|
| 20 |
A |
1099 |
1023 |
40.49 |
|
|
|
| 21 |
A |
1029 |
958 |
87.72 |
|
|
|
| 22 |
A |
997 |
928 |
26.90 |
|
|
|
| 23 |
A |
893 |
831 |
188.51 |
|
|
|
| 24 |
A |
887 |
825 |
106.10 |
|
|
|
| 25 |
A |
834 |
776 |
1.43 |
|
|
|
| 26 |
A |
621 |
578 |
2.30 |
|
|
|
| 27 |
A |
473 |
440 |
2.85 |
|
|
|
| 28 |
A |
375 |
349 |
3.00 |
|
|
|
| 29 |
A |
286 |
266 |
1.15 |
|
|
|
| 30 |
A |
266 |
247 |
2.85 |
|
|
|
| 31 |
A |
160 |
149 |
0.36 |
|
|
|
| 32 |
A |
115 |
107 |
2.07 |
|
|
|
| 33 |
A |
51 |
47 |
0.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24012.8 cm
-1
Scaled (by 0.9305) Zero Point Vibrational Energy (zpe) 22343.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/3-21G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.157 |
-0.957 |
0.189 |
| C2 |
-1.797 |
0.451 |
-0.316 |
| C3 |
-0.448 |
0.923 |
0.222 |
| O4 |
0.546 |
-0.023 |
-0.294 |
| N5 |
1.816 |
0.221 |
0.314 |
| O6 |
2.628 |
-0.552 |
-0.143 |
| H7 |
-3.120 |
-1.280 |
-0.210 |
| H8 |
-2.217 |
-0.972 |
1.280 |
| H9 |
-1.393 |
-1.668 |
-0.123 |
| H10 |
-1.753 |
0.457 |
-1.406 |
| H11 |
-2.559 |
1.174 |
-0.013 |
| H12 |
-0.212 |
1.932 |
-0.117 |
| H13 |
-0.436 |
0.907 |
1.312 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5387 | 2.5410 | 2.9005 | 4.1464 | 4.8132 | 1.0910 | 1.0923 | 1.0890 | 2.1689 | 2.1784 | 3.4960 | 2.7746 |
C2 | 1.5387 | | 1.5271 | 2.3911 | 3.6753 | 4.5404 | 2.1814 | 2.1791 | 2.1661 | 1.0910 | 1.0933 | 2.1780 | 2.1707 | C3 | 2.5410 | 1.5271 | | 1.4667 | 2.3725 | 3.4307 | 3.4899 | 2.8003 | 2.7797 | 2.1382 | 2.1393 | 1.0902 | 1.0904 | O4 | 2.9005 | 2.3911 | 1.4667 | | 1.4295 | 2.1528 | 3.8764 | 3.3189 | 2.5488 | 2.5990 | 3.3405 | 2.1049 | 2.1000 | N5 | 4.1464 | 3.6753 | 2.3725 | 1.4295 | | 1.2106 | 5.1861 | 4.3161 | 3.7502 | 3.9696 | 4.4904 | 2.6886 | 2.5569 | O6 | 4.8132 | 4.5404 | 3.4307 | 2.1528 | 1.2106 | | 5.7936 | 5.0670 | 4.1729 | 4.6696 | 5.4684 | 3.7733 | 3.6919 | H7 | 1.0910 | 2.1814 | 3.4899 | 3.8764 | 5.1861 | 5.7936 | | 1.7683 | 1.7716 | 2.5126 | 2.5253 | 4.3335 | 3.7820 | H8 | 1.0923 | 2.1791 | 2.8003 | 3.3189 | 4.3161 | 5.0670 | 1.7683 | | 1.7690 | 3.0770 | 2.5293 | 3.7956 | 2.5897 | H9 | 1.0890 | 2.1661 | 2.7797 | 2.5488 | 3.7502 | 4.1729 | 1.7716 | 1.7690 | | 2.5079 | 3.0743 | 3.7888 | 3.0993 | H10 | 2.1689 | 1.0910 | 2.1382 | 2.5990 | 3.9696 | 4.6696 | 2.5126 | 3.0770 | 2.5079 | | 1.7617 | 2.4925 | 3.0542 | H11 | 2.1784 | 1.0933 | 2.1393 | 3.3405 | 4.4904 | 5.4684 | 2.5253 | 2.5293 | 3.0743 | 1.7617 | | 2.4685 | 2.5180 | H12 | 3.4960 | 2.1780 | 1.0902 | 2.1049 | 2.6886 | 3.7733 | 4.3335 | 3.7956 | 3.7888 | 2.4925 | 2.4685 | | 1.7733 | H13 | 2.7746 | 2.1707 | 1.0904 | 2.1000 | 2.5569 | 3.6919 | 3.7820 | 2.5897 | 3.0993 | 3.0542 | 2.5180 | 1.7733 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.956 |
|
C1 |
C2 |
H10 |
109.965 |
| C1 |
C2 |
H11 |
110.582 |
|
C2 |
C1 |
H7 |
110.954 |
| C2 |
C1 |
H8 |
110.689 |
|
C2 |
C1 |
H9 |
109.858 |
| C2 |
C3 |
O4 |
105.992 |
|
C2 |
C3 |
H12 |
111.549 |
| C2 |
C3 |
H13 |
110.950 |
|
C3 |
C2 |
H10 |
108.365 |
| C3 |
C2 |
H11 |
108.316 |
|
C3 |
O4 |
N5 |
110.000 |
| O4 |
C3 |
H12 |
109.947 |
|
O4 |
C3 |
H13 |
109.539 |
| O4 |
N5 |
O6 |
108.975 |
|
H7 |
C1 |
H8 |
108.177 |
| H7 |
C1 |
H9 |
108.712 |
|
H8 |
C1 |
H9 |
108.378 |
| H10 |
C2 |
H11 |
107.522 |
|
H12 |
C3 |
H13 |
108.826 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
