Jump to
S1C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -320.581244 |
| Energy at 298.15K | |
| HF Energy | -319.910689 |
| Nuclear repulsion energy | 226.925536 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3164 |
3010 |
23.42 |
|
|
|
| 2 |
A' |
3114 |
2962 |
15.92 |
|
|
|
| 3 |
A' |
3085 |
2935 |
27.75 |
|
|
|
| 4 |
A' |
3082 |
2932 |
8.59 |
|
|
|
| 5 |
A' |
1618 |
1539 |
7.44 |
|
|
|
| 6 |
A' |
1605 |
1527 |
2.79 |
|
|
|
| 7 |
A' |
1594 |
1517 |
3.43 |
|
|
|
| 8 |
A' |
1498 |
1425 |
6.96 |
|
|
|
| 9 |
A' |
1446 |
1375 |
9.50 |
|
|
|
| 10 |
A' |
1385 |
1317 |
4.42 |
|
|
|
| 11 |
A' |
1182 |
1125 |
5.74 |
|
|
|
| 12 |
A' |
1156 |
1099 |
8.27 |
|
|
|
| 13 |
A' |
1036 |
986 |
4.49 |
|
|
|
| 14 |
A' |
994 |
946 |
36.42 |
|
|
|
| 15 |
A' |
926 |
881 |
43.09 |
|
|
|
| 16 |
A' |
786 |
747 |
463.31 |
|
|
|
| 17 |
A' |
591 |
563 |
27.11 |
|
|
|
| 18 |
A' |
364 |
346 |
0.00 |
|
|
|
| 19 |
A' |
344 |
328 |
3.52 |
|
|
|
| 20 |
A' |
144 |
137 |
0.05 |
|
|
|
| 21 |
A" |
3172 |
3017 |
51.75 |
|
|
|
| 22 |
A" |
3147 |
2993 |
0.50 |
|
|
|
| 23 |
A" |
3129 |
2976 |
18.93 |
|
|
|
| 24 |
A" |
1605 |
1527 |
8.22 |
|
|
|
| 25 |
A" |
1384 |
1316 |
0.20 |
|
|
|
| 26 |
A" |
1318 |
1254 |
0.00 |
|
|
|
| 27 |
A" |
1212 |
1153 |
0.22 |
|
|
|
| 28 |
A" |
955 |
909 |
1.53 |
|
|
|
| 29 |
A" |
802 |
763 |
2.88 |
|
|
|
| 30 |
A" |
248 |
236 |
0.00 |
|
|
|
| 31 |
A" |
185 |
176 |
1.91 |
|
|
|
| 32 |
A" |
99 |
95 |
1.06 |
|
|
|
| 33 |
A" |
50i |
48i |
0.54 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23159.0 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 22031.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.904 |
2.301 |
0.000 |
| C2 |
-1.524 |
0.798 |
0.000 |
| C3 |
0.000 |
0.612 |
0.000 |
| O4 |
0.218 |
-0.865 |
0.000 |
| N5 |
1.723 |
-1.080 |
0.000 |
| O6 |
1.967 |
-2.317 |
0.000 |
| H7 |
-2.992 |
2.422 |
0.000 |
| H8 |
-1.502 |
2.799 |
0.889 |
| H9 |
-1.502 |
2.799 |
-0.889 |
| H10 |
-1.934 |
0.301 |
-0.885 |
| H11 |
-1.934 |
0.301 |
0.885 |
| H12 |
0.447 |
1.064 |
0.893 |
| H13 |
0.447 |
1.064 |
-0.893 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5499 | 2.5450 | 3.8115 | 4.9585 | 6.0256 | 1.0944 | 1.0956 | 1.0956 | 2.1872 | 2.1872 | 2.8028 | 2.8028 |
C2 | 1.5499 | | 1.5357 | 2.4089 | 3.7515 | 4.6789 | 2.1884 | 2.1897 | 2.1897 | 1.0946 | 1.0946 | 2.1803 | 2.1803 | C3 | 2.5450 | 1.5357 | | 1.4934 | 2.4150 | 3.5282 | 3.4964 | 2.7980 | 2.7980 | 2.1497 | 2.1497 | 1.0956 | 1.0956 | O4 | 3.8115 | 2.4089 | 1.4934 | | 1.5203 | 2.2728 | 4.5942 | 4.1445 | 4.1445 | 2.6028 | 2.6028 | 2.1376 | 2.1376 | N5 | 4.9585 | 3.7515 | 2.4150 | 1.5203 | | 1.2608 | 5.8730 | 5.1224 | 5.1224 | 4.0082 | 4.0082 | 2.6494 | 2.6494 | O6 | 6.0256 | 4.6789 | 3.5282 | 2.2728 | 1.2608 | | 6.8587 | 6.2448 | 6.2448 | 4.7805 | 4.7805 | 3.8124 | 3.8124 | H7 | 1.0944 | 2.1884 | 3.4964 | 4.5942 | 5.8730 | 6.8587 | | 1.7756 | 1.7756 | 2.5297 | 2.5297 | 3.8036 | 3.8036 | H8 | 1.0956 | 2.1897 | 2.7980 | 4.1445 | 5.1224 | 6.2448 | 1.7756 | | 1.7780 | 3.0944 | 2.5356 | 2.6098 | 3.1599 | H9 | 1.0956 | 2.1897 | 2.7980 | 4.1445 | 5.1224 | 6.2448 | 1.7756 | 1.7780 | | 2.5356 | 3.0944 | 3.1599 | 2.6098 | H10 | 2.1872 | 1.0946 | 2.1497 | 2.6028 | 4.0082 | 4.7805 | 2.5297 | 3.0944 | 2.5356 | | 1.7694 | 3.0678 | 2.5005 | H11 | 2.1872 | 1.0946 | 2.1497 | 2.6028 | 4.0082 | 4.7805 | 2.5297 | 2.5356 | 3.0944 | 1.7694 | | 2.5005 | 3.0678 | H12 | 2.8028 | 2.1803 | 1.0956 | 2.1376 | 2.6494 | 3.8124 | 3.8036 | 2.6098 | 3.1599 | 3.0678 | 2.5005 | | 1.7852 | H13 | 2.8028 | 2.1803 | 1.0956 | 2.1376 | 2.6494 | 3.8124 | 3.8036 | 3.1599 | 2.6098 | 2.5005 | 3.0678 | 1.7852 | |
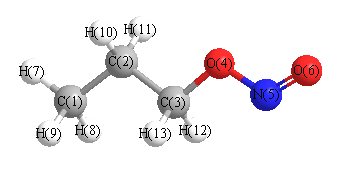 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.137 |
|
C1 |
C2 |
H10 |
110.416 |
| C1 |
C2 |
H11 |
110.416 |
|
C2 |
C1 |
H7 |
110.516 |
| C2 |
C1 |
H8 |
110.550 |
|
C2 |
C1 |
H9 |
110.550 |
| C2 |
C3 |
O4 |
105.352 |
|
C2 |
C3 |
H12 |
110.805 |
| C2 |
C3 |
H13 |
110.805 |
|
C3 |
C2 |
H10 |
108.462 |
| C3 |
C2 |
H11 |
108.462 |
|
C3 |
O4 |
N5 |
106.516 |
| O4 |
C3 |
H12 |
110.361 |
|
O4 |
C3 |
H13 |
110.361 |
| O4 |
N5 |
O6 |
109.259 |
|
H7 |
C1 |
H8 |
108.341 |
| H7 |
C1 |
H9 |
108.341 |
|
H8 |
C1 |
H9 |
108.465 |
| H10 |
C2 |
H11 |
107.852 |
|
H12 |
C3 |
H13 |
109.120 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -320.584581 |
| Energy at 298.15K | -320.593536 |
| HF Energy | -319.911597 |
| Nuclear repulsion energy | 235.556553 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3188 |
3033 |
19.22 |
|
|
|
| 2 |
A |
3163 |
3009 |
34.45 |
|
|
|
| 3 |
A |
3147 |
2994 |
19.89 |
|
|
|
| 4 |
A |
3136 |
2984 |
11.21 |
|
|
|
| 5 |
A |
3095 |
2944 |
18.42 |
|
|
|
| 6 |
A |
3090 |
2940 |
15.17 |
|
|
|
| 7 |
A |
3081 |
2931 |
19.15 |
|
|
|
| 8 |
A |
1616 |
1537 |
7.95 |
|
|
|
| 9 |
A |
1602 |
1524 |
7.23 |
|
|
|
| 10 |
A |
1583 |
1506 |
1.98 |
|
|
|
| 11 |
A |
1581 |
1504 |
2.05 |
|
|
|
| 12 |
A |
1489 |
1416 |
7.51 |
|
|
|
| 13 |
A |
1447 |
1376 |
6.35 |
|
|
|
| 14 |
A |
1431 |
1361 |
3.61 |
|
|
|
| 15 |
A |
1359 |
1292 |
0.26 |
|
|
|
| 16 |
A |
1320 |
1256 |
2.07 |
|
|
|
| 17 |
A |
1226 |
1167 |
0.88 |
|
|
|
| 18 |
A |
1177 |
1119 |
16.46 |
|
|
|
| 19 |
A |
1143 |
1087 |
9.79 |
|
|
|
| 20 |
A |
1062 |
1010 |
9.23 |
|
|
|
| 21 |
A |
973 |
926 |
46.36 |
|
|
|
| 22 |
A |
960 |
913 |
28.01 |
|
|
|
| 23 |
A |
870 |
828 |
2.21 |
|
|
|
| 24 |
A |
811 |
772 |
11.22 |
|
|
|
| 25 |
A |
776 |
738 |
417.40 |
|
|
|
| 26 |
A |
570 |
543 |
17.96 |
|
|
|
| 27 |
A |
448 |
426 |
13.69 |
|
|
|
| 28 |
A |
357 |
340 |
4.26 |
|
|
|
| 29 |
A |
278 |
264 |
3.17 |
|
|
|
| 30 |
A |
270 |
256 |
3.20 |
|
|
|
| 31 |
A |
169 |
161 |
0.36 |
|
|
|
| 32 |
A |
115 |
109 |
1.79 |
|
|
|
| 33 |
A |
63 |
60 |
0.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23297.4 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 22162.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.178 |
-0.962 |
0.190 |
| C2 |
-1.817 |
0.449 |
-0.315 |
| C3 |
-0.465 |
0.923 |
0.224 |
| O4 |
0.537 |
-0.026 |
-0.305 |
| N5 |
1.831 |
0.226 |
0.316 |
| O6 |
2.686 |
-0.547 |
-0.138 |
| H7 |
-3.147 |
-1.285 |
-0.205 |
| H8 |
-2.233 |
-0.977 |
1.286 |
| H9 |
-1.413 |
-1.676 |
-0.128 |
| H10 |
-1.776 |
0.458 |
-1.410 |
| H11 |
-2.582 |
1.176 |
-0.010 |
| H12 |
-0.236 |
1.942 |
-0.110 |
| H13 |
-0.454 |
0.902 |
1.320 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5420 | 2.5473 | 2.9140 | 4.1832 | 4.8926 | 1.0953 | 1.0965 | 1.0933 | 2.1770 | 2.1847 | 3.5064 | 2.7786 |
C2 | 1.5420 | | 1.5308 | 2.4018 | 3.7094 | 4.6154 | 2.1880 | 2.1844 | 2.1713 | 1.0957 | 1.0982 | 2.1844 | 2.1765 | C3 | 2.5473 | 1.5308 | | 1.4787 | 2.4020 | 3.4962 | 3.5002 | 2.8041 | 2.7890 | 2.1460 | 2.1448 | 1.0961 | 1.0963 | O4 | 2.9140 | 2.4018 | 1.4787 | | 1.4573 | 2.2174 | 3.8944 | 3.3324 | 2.5605 | 2.6086 | 3.3559 | 2.1234 | 2.1179 | N5 | 4.1832 | 3.7094 | 2.4020 | 1.4573 | | 1.2382 | 5.2284 | 4.3478 | 3.7868 | 4.0054 | 4.5265 | 2.7200 | 2.5867 | O6 | 4.8926 | 4.6154 | 3.4962 | 2.2174 | 1.2382 | | 5.8799 | 5.1386 | 4.2520 | 4.7469 | 5.5442 | 3.8379 | 3.7534 | H7 | 1.0953 | 2.1880 | 3.5002 | 3.8944 | 5.2284 | 5.8799 | | 1.7753 | 1.7790 | 2.5237 | 2.5316 | 4.3470 | 3.7891 | H8 | 1.0965 | 2.1844 | 2.8041 | 3.3324 | 4.3478 | 5.1386 | 1.7753 | | 1.7766 | 3.0879 | 2.5371 | 3.8023 | 2.5876 | H9 | 1.0933 | 2.1713 | 2.7890 | 2.5605 | 3.7868 | 4.2520 | 1.7790 | 1.7766 | | 2.5155 | 3.0840 | 3.8047 | 3.1083 | H10 | 2.1770 | 1.0957 | 2.1460 | 2.6086 | 4.0054 | 4.7469 | 2.5237 | 3.0879 | 2.5155 | | 1.7681 | 2.5034 | 3.0657 | H11 | 2.1847 | 1.0982 | 2.1448 | 3.3559 | 4.5265 | 5.5442 | 2.5316 | 2.5371 | 3.0840 | 1.7681 | | 2.4708 | 2.5245 | H12 | 3.5064 | 2.1844 | 1.0961 | 2.1234 | 2.7200 | 3.8379 | 4.3470 | 3.8023 | 3.8047 | 2.5034 | 2.4708 | | 1.7814 | H13 | 2.7786 | 2.1765 | 1.0963 | 2.1179 | 2.5867 | 3.7534 | 3.7891 | 2.5876 | 3.1083 | 3.0657 | 2.5245 | 1.7814 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.984 |
|
C1 |
C2 |
H10 |
110.094 |
| C1 |
C2 |
H11 |
110.552 |
|
C2 |
C1 |
H7 |
110.985 |
| C2 |
C1 |
H8 |
110.626 |
|
C2 |
C1 |
H9 |
109.787 |
| C2 |
C3 |
O4 |
105.878 |
|
C2 |
C3 |
H12 |
111.441 |
| C2 |
C3 |
H13 |
110.802 |
|
C3 |
C2 |
H10 |
108.449 |
| C3 |
C2 |
H11 |
108.216 |
|
C3 |
O4 |
N5 |
109.794 |
| O4 |
C3 |
H12 |
110.227 |
|
O4 |
C3 |
H13 |
109.779 |
| O4 |
N5 |
O6 |
110.432 |
|
H7 |
C1 |
H8 |
108.182 |
| H7 |
C1 |
H9 |
108.749 |
|
H8 |
C1 |
H9 |
108.444 |
| H10 |
C2 |
H11 |
107.403 |
|
H12 |
C3 |
H13 |
108.692 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
