Jump to
S1C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -1848.340822 |
| Energy at 298.15K | |
| Nuclear repulsion energy | 166.171575 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3072 |
2923 |
0.00 |
|
|
|
| 2 |
A1' |
1374 |
1308 |
0.00 |
|
|
|
| 3 |
A1' |
539 |
513 |
0.00 |
|
|
|
| 4 |
A1" |
13 |
13 |
0.00 |
|
|
|
| 5 |
A2" |
3071 |
2921 |
21.63 |
|
|
|
| 6 |
A2" |
1363 |
1296 |
20.09 |
|
|
|
| 7 |
A2" |
586 |
558 |
28.55 |
|
|
|
| 8 |
E' |
3136 |
2983 |
39.34 |
|
|
|
| 8 |
E' |
3136 |
2983 |
39.34 |
|
|
|
| 9 |
E' |
1535 |
1460 |
2.45 |
|
|
|
| 9 |
E' |
1535 |
1460 |
2.45 |
|
|
|
| 10 |
E' |
777 |
739 |
108.99 |
|
|
|
| 10 |
E' |
777 |
739 |
108.99 |
|
|
|
| 11 |
E' |
105i |
100i |
4.48 |
|
|
|
| 11 |
E' |
105i |
100i |
4.48 |
|
|
|
| 12 |
E" |
3137 |
2984 |
0.00 |
|
|
|
| 12 |
E" |
3137 |
2984 |
0.00 |
|
|
|
| 13 |
E" |
1534 |
1460 |
0.00 |
|
|
|
| 13 |
E" |
1534 |
1460 |
0.00 |
|
|
|
| 14 |
E" |
657 |
625 |
0.00 |
|
|
|
| 14 |
E" |
657 |
625 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15680.0 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 14916.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Zn1 |
0.000 |
0.000 |
0.000 |
| C2 |
0.000 |
0.000 |
1.919 |
| C3 |
0.000 |
0.000 |
-1.919 |
| H4 |
0.000 |
1.017 |
2.331 |
| H5 |
0.000 |
1.017 |
-2.331 |
| H6 |
-0.881 |
-0.508 |
2.331 |
| H7 |
0.881 |
-0.508 |
2.331 |
| H8 |
0.881 |
-0.508 |
-2.331 |
| H9 |
-0.881 |
-0.508 |
-2.331 |
Atom - Atom Distances (Å)
| |
Zn1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| Zn1 | | 1.9194 | 1.9194 | 2.5433 | 2.5433 | 2.5433 | 2.5433 | 2.5433 | 2.5433 |
C2 | 1.9194 | | 3.8388 | 1.0970 | 4.3705 | 1.0970 | 1.0970 | 4.3705 | 4.3705 | C3 | 1.9194 | 3.8388 | | 4.3705 | 1.0970 | 4.3705 | 4.3705 | 1.0970 | 1.0970 | H4 | 2.5433 | 1.0970 | 4.3705 | | 4.6624 | 1.7611 | 1.7611 | 4.9840 | 4.9840 | H5 | 2.5433 | 4.3705 | 1.0970 | 4.6624 | | 4.9840 | 4.9840 | 1.7611 | 1.7611 | H6 | 2.5433 | 1.0970 | 4.3705 | 1.7611 | 4.9840 | | 1.7611 | 4.9840 | 4.6624 | H7 | 2.5433 | 1.0970 | 4.3705 | 1.7611 | 4.9840 | 1.7611 | | 4.6624 | 4.9840 | H8 | 2.5433 | 4.3705 | 1.0970 | 4.9840 | 1.7611 | 4.9840 | 4.6624 | | 1.7611 | H9 | 2.5433 | 4.3705 | 1.0970 | 4.9840 | 1.7611 | 4.6624 | 4.9840 | 1.7611 | |
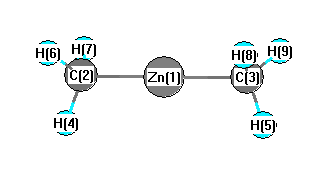 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Zn1 |
C2 |
H4 |
112.049 |
|
Zn1 |
C2 |
H6 |
112.049 |
| Zn1 |
C2 |
H7 |
112.049 |
|
Zn1 |
C3 |
H5 |
112.049 |
| Zn1 |
C3 |
H8 |
112.049 |
|
Zn1 |
C3 |
H9 |
112.049 |
| C2 |
Zn1 |
C3 |
180.000 |
|
H4 |
C2 |
H6 |
106.774 |
| H4 |
C2 |
H7 |
106.774 |
|
H5 |
C3 |
H8 |
106.774 |
| H5 |
C3 |
H9 |
106.774 |
|
H6 |
C2 |
H7 |
106.774 |
| H8 |
C3 |
H9 |
106.774 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -1848.341952 |
| Energy at 298.15K | -1848.347343 |
| Nuclear repulsion energy | 166.580770 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3131 |
2978 |
27.21 |
|
|
|
| 2 |
A1 |
3067 |
2917 |
7.17 |
|
|
|
| 3 |
A1 |
1544 |
1469 |
2.77 |
|
|
|
| 4 |
A1 |
1367 |
1300 |
1.44 |
|
|
|
| 5 |
A1 |
780 |
742 |
112.63 |
|
|
|
| 6 |
A1 |
553 |
526 |
1.93 |
|
|
|
| 7 |
A1 |
130 |
124 |
2.88 |
|
|
|
| 8 |
A2 |
3149 |
2996 |
0.00 |
|
|
|
| 9 |
A2 |
1534 |
1459 |
0.00 |
|
|
|
| 10 |
A2 |
628 |
597 |
0.00 |
|
|
|
| 11 |
A2 |
37 |
35 |
0.00 |
|
|
|
| 12 |
B1 |
3149 |
2995 |
38.94 |
|
|
|
| 13 |
B1 |
1536 |
1461 |
0.89 |
|
|
|
| 14 |
B1 |
754 |
717 |
109.76 |
|
|
|
| 15 |
B1 |
52 |
50 |
2.89 |
|
|
|
| 16 |
B2 |
3131 |
2978 |
15.10 |
|
|
|
| 17 |
B2 |
3065 |
2916 |
11.35 |
|
|
|
| 18 |
B2 |
1538 |
1463 |
1.67 |
|
|
|
| 19 |
B2 |
1352 |
1286 |
14.71 |
|
|
|
| 20 |
B2 |
664 |
632 |
5.48 |
|
|
|
| 21 |
B2 |
599 |
569 |
24.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15879.3 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 15106.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Zn1 |
0.000 |
0.000 |
0.148 |
| C2 |
0.000 |
1.885 |
-0.207 |
| C3 |
0.000 |
-1.885 |
-0.207 |
| H4 |
0.000 |
1.990 |
-1.301 |
| H5 |
0.000 |
-1.990 |
-1.301 |
| H6 |
-0.881 |
2.419 |
0.164 |
| H7 |
0.881 |
2.419 |
0.164 |
| H8 |
0.881 |
-2.419 |
0.164 |
| H9 |
-0.881 |
-2.419 |
0.164 |
Atom - Atom Distances (Å)
| |
Zn1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| Zn1 | | 1.9177 | 1.9177 | 2.4622 | 2.4622 | 2.5751 | 2.5751 | 2.5751 | 2.5751 |
C2 | 1.9177 | | 3.7691 | 1.0993 | 4.0265 | 1.0957 | 1.0957 | 4.4089 | 4.4089 | C3 | 1.9177 | 3.7691 | | 4.0265 | 1.0993 | 4.4089 | 4.4089 | 1.0957 | 1.0957 | H4 | 2.4622 | 1.0993 | 4.0265 | | 3.9808 | 1.7628 | 1.7628 | 4.7297 | 4.7297 | H5 | 2.4622 | 4.0265 | 1.0993 | 3.9808 | | 4.7297 | 4.7297 | 1.7628 | 1.7628 | H6 | 2.5751 | 1.0957 | 4.4089 | 1.7628 | 4.7297 | | 1.7628 | 5.1500 | 4.8389 | H7 | 2.5751 | 1.0957 | 4.4089 | 1.7628 | 4.7297 | 1.7628 | | 4.8389 | 5.1500 | H8 | 2.5751 | 4.4089 | 1.0957 | 4.7297 | 1.7628 | 5.1500 | 4.8389 | | 1.7628 | H9 | 2.5751 | 4.4089 | 1.0957 | 4.7297 | 1.7628 | 4.8389 | 5.1500 | 1.7628 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Zn1 |
C2 |
H4 |
106.198 |
|
Zn1 |
C2 |
H6 |
114.651 |
| Zn1 |
C2 |
H7 |
114.651 |
|
Zn1 |
C3 |
H5 |
106.198 |
| Zn1 |
C3 |
H8 |
114.651 |
|
Zn1 |
C3 |
H9 |
114.651 |
| C2 |
Zn1 |
C3 |
158.657 |
|
H4 |
C2 |
H6 |
106.850 |
| H4 |
C2 |
H7 |
106.850 |
|
H5 |
C3 |
H8 |
106.850 |
| H5 |
C3 |
H9 |
106.850 |
|
H6 |
C2 |
H7 |
107.109 |
| H8 |
C3 |
H9 |
107.109 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
