Jump to
S1C2
Energy calculated at MP2=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -93.045344 |
| Energy at 298.15K | -93.046626 |
| HF Energy | -92.859842 |
| Nuclear repulsion energy | 28.501209 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3407 |
3229 |
36.92 |
|
|
|
| 2 |
A' |
2978 |
2823 |
546.32 |
|
|
|
| 3 |
A' |
2567 |
2433 |
186.29 |
|
|
|
| 4 |
A' |
1036 |
982 |
45.26 |
|
|
|
| 5 |
A' |
760 |
721 |
481.23 |
|
|
|
| 6 |
A" |
1009 |
957 |
29.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5878.6 cm
-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 5573.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.099 |
0.635 |
0.000 |
| N2 |
0.099 |
-0.563 |
0.000 |
| H3 |
-0.672 |
1.430 |
0.000 |
| H4 |
-0.613 |
-1.298 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.1987 | 1.1066 | 2.0602 |
N2 | 1.1987 | | 2.1368 | 1.0228 | H3 | 1.1066 | 2.1368 | | 2.7286 | H4 | 2.0602 | 1.0228 | 2.7286 | |
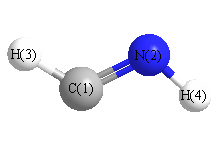 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
135.922 |
|
H3 |
C1 |
N2 |
135.886 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -93.052105 |
| Energy at 298.15K | -93.053444 |
| HF Energy | -92.866425 |
| Nuclear repulsion energy | 28.422072 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3507 |
3325 |
70.28 |
|
|
|
| 2 |
A' |
3011 |
2855 |
354.94 |
|
|
|
| 3 |
A' |
2586 |
2452 |
325.48 |
|
|
|
| 4 |
A' |
1227 |
1163 |
14.63 |
|
|
|
| 5 |
A' |
846 |
802 |
560.60 |
|
|
|
| 6 |
A" |
1038 |
984 |
225.45 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6107.3 cm
-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 5789.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.007 |
0.646 |
0.000 |
| N2 |
0.007 |
-0.565 |
0.000 |
| H3 |
-0.897 |
1.278 |
0.000 |
| H4 |
0.809 |
-1.195 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2112 | 1.1029 | 2.0080 |
N2 | 1.2112 | | 2.0529 | 1.0198 | H3 | 1.1029 | 2.0529 | | 3.0043 | H4 | 2.0080 | 1.0198 | 3.0043 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
128.121 |
|
H3 |
C1 |
N2 |
124.961 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
