return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6S (Thiophene, 2-methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-587.983694
Energy at 298.15K-587.989872
HF Energy-587.410723
Nuclear repulsion energy270.593259
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3282 3123 1.51      
2 A 3246 3088 2.75      
3 A 3227 3070 4.42      
4 A 3154 3001 9.41      
5 A 3080 2930 17.74      
6 A 1605 1527 4.54      
7 A 1542 1467 1.90      
8 A 1496 1423 0.91      
9 A 1439 1369 8.64      
10 A 1403 1334 0.12      
11 A 1309 1245 14.79      
12 A 1187 1129 16.69      
13 A 1153 1096 1.62      
14 A 1076 1024 1.08      
15 A 1024 974 1.98      
16 A 877 834 13.93      
17 A 758 721 1.33      
18 A 667 635 2.72      
19 A 565 537 1.33      
20 A 311 296 0.98      
21 A 3146 2993 12.70      
22 A 1583 1506 8.68      
23 A 1123 1069 1.07      
24 A 860 818 0.01      
25 A 807 767 11.66      
26 A 710 676 86.55      
27 A 578 550 0.51      
28 A 481 458 5.89      
29 A 228 217 3.67      
30 A 96 92 0.13      

Unscaled Zero Point Vibrational Energy (zpe) 21006.5 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 19983.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.17288 0.10217 0.06501

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -2.395 -1.572 0.000
C2 -1.447 -1.046 0.000
H3 -2.154 1.076 0.000
C4 -1.315 0.389 0.000
C5 0.000 0.814 0.000
H6 -0.030 -2.749 0.000
C7 -0.224 -1.686 0.000
S8 1.091 -0.551 0.000
H9 -0.333 2.925 0.000
H10 1.126 2.442 0.890
H11 1.126 2.442 -0.890
C12 0.520 2.237 0.000

Atom - Atom Distances (Å)
  H1 C2 H3 C4 C5 H6 C7 S8 H9 H10 H11 C12
H11.08352.65852.23783.37992.64192.17413.63214.94665.41325.41324.7965
C21.08352.23701.44092.35652.21561.38052.58604.12444.42564.42563.8280
H32.65852.23701.08472.17024.37563.36983.63002.59523.66363.66362.9160
C42.23781.44091.08471.38223.39092.34412.58292.71983.31233.31232.6053
C53.37992.35652.17021.38223.56302.50941.74672.13742.17112.17111.5160
H62.64192.21564.37563.39093.56301.08102.46795.68225.39285.39285.0170
C72.17411.38053.36982.34412.50941.08101.73694.61204.43374.43373.9932
S83.63212.58603.63002.58291.74672.46791.73693.75573.12273.12272.8457
H94.94664.12442.59522.71982.13745.68224.61203.75571.77591.77591.0958
H105.41324.42563.66363.31232.17115.39284.43373.12271.77591.77931.0958
H115.41324.42563.66363.31232.17115.39284.43373.12271.77591.77931.0958
C124.79653.82802.91602.60531.51605.01703.99322.84571.09581.09581.0958

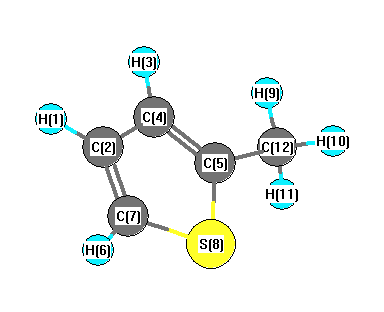
picture of Thiophene, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C4 124.256 H1 C2 C7 123.398
C2 C4 H3 124.074 C2 C4 C5 113.160
C2 C7 H6 127.928 C2 C7 S8 111.587
H3 C4 C5 122.766 C4 C2 C7 112.346
C4 C5 S8 110.742 C4 C5 C12 127.984
C5 S8 C7 92.165 C5 C12 H9 108.774
C5 C12 H10 111.446 C5 C12 H11 111.446
H6 C7 S8 120.485 S8 C5 C12 121.274
H9 C12 H10 108.258 H9 C12 H11 108.258
H10 C12 H11 108.559
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability