return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (2,3-dimethylbut-1-ene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-233.434132
Energy at 298.15K-233.446516
HF Energy-232.882340
Nuclear repulsion energy242.107671
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3243 3085 19.00      
2 A' 3162 3008 47.14      
3 A' 3160 3006 3.39      
4 A' 3152 2998 9.02      
5 A' 3150 2996 53.09      
6 A' 3081 2931 14.16      
7 A' 3076 2926 9.78      
8 A' 3067 2917 14.25      
9 A' 1697 1614 3.89      
10 A' 1609 1531 6.46      
11 A' 1607 1529 9.18      
12 A' 1596 1519 7.67      
13 A' 1528 1453 0.22      
14 A' 1496 1423 5.55      
15 A' 1483 1411 8.15      
16 A' 1394 1327 2.68      
17 A' 1324 1260 0.56      
18 A' 1241 1181 2.89      
19 A' 1160 1103 13.48      
20 A' 1063 1011 1.58      
21 A' 974 926 0.34      
22 A' 894 851 1.34      
23 A' 719 684 0.96      
24 A' 527 501 1.70      
25 A' 454 432 0.36      
26 A' 337 321 0.30      
27 A' 288 274 0.20      
28 A' 265 252 0.16      
29 A" 3158 3004 4.15      
30 A" 3149 2995 18.90      
31 A" 3142 2989 13.23      
32 A" 3076 2926 17.66      
33 A" 1602 1524 0.33      
34 A" 1590 1513 0.74      
35 A" 1582 1505 8.55      
36 A" 1473 1402 5.41      
37 A" 1401 1333 1.57      
38 A" 1157 1101 1.94      
39 A" 1120 1065 2.23      
40 A" 1001 952 0.66      
41 A" 974 927 1.13      
42 A" 935 889 45.28      
43 A" 758 721 0.68      
44 A" 559 532 8.04      
45 A" 279 265 0.07      
46 A" 221 210 0.56      
47 A" 162 154 0.26      
48 A" 27 26 0.06      

Unscaled Zero Point Vibrational Energy (zpe) 37055.6 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 35251.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.14520 0.08225 0.08137

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.769 -1.074 0.000
C2 -0.486 -0.668 0.000
C3 -0.127 0.820 0.000
C4 0.665 -1.671 0.000
C5 0.665 1.214 1.276
C6 0.665 1.214 -1.276
H7 -2.038 -2.128 0.000
H8 -2.593 -0.364 0.000
H9 -1.073 1.381 0.000
H10 0.273 -2.694 0.000
H11 1.297 -1.545 -0.887
H12 1.297 -1.545 0.887
H13 0.834 2.297 1.294
H14 0.107 0.929 2.175
H15 1.639 0.714 1.292
H16 0.834 2.297 -1.294
H17 0.107 0.929 -2.175
H18 1.639 0.714 -1.292

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.34572.50702.50653.57643.57641.08741.08752.55172.60633.22623.22624.45173.50184.06014.45173.50184.0601
C21.34571.53041.52732.54872.54872.13062.12822.13062.16382.17632.17633.49412.76262.84553.49412.76262.8455
C32.50701.53042.61371.55281.55283.51312.73551.09983.53652.89942.89942.18642.19002.19092.18642.19002.1909
C42.50651.52732.61373.15463.15462.74153.51033.51181.09581.09611.09614.17743.43562.88194.17743.43562.8819
C53.57642.54871.55283.15462.55254.48373.83842.16274.12983.56252.85711.09651.09561.09532.79463.50752.7921
C63.57642.54871.55283.15462.55254.48373.83842.16274.12982.85713.56252.79463.50752.79211.09651.09561.0953
H71.08742.13063.51312.74154.48374.48371.84903.63882.37893.49953.49955.43174.32164.82365.43174.32164.8236
H81.08752.12822.73553.51033.83843.83841.84902.31393.69314.16054.16054.52763.70004.55444.52763.70004.5544
H92.55172.13061.09983.51182.16272.16273.63882.31394.29103.86803.86802.48002.51493.07742.48002.51493.0774
H102.60632.16383.53651.09584.12984.12982.37893.69314.29101.77641.77645.18684.22923.89235.18684.22923.8923
H113.22622.17632.89941.09613.56252.85713.49954.16053.86801.77641.77414.44264.11273.15733.89163.03282.3203
H123.22622.17632.89941.09612.85713.56253.49954.16053.86801.77641.77413.89163.03282.32034.44264.11273.1573
H134.45173.49412.18644.17741.09652.79465.43174.52762.48005.18684.44263.89161.78191.77682.58863.79933.1379
H143.50182.76262.19003.43561.09563.50754.32163.70002.51494.22924.11273.03281.78191.78183.79934.34963.7968
H154.06012.84552.19092.88191.09532.79214.82364.55443.07743.89233.15732.32031.77681.78183.13793.79682.5843
H164.45173.49412.18644.17742.79461.09655.43174.52762.48005.18683.89164.44262.58863.79933.13791.78191.7768
H173.50182.76262.19003.43563.50751.09564.32163.70002.51494.22923.03284.11273.79934.34963.79681.78191.7818
H184.06012.84552.19092.88192.79211.09534.82364.55443.07743.89232.32033.15733.13793.79682.58431.77681.7818

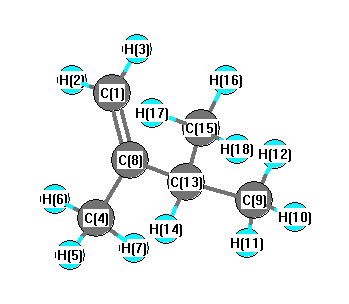
picture of 2,3-dimethylbut-1-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.173 C1 C2 C4 121.356
C2 C1 H7 121.892 C2 C1 H8 121.647
C2 C3 C5 111.502 C2 C3 C6 111.502
C2 C3 H9 107.065 C2 C4 H10 110.067
C2 C4 H11 111.039 C2 C4 H12 111.039
C3 C2 C4 117.471 C3 C5 H13 110.039
C3 C5 H14 110.376 C3 C5 H15 110.463
C3 C6 H16 110.039 C3 C6 H17 110.376
C3 C6 H18 110.463 C5 C3 C6 110.549
C5 C3 H9 108.017 C6 C3 H9 108.017
H7 C1 H8 116.461 H10 C4 H11 108.274
H10 C4 H12 108.274 H11 C4 H12 108.050
H13 C5 H14 108.756 H13 C5 H15 108.319
H14 C5 H15 108.836 H16 C6 H17 108.756
H16 C6 H18 108.319 H17 C6 H18 108.836
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability