return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HO2CCHNH2CH2OH (Serine)

using model chemistry: MP2=FULL/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/3-21G*
 hartrees
Energy at 0K-395.251783
Energy at 298.15K-395.261853
HF Energy-394.497635
Nuclear repulsion energy321.384250
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3541 3356 1.26      
2 A 3503 3321 5.04      
3 A 3498 3316 72.09      
4 A 3439 3260 0.74      
5 A 3186 3020 18.90      
6 A 3122 2960 5.86      
7 A 3032 2874 55.70      
8 A 1768 1676 32.16      
9 A 1699 1611 99.90      
10 A 1609 1525 0.73      
11 A 1477 1401 69.87      
12 A 1436 1361 2.25      
13 A 1423 1349 6.73      
14 A 1389 1317 36.02      
15 A 1383 1311 11.66      
16 A 1276 1209 14.73      
17 A 1216 1153 6.31      
18 A 1168 1107 25.24      
19 A 1115 1057 199.82      
20 A 1051 996 73.36      
21 A 1031 977 45.07      
22 A 983 932 29.19      
23 A 843 799 203.67      
24 A 792 751 54.41      
25 A 713 676 42.92      
26 A 647 613 123.39      
27 A 623 591 235.49      
28 A 562 532 1.67      
29 A 530 503 51.79      
30 A 416 394 6.19      
31 A 314 298 6.92      
32 A 297 282 10.90      
33 A 254 241 30.37      
34 A 240 227 20.63      
35 A 173 164 0.99      
36 A 48 45 1.11      

Unscaled Zero Point Vibrational Energy (zpe) 24895.8 cm-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 23601.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/3-21G*
ABC
0.11691 0.07807 0.04942

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.734 -0.583 0.010
O2 2.056 -0.441 -0.347
O3 0.267 -1.597 0.544
C4 -0.050 0.691 -0.319
C5 -1.487 0.526 0.195
O6 -2.135 -0.627 -0.389
N7 0.557 1.929 0.221
H8 2.570 -1.259 -0.131
H9 -0.085 0.774 -1.410
H10 -1.450 0.485 1.296
H11 -2.044 1.420 -0.089
H12 -1.708 -1.436 -0.026
H13 1.466 2.125 -0.203
H14 0.658 1.894 1.239

Atom - Atom Distances (Å)
  C1 O2 O3 C4 C5 O6 N7 H8 H9 H10 H11 H12 H13 H14
C11.37691.23731.53192.48922.89662.52731.96162.12792.75053.42612.58682.81292.7660
O21.37692.30872.39143.71234.19512.86130.98982.68103.98144.50993.90652.63643.1500
O31.23732.30872.46622.77622.75363.55302.42323.09242.80223.85312.06223.98063.5809
C41.53192.39142.46621.53472.46751.48083.27151.09532.14702.13522.71312.08962.0911
C52.48923.71232.77621.53471.44552.47964.44412.14551.10251.09091.98713.38132.7496
O62.89664.19512.75362.46751.44553.76214.75412.68452.13162.07070.98454.53564.0989
N72.52732.86133.55301.48082.47963.76213.78652.09992.69642.66874.06441.02161.0232
H81.96160.98982.42323.27154.44414.75413.78653.57974.60875.33534.28293.55943.9335
H92.12792.68103.09241.09532.14552.68452.09993.57973.04482.44993.07142.38522.9706
H102.75053.98142.80222.14701.10252.13162.69644.60873.04481.77312.34663.66582.5359
H113.42614.50993.85312.13521.09092.07072.66875.33532.44991.77312.87673.58193.0473
H122.58683.90652.06222.71311.98710.98454.06444.28293.07142.34662.87674.77344.2762
H132.81292.63643.98062.08963.38134.53561.02163.55942.38523.66583.58194.77341.6686
H142.76603.15003.58092.09112.74964.09891.02323.93352.97062.53593.04734.27621.6686

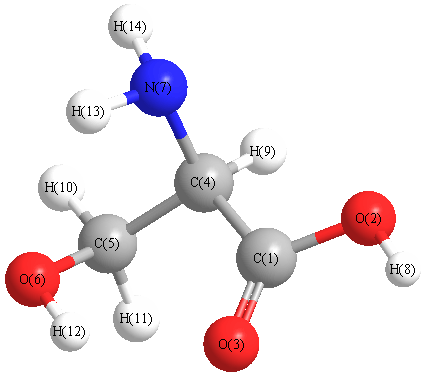
picture of Serine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 H8 110.893 C1 C4 C5 108.525
C1 C4 N7 114.033 C1 C4 H9 107.008
O2 C1 O3 123.957 O2 C1 C4 110.476
O3 C1 C4 125.554 C4 C5 O6 111.745
C4 C5 H10 107.877 C4 C5 H11 107.626
C4 N7 H13 111.935 C4 N7 H14 111.957
C5 C4 N7 110.620 C5 C4 H9 108.168
C5 O6 H12 108.217 O6 C5 H10 112.872
O6 C5 H11 108.646 N7 C4 H9 108.277
H10 C5 H11 107.878 H13 N7 H14 109.385
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability