return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H6O (2,5-Cyclohexadienone)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-304.473936
Energy at 298.15K-304.480538
HF Energy-303.822687
Nuclear repulsion energy265.689844
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3218 3061 0.15      
2 A1 3182 3027 5.49      
3 A1 3054 2905 4.50      
4 A1 1690 1608 24.37      
5 A1 1608 1530 29.86      
6 A1 1547 1472 9.93      
7 A1 1451 1381 1.56      
8 A1 1254 1192 3.26      
9 A1 979 931 2.35      
10 A1 860 818 5.35      
11 A1 744 708 2.46      
12 A1 514 489 0.92      
13 A2 1274 1212 0.00      
14 A2 1024 974 0.00      
15 A2 767 730 0.00      
16 A2 369 351 0.00      
17 B1 3078 2928 6.56      
18 B1 1036 986 2.55      
19 B1 999 950 15.21      
20 B1 869 827 51.86      
21 B1 592 563 33.81      
22 B1 305 290 1.55      
23 B1 118 112 0.45      
24 B2 3215 3059 19.86      
25 B2 3181 3026 11.08      
26 B2 1645 1565 3.67      
27 B2 1439 1369 11.43      
28 B2 1385 1318 5.41      
29 B2 1291 1228 23.28      
30 B2 1169 1112 29.05      
31 B2 982 934 10.64      
32 B2 586 557 2.02      
33 B2 438 416 10.43      

Unscaled Zero Point Vibrational Energy (zpe) 22930.7 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 21813.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.17279 0.08609 0.05808

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.126
C2 0.000 0.000 -1.857
C3 0.000 1.259 0.327
C4 0.000 -1.259 0.327
C5 0.000 1.267 -1.022
C6 0.000 -1.267 -1.022
O7 0.000 0.000 2.387
H8 0.000 2.184 0.897
H9 0.000 -2.184 0.897
H10 0.000 2.209 -1.566
H11 0.000 -2.209 -1.566
H12 0.880 0.000 -2.519
H13 -0.880 0.000 -2.519

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 O7 H8 H9 H10 H11 H12 H13
C12.98321.49091.49092.49322.49321.26072.19612.19613.48223.48223.74983.7498
C22.98322.52142.52141.51751.51754.24403.51503.51502.22842.22841.10141.1014
C31.49092.52142.51791.34882.86322.41371.08643.48982.11813.95123.23443.2344
C41.49092.52142.51792.86321.34882.41373.48981.08643.95122.11813.23443.2344
C52.49321.51751.34882.86322.53333.63602.12653.94811.08843.51832.14992.1499
C62.49321.51752.86321.34882.53333.63603.94812.12653.51831.08842.14992.1499
O71.26074.24402.41372.41373.63603.63602.64382.64384.52794.52794.98404.9840
H82.19613.51501.08643.48982.12653.94812.64384.36812.46275.03644.14894.1489
H92.19613.51503.48981.08643.94812.12652.64384.36815.03642.46274.14894.1489
H103.48222.22842.11813.95121.08843.51834.52792.46275.03644.41852.56222.5622
H113.48222.22843.95122.11813.51831.08844.52795.03642.46274.41852.56222.5622
H123.74981.10143.23443.23442.14992.14994.98404.14894.14892.56222.56221.7609
H133.74981.10143.23443.23442.14992.14994.98404.14894.14892.56222.56221.7609

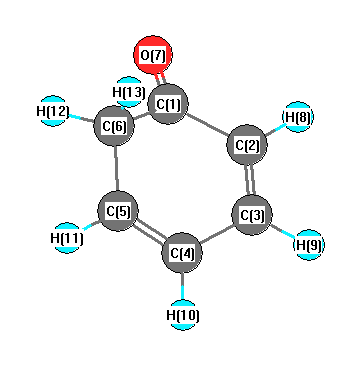
picture of 2,5-Cyclohexadienone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C5 122.719 C1 C3 H8 115.988
C1 C4 C6 122.719 C1 C4 H9 115.988
C2 C5 C3 123.089 C2 C5 H10 116.585
C2 C6 C4 123.089 C2 C6 H11 116.585
C3 C1 C4 115.217 C3 C1 O7 122.391
C3 C5 H10 120.326 C4 C1 O7 122.391
C4 C6 H11 120.326 C5 C2 C6 113.166
C5 C2 H12 109.322 C5 C2 H13 109.322
C5 C3 H8 121.293 C6 C2 H12 109.322
C6 C2 H13 109.322 C6 C4 H9 121.293
H12 C2 H13 106.146
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability