return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H5NO (Isoxazole, 5-methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-282.637631
Energy at 298.15K-282.643921
HF Energy-282.032113
Nuclear repulsion energy218.314701
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3311 3150 0.23      
2 A 3284 3124 1.73      
3 A 3177 3023 6.13      
4 A 3163 3009 7.08      
5 A 3096 2946 7.81      
6 A 1621 1542 8.61      
7 A 1579 1502 7.93      
8 A 1578 1501 21.96      
9 A 1499 1426 4.40      
10 A 1402 1334 16.58      
11 A 1348 1282 14.58      
12 A 1233 1173 11.20      
13 A 1174 1117 6.53      
14 A 1138 1082 4.01      
15 A 1070 1018 2.64      
16 A 1045 994 3.45      
17 A 947 901 6.73      
18 A 923 878 4.02      
19 A 867 825 8.72      
20 A 846 805 9.33      
21 A 808 769 43.93      
22 A 649 617 1.05      
23 A 641 610 1.08      
24 A 607 577 0.63      
25 A 326 310 1.74      
26 A 241 229 3.47      
27 A 92 87 0.73      

Unscaled Zero Point Vibrational Energy (zpe) 18831.5 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 17914.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.29358 0.11345 0.08311

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.144 0.016 -0.000
H2 -2.497 -0.517 0.889
H3 -2.566 1.025 -0.000
H4 -2.497 -0.518 -0.888
C5 -0.645 0.108 -0.000
O6 0.061 -1.109 0.000
N7 1.519 -0.783 -0.000
C8 1.543 0.566 -0.000
H9 2.501 1.063 0.000
C10 0.236 1.158 0.000
H11 -0.004 2.209 -0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 O6 N7 C8 H9 C10 H11
C11.09461.09381.09461.50172.47513.74883.72724.76172.64013.0641
H21.09461.78091.77672.14742.77204.12194.27585.31703.32693.7994
H31.09381.78091.78092.12873.38464.46724.13425.06782.80562.8224
H41.09461.77671.78092.14742.77174.12174.27585.31713.32713.7998
C51.50172.14742.12872.14741.40732.34022.23493.28791.37102.1966
O62.47512.77203.38462.77171.40731.49432.23683.26722.27473.3194
N73.74884.12194.46724.12172.34021.49431.34932.09072.32683.3576
C83.72724.27584.13424.27582.23492.23681.34931.07981.43422.2567
H94.76175.31705.06785.31713.28793.26722.09071.07982.26722.7557
C102.64013.32692.80563.32711.37102.27472.32681.43422.26721.0780
H113.06413.79942.82243.79982.19663.31943.35762.25672.75571.0780

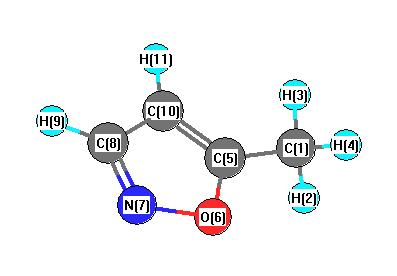
picture of Isoxazole, 5-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 O6 116.570 C1 C5 C10 133.523
H2 C1 H3 108.935 H2 C1 H4 108.498
H2 C1 C5 110.623 H3 C1 H4 108.935
H3 C1 C5 109.190 H4 C1 C5 110.621
C5 O6 N7 107.474 C5 C10 C8 105.608
C5 C10 H11 127.104 O6 C5 C10 109.907
O6 N7 C8 103.624 N7 C8 H9 118.370
N7 C8 C10 113.388 C8 C10 H11 127.288
H9 C8 C10 128.242
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability