return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHCHCH3 (2-Butene, (Z)-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-155.606367
Energy at 298.15K-155.614074
HF Energy-155.242047
Nuclear repulsion energy116.665070
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3188 3033 55.12      
2 A 3171 3017 0.00      
3 A 3080 2930 9.22      
4 A 1711 1628 0.96      
5 A 1599 1521 6.22      
6 A 1501 1428 8.08      
7 A 1353 1287 0.40      
8 A 1071 1018 1.62      
9 A 856 814 0.47      
10 A 311 295 0.03      
11 A 3131 2979 0.00      
12 A 1590 1512 0.00      
13 A 1125 1070 0.00      
14 A 1024 974 0.00      
15 A 387 368 0.00      
16 A 122 117 0.00      
17 A 3132 2979 40.08      
18 A 1594 1517 11.45      
19 A 1134 1078 0.48      
20 A 718 683 45.87      
21 A 100 95 0.68      
22 A 3168 3013 21.66      
23 A 3149 2996 0.01      
24 A 3077 2928 23.76      
25 A 1595 1517 3.85      
26 A 1489 1417 5.74      
27 A 1481 1408 0.01      
28 A 1195 1137 3.29      
29 A 978 931 8.39      
30 A 590 561 6.59      

Unscaled Zero Point Vibrational Energy (zpe) 24309.3 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 23125.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.53041 0.16790 0.13394

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.673 0.677
C2 0.000 -0.673 0.677
C3 0.000 1.596 -0.532
C4 0.000 -1.596 -0.532
H5 0.000 1.184 1.640
H6 0.000 -1.184 1.640
H7 0.000 1.035 -1.471
H8 0.000 -1.035 -1.471
H9 -0.886 2.243 -0.520
H10 0.886 2.243 -0.520
H11 0.886 -2.243 -0.520
H12 -0.886 -2.243 -0.520

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.34581.52132.57091.08982.09172.17832.74422.16362.16363.27383.2738
C21.34582.57091.52132.09171.08982.74422.17833.27383.27382.16362.1636
C31.52132.57093.19142.21063.52771.09332.79311.09701.09703.93943.9394
C42.57091.52133.19143.52772.21062.79311.09333.93943.93941.09701.0970
H51.08982.09172.21063.52772.36833.11413.82092.56272.56274.14624.1462
H62.09171.08983.52772.21062.36833.82093.11414.14624.14622.56272.5627
H72.17832.74421.09332.79313.11413.82092.06991.77431.77433.52603.5260
H82.74422.17832.79311.09333.82093.11412.06993.52603.52601.77431.7743
H92.16363.27381.09703.93942.56274.14621.77433.52601.77144.82274.4856
H102.16363.27381.09703.93942.56274.14621.77433.52601.77144.48564.8227
H113.27382.16363.93941.09704.14622.56273.52601.77434.82274.48561.7714
H123.27382.16363.93941.09704.14622.56273.52601.77434.48564.82271.7714

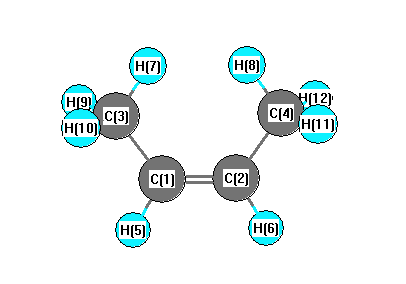
picture of 2-Butene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 127.344 C1 C2 H6 117.975
C1 C3 H7 111.799 C1 C3 H9 110.403
C1 C3 H10 110.403 C2 C1 C3 127.344
C2 C1 H5 117.975 C2 C4 H8 111.799
C2 C4 H11 110.403 C2 C4 H12 110.403
C3 C1 H5 114.681 C4 C2 H6 114.681
H7 C3 H9 108.208 H7 C3 H10 108.208
H8 C4 H11 108.208 H8 C4 H12 108.208
H9 C3 H10 107.687 H11 C4 H12 107.687
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability