return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,4-Pentadiene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-193.318263
Energy at 298.15K-193.325826
HF Energy-192.870275
Nuclear repulsion energy154.368312
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3256 3097 15.56      
2 A 3180 3025 3.13      
3 A 3166 3012 3.43      
4 A 3059 2910 17.34      
5 A 1696 1613 0.08      
6 A 1562 1486 7.47      
7 A 1533 1459 0.03      
8 A 1374 1307 0.00      
9 A 1315 1251 0.00      
10 A 1125 1070 0.34      
11 A 1047 996 17.54      
12 A 964 917 43.06      
13 A 929 884 5.64      
14 A 684 651 8.48      
15 A 382 363 0.30      
16 A 313 297 0.49      
17 A 84 80 0.04      
18 A 3256 3097 20.15      
19 A 3179 3024 8.44      
20 A 3167 3013 21.16      
21 A 3107 2956 10.34      
22 A 1682 1600 8.39      
23 A 1524 1450 1.87      
24 A 1376 1309 0.58      
25 A 1361 1295 1.67      
26 A 1201 1142 2.79      
27 A 1047 996 18.28      
28 A 965 918 28.63      
29 A 962 915 13.99      
30 A 925 880 6.69      
31 A 616 586 14.21      
32 A 463 440 1.80      
33 A 88 83 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 25292.2 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 24060.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.62576 0.07839 0.07769

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.706
C2 0.000 1.252 -0.166
C3 0.000 -1.252 -0.166
C4 0.998 2.148 -0.206
C5 -0.998 -2.148 -0.206
H6 -0.889 -0.010 1.352
H7 0.889 0.010 1.352
H8 -0.876 1.379 -0.801
H9 0.876 -1.379 -0.801
H10 0.964 3.018 -0.854
H11 -0.964 -3.018 -0.854
H12 1.885 2.041 0.415
H13 -1.885 -2.041 0.415

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.52521.52522.53802.53801.09921.09922.22232.22233.53173.53172.79352.7935
C21.52522.50371.34223.54382.16452.15321.08952.84462.12704.43152.12443.8384
C31.52522.50373.54381.34222.15322.16452.84461.08954.43152.12703.83842.1244
C42.53801.34223.54384.73753.26252.64772.11173.57901.08585.56461.08765.1232
C52.53803.54381.34224.73752.64773.26253.57902.11175.56461.08585.12321.0876
H61.09922.16452.15323.26252.64771.77862.56173.10244.17973.73143.57482.4486
H71.09922.15322.16452.64773.26251.77863.10242.56173.73144.17972.44863.5748
H82.22231.08952.84462.11173.57902.56173.10243.26732.46514.39853.08873.7672
H92.22232.84461.08953.57902.11173.10242.56173.26734.39852.46513.76723.0887
H103.53172.12704.43151.08585.56464.17973.73142.46514.39856.33721.84765.9435
H113.53174.43152.12705.56461.08583.73144.17974.39852.46516.33725.94351.8476
H122.79352.12443.83841.08765.12323.57482.44863.08873.76721.84765.94355.5566
H132.79353.83842.12445.12321.08762.44863.57483.76723.08875.94351.84765.5566

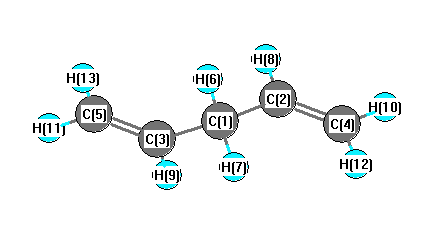
picture of 1,4-Pentadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 124.406 C1 C2 H8 115.388
C1 C3 C5 124.406 C1 C3 H9 115.388
C2 C1 C3 110.322 C2 C1 H6 110.061
C2 C1 H7 109.182 C2 C4 H10 121.975
C2 C4 H12 121.581 C3 C1 H6 109.182
C3 C1 H7 110.061 C3 C5 H11 121.975
C3 C5 H13 121.581 C4 C2 H8 120.189
C5 C3 H9 120.189 H6 C1 H7 107.999
H10 C4 H12 116.444 H11 C5 H13 116.444
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2/3-21G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.511      
2 C -0.198      
3 C -0.198      
4 C -0.419      
5 C -0.419      
6 H 0.228      
7 H 0.228      
8 H 0.229      
9 H 0.229      
10 H 0.211      
11 H 0.211      
12 H 0.204      
13 H 0.204      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 7.236 2.681 0.000
y 2.681 8.364 0.000
z 0.000 0.000 4.754


<r2> (average value of r2) Å2
<r2> 159.216
(<r2>)1/2 12.618