return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (1,5-Hexadiene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1Ag
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-232.229349
Energy at 298.15K-232.239154
HF Energy-231.689375
Nuclear repulsion energy211.156771
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3255 3097 8.11      
2 A 3174 3019 0.11      
3 A 3161 3007 35.71      
4 A 3129 2977 16.99      
5 A 3065 2916 1.83      
6 A 1689 1607 3.81      
7 A 1570 1494 1.55      
8 A 1531 1456 1.73      
9 A 1404 1335 1.42      
10 A 1379 1312 0.41      
11 A 1294 1231 0.46      
12 A 1128 1073 0.13      
13 A 1050 999 6.44      
14 A 1012 963 3.66      
15 A 976 928 0.66      
16 A 960 913 12.38      
17 A 802 763 0.77      
18 A 639 608 5.97      
19 A 440 418 0.00      
20 A 353 336 0.11      
21 A 93 89 0.00      
22 A 72 69 0.01      
23 B 3255 3097 28.96      
24 B 3173 3019 8.85      
25 B 3160 3006 6.58      
26 B 3108 2957 0.25      
27 B 3073 2923 36.13      
28 B 1691 1609 4.06      
29 B 1581 1504 7.46      
30 B 1533 1459 0.16      
31 B 1386 1319 0.91      
32 B 1363 1296 0.06      
33 B 1333 1268 0.11      
34 B 1216 1156 3.24      
35 B 1050 999 29.83      
36 B 1007 958 3.39      
37 B 961 914 72.65      
38 B 925 880 0.10      
39 B 678 645 24.11      
40 B 445 424 0.99      
41 B 231 220 0.19      
42 B 103 98 0.15      

Unscaled Zero Point Vibrational Energy (zpe) 31723.3 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 30178.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.39343 0.04704 0.04537

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.435 0.649 -0.328
C2 0.435 -0.649 -0.328
C3 0.435 1.894 -0.360
C4 -0.435 -1.894 -0.360
C5 0.529 2.780 0.644
C6 -0.529 -2.780 0.644
H7 -1.104 0.632 -1.199
H8 -1.058 0.655 0.576
H9 1.104 -0.632 -1.199
H10 1.058 -0.655 0.576
H11 1.032 2.033 -1.262
H12 -1.032 -2.033 -1.262
H13 1.180 3.647 0.589
H14 -0.051 2.668 1.557
H15 -1.180 -3.647 0.589
H16 0.051 -2.668 1.557

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.56291.51872.54392.53253.56601.09891.09772.18362.17852.22222.90283.52632.78804.45613.8460
C21.56292.54391.51873.56602.53252.18362.17851.09891.09772.90282.22224.45613.84603.52632.7880
C31.51872.54393.88701.34274.87742.15992.15422.74442.78631.09054.28822.12832.12395.84954.9634
C42.54391.51873.88704.87741.34272.74442.78632.15992.15424.28821.09055.84954.96342.12832.1239
C52.53253.56601.34274.87745.66033.26802.65343.92033.47642.10835.40731.08591.08756.65115.5446
C63.56602.53254.87741.34275.66033.92033.47643.26802.65345.40732.10836.65115.54461.08591.0875
H71.09892.18362.15992.74443.26803.92031.77522.54453.07902.55562.66664.18433.58464.63784.4514
H81.09772.17852.15422.78632.65343.47641.77523.07902.48883.10573.25653.73702.45524.30393.6377
H92.18361.09892.74442.15993.92033.26802.54453.07901.77522.66662.55564.63784.45144.18433.5846
H102.17851.09772.78632.15423.47642.65343.07902.48881.77523.25653.10574.30393.63773.73702.4552
H112.22222.90281.09054.28822.10835.40732.55563.10572.66663.25654.56002.46023.08616.37075.5685
H122.90282.22224.28821.09055.40732.10832.66663.25652.55563.10574.56006.37075.56852.46023.0861
H133.52634.45612.12835.84951.08596.65114.18433.73704.63784.30392.46026.37071.84787.66696.4877
H142.78803.84602.12394.96341.08755.54463.58462.45524.45143.63773.08615.56851.84786.48775.3363
H154.45613.52635.84952.12836.65111.08594.63784.30394.18433.73706.37072.46027.66696.48771.8478
H163.84602.78804.96342.12395.54461.08754.45143.63773.58462.45525.56853.08616.48775.33631.8478

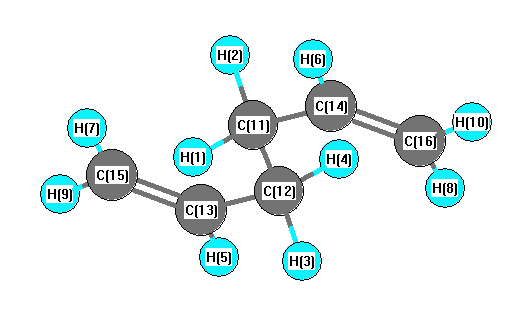
picture of 1,5-Hexadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 111.278 C1 C2 H9 108.999
C1 C2 H10 108.666 C1 C3 C5 124.401
C1 C3 H11 115.825 C2 C1 C3 111.278
C2 C1 H7 108.999 C2 C1 H8 108.666
C2 C4 C6 124.401 C2 C4 H12 115.825
C3 C1 H7 110.182 C3 C1 H8 109.799
C3 C5 H13 122.052 C3 C5 H14 121.485
C4 C2 H9 110.182 C4 C2 H10 109.799
C4 C6 H15 122.052 C4 C6 H16 121.485
C5 C3 H11 119.748 C6 C4 H12 119.748
H7 C1 H8 107.836 H9 C2 H10 107.836
H13 C5 H14 116.462 H15 C6 H16 116.462
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability