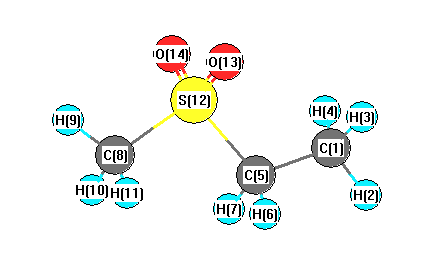
Vibrational Frequencies calculated at B3PW91/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3180 |
3051 |
1.97 |
|
|
|
| 2 |
A |
3169 |
3040 |
1.35 |
|
|
|
| 3 |
A |
3163 |
3035 |
4.49 |
|
|
|
| 4 |
A |
3145 |
3017 |
11.51 |
|
|
|
| 5 |
A |
3129 |
3002 |
4.24 |
|
|
|
| 6 |
A |
3081 |
2956 |
3.02 |
|
|
|
| 7 |
A |
3079 |
2954 |
2.29 |
|
|
|
| 8 |
A |
3071 |
2947 |
11.78 |
|
|
|
| 9 |
A |
1565 |
1501 |
12.14 |
|
|
|
| 10 |
A |
1560 |
1497 |
16.82 |
|
|
|
| 11 |
A |
1516 |
1455 |
16.95 |
|
|
|
| 12 |
A |
1515 |
1454 |
9.13 |
|
|
|
| 13 |
A |
1502 |
1441 |
6.20 |
|
|
|
| 14 |
A |
1462 |
1402 |
12.24 |
|
|
|
| 15 |
A |
1420 |
1362 |
34.78 |
|
|
|
| 16 |
A |
1389 |
1333 |
159.71 |
|
|
|
| 17 |
A |
1334 |
1280 |
21.32 |
|
|
|
| 18 |
A |
1302 |
1249 |
8.23 |
|
|
|
| 19 |
A |
1169 |
1122 |
132.70 |
|
|
|
| 20 |
A |
1099 |
1054 |
1.60 |
|
|
|
| 21 |
A |
1088 |
1043 |
17.86 |
|
|
|
| 22 |
A |
1017 |
975 |
22.82 |
|
|
|
| 23 |
A |
1004 |
963 |
2.86 |
|
|
|
| 24 |
A |
993 |
952 |
19.18 |
|
|
|
| 25 |
A |
827 |
794 |
63.43 |
|
|
|
| 26 |
A |
743 |
713 |
47.82 |
|
|
|
| 27 |
A |
663 |
636 |
16.91 |
|
|
|
| 28 |
A |
506 |
486 |
17.85 |
|
|
|
| 29 |
A |
443 |
425 |
21.30 |
|
|
|
| 30 |
A |
405 |
388 |
12.46 |
|
|
|
| 31 |
A |
315 |
302 |
1.78 |
|
|
|
| 32 |
A |
297 |
285 |
1.13 |
|
|
|
| 33 |
A |
236 |
226 |
2.53 |
|
|
|
| 34 |
A |
217 |
208 |
1.11 |
|
|
|
| 35 |
A |
198 |
190 |
0.26 |
|
|
|
| 36 |
A |
101 |
97 |
2.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25450.0 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 24416.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.656 |
|
|
|
| 2 |
H |
0.256 |
|
|
|
| 3 |
H |
0.235 |
|
|
|
| 4 |
H |
0.219 |
|
|
|
| 5 |
C |
-0.728 |
|
|
|
| 6 |
H |
0.279 |
|
|
|
| 7 |
H |
0.253 |
|
|
|
| 8 |
C |
-0.916 |
|
|
|
| 9 |
H |
0.252 |
|
|
|
| 10 |
H |
0.279 |
|
|
|
| 11 |
H |
0.255 |
|
|
|
| 12 |
S |
1.219 |
|
|
|
| 13 |
O |
-0.474 |
|
|
|
| 14 |
O |
-0.473 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.750 |
2.696 |
-2.008 |
4.344 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.111 |
-4.747 |
-1.539 |
| y |
-4.747 |
-41.538 |
-0.135 |
| z |
-1.539 |
-0.135 |
-47.319 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.317 |
-4.747 |
-1.539 |
| y |
-4.747 |
4.177 |
-0.135 |
| z |
-1.539 |
-0.135 |
-4.494 |
|
| Polar |
|---|
| 3z2-r2 | -8.989 |
|---|
| x2-y2 | -2.573 |
|---|
| xy | -4.747 |
|---|
| xz | -1.539 |
|---|
| yz | -0.135 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
190.200 |
| (<r2>)1/2 |
13.791 |