return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H8 ((Z)-hexa-2,3,4-triene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-231.016772
Energy at 298.15K-231.023106
HF Energy-230.480771
Nuclear repulsion energy189.050483
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3169 3015 25.48      
2 A1 3150 2997 3.71      
3 A1 3073 2923 10.41      
4 A1 2171 2065 0.21      
5 A1 1596 1519 4.48      
6 A1 1487 1415 6.90      
7 A1 1428 1358 0.11      
8 A1 1141 1086 2.64      
9 A1 1087 1034 0.38      
10 A1 761 724 0.32      
11 A1 476 453 5.37      
12 A1 89 84 1.46      
13 A2 3132 2979 0.00      
14 A2 1585 1507 0.00      
15 A2 1117 1062 0.00      
16 A2 883 840 0.00      
17 A2 240 229 0.00      
18 A2 141 134 0.00      
19 B1 3132 2980 34.66      
20 B1 1585 1508 13.10      
21 B1 1115 1061 0.04      
22 B1 730 694 52.94      
23 B1 167 159 1.79      
24 B1 110 105 1.69      
25 B2 3169 3015 12.13      
26 B2 3151 2997 0.03      
27 B2 3073 2923 32.48      
28 B2 1675 1593 2.24      
29 B2 1585 1508 9.21      
30 B2 1481 1409 5.90      
31 B2 1353 1287 10.46      
32 B2 1125 1071 14.85      
33 B2 930 884 13.78      
34 B2 537 510 3.07      
35 B2 161 153 0.14      

Unscaled Zero Point Vibrational Energy (zpe) 25902.4 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 24640.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.40566 0.04592 0.04191

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 0.000 2.179 1.636
H2 0.887 3.464 0.778
H3 -0.887 3.464 0.778
H4 0.000 -2.179 1.636
H5 0.887 -3.464 0.778
H6 -0.887 -3.464 0.778
C7 0.000 2.819 0.749
C8 0.000 -2.819 0.749
H9 0.000 2.510 -1.462
H10 0.000 -2.510 -1.462
C11 0.000 -0.638 -0.521
C12 0.000 0.638 -0.521
C13 0.000 1.968 -0.516
C14 0.000 -1.968 -0.516

Atom - Atom Distances (Å)
  H1 H2 H3 H4 H5 H6 C7 C8 H9 H10 C11 C12 C13 C14
H11.78191.78194.35755.77625.77621.09415.07623.11595.62003.54742.65092.16214.6716
H21.78191.77415.77626.92817.15161.09696.34582.59146.44174.39293.23462.16785.6538
H31.78191.77415.77627.15166.92811.09696.34582.59146.44174.39293.23462.16785.6538
H44.35755.77625.77621.78191.78195.07621.09415.62003.11592.65093.54744.67162.1621
H55.77626.92817.15161.78191.77416.34581.09696.44172.59143.23464.39295.65382.1678
H65.77627.15166.92811.78191.77416.34581.09696.44172.59143.23464.39295.65382.1678
C71.09411.09691.09695.07626.34586.34585.63882.23295.77013.68292.52451.52494.9515
C85.07626.34586.34581.09411.09691.09695.63885.77012.23292.52453.68294.95151.5249
H93.11592.59142.59145.62006.44176.44172.23295.77015.02023.28542.09581.09084.5768
H105.62006.44176.44173.11592.59142.59145.77012.23295.02022.09583.28544.57681.0908
C113.54744.39294.39292.65093.23463.23463.68292.52453.28542.09581.27522.60541.3302
C122.65093.23463.23463.54744.39294.39292.52453.68292.09583.28541.27521.33022.6054
C132.16212.16782.16784.67165.65385.65381.52494.95151.09084.57682.60541.33023.9355
C144.67165.65385.65382.16212.16782.16784.95151.52494.57681.09081.33022.60543.9355

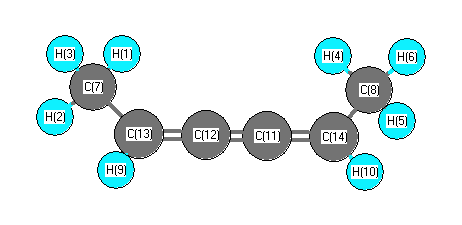
picture of (Z)-hexa-2,3,4-triene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C7 H2 108.834 H1 C7 H3 108.834
H1 C7 C13 110.203 H2 C7 H3 107.933
H2 C7 C13 110.488 H3 C7 C13 110.488
H4 C8 H5 108.834 H4 C8 H6 108.834
H4 C8 C14 110.203 H5 C8 H6 107.933
H5 C8 C14 110.488 H6 C8 C14 110.488
C7 C13 H9 116.233 C7 C13 C12 124.173
C8 C14 H10 116.233 C8 C14 C11 124.173
H9 C13 C12 119.594 H10 C14 C11 119.594
C11 C12 C13 179.780 C12 C11 C14 179.780
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability