return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,2-Butadiene, 3-methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-193.311661
Energy at 298.15K-193.318739
HF Energy-192.861093
Nuclear repulsion energy156.066327
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3166 3012 17.09      
2 A1 3158 3004 3.87      
3 A1 3072 2922 31.56      
4 A1 2043 1943 9.18      
5 A1 1606 1528 0.21      
6 A1 1560 1484 0.61      
7 A1 1490 1418 0.78      
8 A1 1348 1282 11.06      
9 A1 1054 1003 7.95      
10 A1 714 679 0.04      
11 A1 349 332 0.23      
12 A2 3131 2978 0.00      
13 A2 1580 1503 0.00      
14 A2 1070 1018 0.00      
15 A2 648 617 0.00      
16 A2 168 160 0.00      
17 B1 3229 3071 6.62      
18 B1 3133 2981 38.70      
19 B1 1594 1517 16.00      
20 B1 1166 1109 0.78      
21 B1 1106 1052 0.01      
22 B1 430 409 1.57      
23 B1 232 221 5.34      
24 B1 165 157 1.00      
25 B2 3165 3011 15.03      
26 B2 3070 2920 13.71      
27 B2 1591 1514 5.36      
28 B2 1475 1403 4.75      
29 B2 1203 1144 5.81      
30 B2 1001 953 0.28      
31 B2 941 895 59.46      
32 B2 613 583 8.34      
33 B2 174 165 0.59      

Unscaled Zero Point Vibrational Energy (zpe) 25221.4 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 23993.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.27298 0.11817 0.08662

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 2.261
C2 0.000 0.000 0.943
H3 -0.923 0.000 2.836
H4 0.923 0.000 2.836
C5 0.000 0.000 -0.374
C6 0.000 1.295 -1.189
C7 0.000 -1.295 -1.189
H8 0.000 2.167 -0.529
H9 0.000 -2.167 -0.529
H10 0.889 1.334 -1.831
H11 -0.889 1.334 -1.831
H12 -0.889 -1.334 -1.831
H13 0.889 -1.334 -1.831

Atom - Atom Distances (Å)
  C1 C2 H3 H4 C5 C6 C7 H8 H9 H10 H11 H12 H13
C11.31761.08761.08762.63493.68413.68413.53203.53204.39424.39424.39424.3942
C21.31762.10592.10591.31732.49392.49392.61942.61943.20363.20363.20363.2036
H31.08762.10591.84623.34014.32704.32704.10684.10685.18064.85374.85375.1806
H41.08762.10591.84623.34014.32704.32704.10684.10684.85375.18065.18064.8537
C52.63491.31733.34013.34011.52931.52932.17252.17252.16572.16572.16572.1657
C63.68412.49394.32704.32701.52932.58901.09423.52401.09721.09722.84772.8477
C73.68412.49394.32704.32701.52932.58903.52401.09422.84772.84771.09721.0972
H83.53202.61944.10684.10682.17251.09423.52404.33411.78341.78343.83943.8394
H93.53202.61944.10684.10682.17253.52401.09424.33413.83943.83941.78341.7834
H104.39423.20365.18064.85372.16571.09722.84771.78343.83941.77723.20522.6673
H114.39423.20364.85375.18062.16571.09722.84771.78343.83941.77722.66733.2052
H124.39423.20364.85375.18062.16572.84771.09723.83941.78343.20522.66731.7772
H134.39423.20365.18064.85372.16572.84771.09723.83941.78342.66733.20521.7772

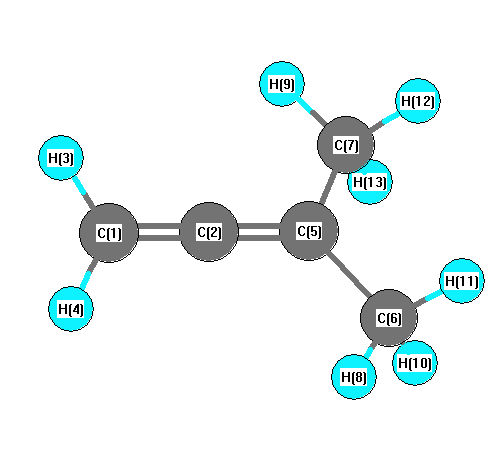
picture of 1,2-Butadiene, 3-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C5 180.000 C2 C1 H3 121.926
C2 C1 H4 121.926 C2 C5 C6 122.172
C2 C5 C7 122.172 H3 C1 H4 116.147
C5 C6 H8 110.718 C5 C6 H10 109.996
C5 C6 H11 109.996 C5 C7 H9 110.718
C5 C7 H12 109.996 C5 C7 H13 109.996
C6 C5 C7 115.656 H8 C6 H10 108.951
H8 C6 H11 108.951 H9 C7 H12 108.951
H9 C7 H13 108.951 H10 C6 H11 108.176
H12 C7 H13 108.176
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability