return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 ((E)-3-methylpent-2-ene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-233.432432
Energy at 298.15K-233.444437
HF Energy-232.881750
Nuclear repulsion energy235.433968
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3181 3026 29.70      
2 A 3168 3014 47.14      
3 A 3165 3011 4.20      
4 A 3156 3003 32.61      
5 A 3140 2987 11.88      
6 A 3131 2979 23.27      
7 A 3127 2974 17.43      
8 A 3112 2961 10.73      
9 A 3085 2935 18.89      
10 A 3078 2928 10.31      
11 A 3075 2925 27.20      
12 A 3067 2918 21.64      
13 A 1719 1635 0.45      
14 A 1607 1529 3.16      
15 A 1604 1526 9.59      
16 A 1599 1521 4.45      
17 A 1594 1516 6.33      
18 A 1592 1514 9.71      
19 A 1588 1511 1.28      
20 A 1575 1498 1.98      
21 A 1500 1427 7.41      
22 A 1486 1413 2.90      
23 A 1476 1404 2.24      
24 A 1446 1376 1.15      
25 A 1400 1332 1.52      
26 A 1346 1281 0.02      
27 A 1253 1192 2.62      
28 A 1182 1125 2.29      
29 A 1146 1090 10.64      
30 A 1124 1069 0.25      
31 A 1115 1061 0.68      
32 A 1100 1047 3.68      
33 A 1024 975 3.29      
34 A 998 950 5.42      
35 A 929 883 0.63      
36 A 869 827 15.45      
37 A 815 776 2.04      
38 A 742 706 2.00      
39 A 566 539 5.40      
40 A 496 472 3.58      
41 A 403 384 0.50      
42 A 319 304 0.19      
43 A 317 302 0.35      
44 A 234 223 0.14      
45 A 195 185 1.04      
46 A 115 109 0.30      
47 A 90 86 0.57      
48 A 53 50 0.05      

Unscaled Zero Point Vibrational Energy (zpe) 37051.8 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 35247.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.19592 0.06750 0.05710

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.523 -0.388 0.257
H2 -2.806 -0.938 1.163
H3 -3.216 -0.682 -0.541
H4 -2.659 0.680 0.452
C5 -0.107 1.635 -0.012
H6 -1.122 1.976 0.208
H7 0.248 2.167 -0.904
H8 0.538 1.920 0.827
C9 2.309 -0.411 0.589
H10 1.894 -1.039 1.383
H11 2.450 0.601 0.983
C12 1.339 -0.401 -0.627
H13 1.762 0.219 -1.430
H14 1.239 -1.424 -1.015
C15 -1.090 -0.716 -0.135
H16 -0.911 -1.772 -0.344
C17 -0.046 0.130 -0.252
H18 3.289 -0.808 0.300

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 H11 C12 H13 H14 C15 H16 C17 H18
C11.09741.09721.09323.16212.74783.94373.87574.84314.60415.12123.96154.64454.10401.52092.20812.58105.8264
H21.09741.77141.77313.91003.49854.82144.41235.17444.70635.47954.54715.37874.62012.16332.56133.28076.1570
H31.09721.77141.77433.91293.46514.49954.76825.64565.47156.00524.56395.13594.54102.16432.55723.28476.5595
H41.09321.77311.77432.76422.02543.53623.45035.08824.95495.13694.27974.82684.66592.17993.11452.76176.1326
C53.16213.91003.91292.76421.09311.09771.09683.22293.62032.93242.57252.74053.48952.55113.51691.52614.1951
H62.74783.49853.46512.02541.09311.77571.77324.19724.42403.90533.52243.75364.31642.71363.79472.18645.2168
H73.94374.82144.49953.53621.09771.77571.77313.62224.26863.29552.80372.52223.72643.26984.14422.15924.4206
H83.87574.41234.76823.45031.09681.77321.77312.93703.30252.32782.85443.08013.88173.24434.13632.17123.9097
C94.84315.17445.64565.08823.22294.19723.62222.93701.09491.09451.55602.18422.17783.48873.61862.55841.0958
H104.60414.70635.47154.95493.62034.42404.26863.30251.09491.77752.18143.08442.51533.36353.37472.79371.7813
H115.12125.47956.00525.13692.93243.90533.29552.32781.09451.77752.19802.53763.09103.93854.32282.82411.7761
C123.96154.54714.56394.27972.57253.52242.80372.85441.55602.18142.19801.09871.09792.49832.64991.52972.1969
H134.64455.37875.13594.82682.74053.75362.52223.08012.18423.08442.53761.09871.77303.26873.50542.15932.5250
H144.10404.62014.54104.66593.48954.31643.72643.88172.17782.51533.09101.09791.77302.58872.27922.15532.5114
C151.52092.16332.16432.17992.55112.71363.26983.24433.48873.36353.93852.49833.26872.58871.09201.34854.4011
H162.20812.56132.55723.11453.51693.79474.14424.13633.61863.37474.32282.64993.50542.27921.09202.09144.3567
C172.58103.28073.28472.76171.52612.18642.15922.17122.55842.79372.82411.52972.15932.15531.34852.09143.5074
H185.82646.15706.55956.13264.19515.21684.42063.90971.09581.78131.77612.19692.52502.51144.40114.35673.5074

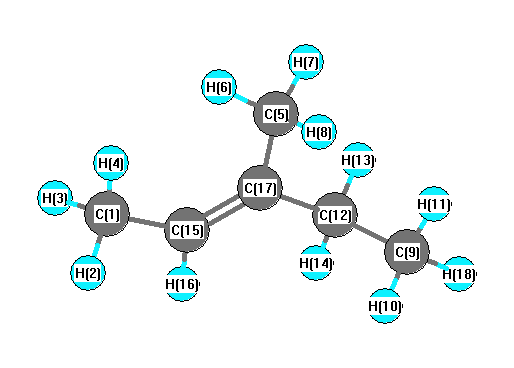
picture of (E)-3-methylpent-2-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C15 H16 114.360 C1 C15 C17 128.072
H2 C1 H3 107.636 H2 C1 H4 108.077
H2 C1 C15 110.376 H3 C1 H4 108.190
H3 C1 C15 110.462 H4 C1 C15 111.957
C5 C17 C12 114.673 C5 C17 C15 124.996
H6 C5 H7 108.292 H6 C5 H8 108.142
H6 C5 C17 112.122 H7 C5 H8 107.802
H7 C5 C17 109.673 H8 C5 C17 110.679
C9 C12 H13 109.518 C9 C12 H14 109.068
C9 C12 C17 112.016 H10 C9 H11 108.561
H10 C9 C12 109.523 H10 C9 H18 108.804
H11 C9 C12 110.845 H11 C9 H18 108.369
C12 C9 H18 110.684 C12 C17 C15 120.322
H13 C12 H14 107.639 H13 C12 C17 109.379
H14 C12 C17 109.118 H16 C15 C17 117.568
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability