return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHCHCH3 (2-Butene, (E)-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2h 1Ag
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-155.608542
Energy at 298.15K-155.616291
HF Energy-155.244517
Nuclear repulsion energy114.952827
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3158 3004 0.00      
2 A 3143 2990 0.00      
3 A 3072 2922 0.00      
4 A 1721 1637 0.00      
5 A 1594 1517 0.00      
6 A 1494 1421 0.00      
7 A 1390 1322 0.00      
8 A 1198 1140 0.00      
9 A 886 842 0.00      
10 A 510 486 0.00      
11 A 3133 2980 43.24      
12 A 1586 1509 12.56      
13 A 1137 1082 0.10      
14 A 1026 976 37.76      
15 A 246 234 4.06      
16 A 166 158 0.85      
17 A 3133 2980 0.00      
18 A 1587 1509 0.00      
19 A 1127 1072 0.00      
20 A 763 725 0.00      
21 A 210 200 0.00      
22 A 3166 3012 61.56      
23 A 3142 2989 13.09      
24 A 3072 2922 39.60      
25 A 1603 1525 13.63      
26 A 1494 1421 7.46      
27 A 1398 1330 2.13      
28 A 1073 1021 4.95      
29 A 1029 979 8.05      
30 A 291 277 0.69      

Unscaled Zero Point Vibrational Energy (zpe) 24273.8 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 23091.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
1.14138 0.12221 0.11517

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.671 0.000
C2 0.000 -0.671 0.000
C3 -1.252 1.533 0.000
C4 1.252 -1.533 0.000
H5 0.952 1.205 0.000
H6 -0.952 -1.205 0.000
H7 -2.148 0.902 0.000
H8 2.148 -0.902 0.000
H9 -1.284 2.179 0.886
H10 -1.284 2.179 -0.886
H11 1.284 -2.179 0.886
H12 1.284 -2.179 -0.886

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.34281.52032.53561.09102.10392.16082.66312.16952.16953.24943.2494
C21.34282.53561.52032.10391.09102.66312.16083.24943.24942.16952.1695
C31.52032.53563.95962.22822.75491.09614.18301.09681.09684.58264.5826
C42.53561.52033.95962.75492.22824.18301.09614.58264.58261.09681.0968
H51.09102.10392.22822.75493.07093.11472.42352.59462.59463.51383.5138
H62.10391.09102.75492.22823.07092.42353.11473.51383.51382.59462.5946
H72.16082.66311.09614.18303.11472.42354.66041.77821.77824.69704.6970
H82.66312.16084.18301.09612.42353.11474.66044.69704.69701.77821.7782
H92.16953.24941.09684.58262.59463.51381.77824.69701.77245.05835.3598
H102.16953.24941.09684.58262.59463.51381.77824.69701.77245.35985.0583
H113.24942.16954.58261.09683.51382.59464.69701.77825.05835.35981.7724
H123.24942.16954.58261.09683.51382.59464.69701.77825.35985.05831.7724

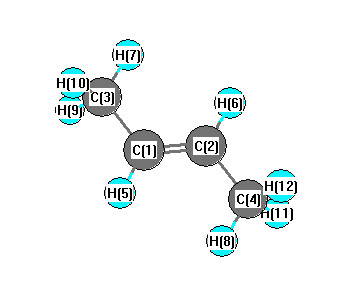
picture of 2-Butene, (E)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 124.541 C1 C2 H6 119.286
C1 C3 H7 110.301 C1 C3 H9 110.953
C1 C3 H10 110.953 C2 C1 C3 124.541
C2 C1 H5 119.286 C2 C4 H8 110.301
C2 C4 H11 110.953 C2 C4 H12 110.953
C3 C1 H5 116.174 C4 C2 H6 116.174
H7 C3 H9 108.368 H7 C3 H10 108.368
H8 C4 H11 108.368 H8 C4 H12 108.368
H9 C3 H10 107.799 H11 C4 H12 107.799
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability