Jump to
S1C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -275.831325 |
| Energy at 298.15K | |
| HF Energy | -275.418070 |
| Nuclear repulsion energy | 123.557959 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3125 |
2972 |
0.00 |
|
|
|
| 2 |
Ag |
1628 |
1548 |
0.00 |
|
|
|
| 3 |
Ag |
1509 |
1436 |
0.00 |
|
|
|
| 4 |
Ag |
1079 |
1026 |
0.00 |
|
|
|
| 5 |
Ag |
1063 |
1011 |
0.00 |
|
|
|
| 6 |
Ag |
440 |
418 |
0.00 |
|
|
|
| 7 |
Au |
3192 |
3037 |
63.44 |
|
|
|
| 8 |
Au |
1263 |
1201 |
1.54 |
|
|
|
| 9 |
Au |
872 |
829 |
0.02 |
|
|
|
| 10 |
Au |
125 |
119 |
14.93 |
|
|
|
| 11 |
Bg |
3167 |
3013 |
0.00 |
|
|
|
| 12 |
Bg |
1375 |
1308 |
0.00 |
|
|
|
| 13 |
Bg |
1210 |
1151 |
0.00 |
|
|
|
| 14 |
Bu |
3133 |
2980 |
44.84 |
|
|
|
| 15 |
Bu |
1642 |
1562 |
1.16 |
|
|
|
| 16 |
Bu |
1424 |
1355 |
13.33 |
|
|
|
| 17 |
Bu |
1097 |
1043 |
82.08 |
|
|
|
| 18 |
Bu |
236 |
224 |
20.92 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13788.6 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 13117.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
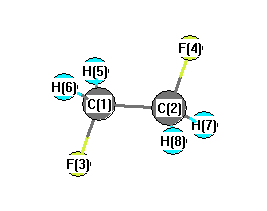
Geometric Data calculated at MP2/3-21G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.438 |
0.626 |
0.000 |
| C2 |
-0.438 |
-0.626 |
0.000 |
| F3 |
-0.438 |
1.756 |
0.000 |
| F4 |
0.438 |
-1.756 |
0.000 |
| H5 |
1.070 |
0.642 |
0.890 |
| H6 |
1.070 |
0.642 |
-0.890 |
| H7 |
-1.070 |
-0.642 |
0.890 |
| H8 |
-1.070 |
-0.642 |
-0.890 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5285 | 1.4294 | 2.3827 | 1.0915 | 1.0915 | 2.1616 | 2.1616 |
C2 | 1.5285 | | 2.3827 | 1.4294 | 2.1616 | 2.1616 | 1.0915 | 1.0915 | F3 | 1.4294 | 2.3827 | | 3.6200 | 2.0753 | 2.0753 | 2.6346 | 2.6346 | F4 | 2.3827 | 1.4294 | 3.6200 | | 2.6346 | 2.6346 | 2.0753 | 2.0753 | H5 | 1.0915 | 2.1616 | 2.0753 | 2.6346 | | 1.7797 | 2.4948 | 3.0646 | H6 | 1.0915 | 2.1616 | 2.0753 | 2.6346 | 1.7797 | | 3.0646 | 2.4948 | H7 | 2.1616 | 1.0915 | 2.6346 | 2.0753 | 2.4948 | 3.0646 | | 1.7797 | H8 | 2.1616 | 1.0915 | 2.6346 | 2.0753 | 3.0646 | 2.4948 | 1.7797 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.274 |
|
C1 |
C2 |
H7 |
110.066 |
| C1 |
C2 |
H8 |
110.066 |
|
C2 |
C1 |
F3 |
107.274 |
| C2 |
C1 |
H5 |
110.066 |
|
C2 |
C1 |
H6 |
110.066 |
| F3 |
C1 |
H5 |
110.099 |
|
F3 |
C1 |
H6 |
110.099 |
| F4 |
C2 |
H7 |
110.099 |
|
F4 |
C2 |
H8 |
110.099 |
| H5 |
C1 |
H6 |
109.221 |
|
H7 |
C2 |
H8 |
109.221 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -275.831079 |
| Energy at 298.15K | |
| HF Energy | -275.416738 |
| Nuclear repulsion energy | 125.479303 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3161 |
3007 |
21.33 |
|
|
|
| 2 |
A |
3116 |
2965 |
24.97 |
|
|
|
| 3 |
A |
1609 |
1530 |
0.14 |
|
|
|
| 4 |
A |
1488 |
1416 |
11.41 |
|
|
|
| 5 |
A |
1358 |
1292 |
0.00 |
|
|
|
| 6 |
A |
1167 |
1110 |
0.76 |
|
|
|
| 7 |
A |
1118 |
1063 |
35.01 |
|
|
|
| 8 |
A |
882 |
839 |
14.38 |
|
|
|
| 9 |
A |
311 |
296 |
0.66 |
|
|
|
| 10 |
A |
143 |
137 |
4.23 |
|
|
|
| 11 |
B |
3172 |
3018 |
41.70 |
|
|
|
| 12 |
B |
3106 |
2955 |
14.87 |
|
|
|
| 13 |
B |
1610 |
1532 |
3.83 |
|
|
|
| 14 |
B |
1505 |
1432 |
9.77 |
|
|
|
| 15 |
B |
1298 |
1234 |
1.22 |
|
|
|
| 16 |
B |
1107 |
1053 |
12.66 |
|
|
|
| 17 |
B |
948 |
902 |
25.38 |
|
|
|
| 18 |
B |
462 |
440 |
17.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13780.7 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 13109.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.284 |
0.705 |
0.523 |
| C2 |
-0.284 |
-0.705 |
0.523 |
| F3 |
-0.284 |
1.449 |
-0.557 |
| F4 |
0.284 |
-1.449 |
-0.557 |
| H5 |
0.038 |
1.197 |
1.468 |
| H6 |
1.369 |
0.660 |
0.405 |
| H7 |
-0.038 |
-1.197 |
1.468 |
| H8 |
-1.369 |
-0.660 |
0.405 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5197 | 1.4290 | 2.4099 | 1.0937 | 1.0924 | 2.1484 | 2.1460 |
C2 | 1.5197 | | 2.4099 | 1.4290 | 2.1484 | 2.1460 | 1.0937 | 1.0924 | F3 | 1.4290 | 2.4099 | | 2.9538 | 2.0658 | 2.0682 | 3.3417 | 2.5591 | F4 | 2.4099 | 1.4290 | 2.9538 | | 3.3417 | 2.5591 | 2.0658 | 2.0682 | H5 | 1.0937 | 2.1484 | 2.0658 | 3.3417 | | 1.7867 | 2.3961 | 2.5607 | H6 | 1.0924 | 2.1460 | 2.0682 | 2.5591 | 1.7867 | | 2.5607 | 3.0383 | H7 | 2.1484 | 1.0937 | 3.3417 | 2.0658 | 2.3961 | 2.5607 | | 1.7867 | H8 | 2.1460 | 1.0924 | 2.5591 | 2.0682 | 2.5607 | 3.0383 | 1.7867 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
109.584 |
|
C1 |
C2 |
H7 |
109.501 |
| C1 |
C2 |
H8 |
109.395 |
|
C2 |
C1 |
F3 |
109.584 |
| C2 |
C1 |
H5 |
109.501 |
|
C2 |
C1 |
H6 |
109.395 |
| F3 |
C1 |
H5 |
109.220 |
|
F3 |
C1 |
H6 |
109.500 |
| F4 |
C2 |
H7 |
109.220 |
|
F4 |
C2 |
H8 |
109.500 |
| H5 |
C1 |
H6 |
109.627 |
|
H7 |
C2 |
H8 |
109.627 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
