return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (2-Pentene, (Z)-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-194.517048
Energy at 298.15K-194.527118
HF Energy-194.060910
Nuclear repulsion energy170.485131
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3182 3027 54.58      
2 A 3166 3012 1.15      
3 A 3165 3011 40.32      
4 A 3156 3003 25.75      
5 A 3147 2993 4.67      
6 A 3131 2979 16.69      
7 A 3131 2979 4.99      
8 A 3085 2935 16.87      
9 A 3082 2932 23.13      
10 A 3078 2928 17.74      
11 A 1703 1620 0.36      
12 A 1605 1527 6.19      
13 A 1602 1524 6.10      
14 A 1599 1521 4.28      
15 A 1592 1514 5.40      
16 A 1580 1503 1.41      
17 A 1495 1422 4.33      
18 A 1491 1418 5.68      
19 A 1480 1408 2.26      
20 A 1397 1329 1.48      
21 A 1360 1293 0.42      
22 A 1343 1277 0.31      
23 A 1210 1151 1.43      
24 A 1140 1084 5.82      
25 A 1129 1074 0.28      
26 A 1099 1045 0.58      
27 A 1022 972 0.28      
28 A 1013 964 2.35      
29 A 938 892 7.47      
30 A 866 823 1.40      
31 A 817 777 4.98      
32 A 743 707 37.69      
33 A 581 552 6.17      
34 A 489 465 1.75      
35 A 321 305 0.05      
36 A 259 247 0.05      
37 A 231 220 0.01      
38 A 110 105 0.26      
39 A 41 39 0.08      

Unscaled Zero Point Vibrational Energy (zpe) 30788.7 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 29289.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.33754 0.08824 0.08015

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.115 -0.383 -0.441
H2 2.590 0.543 -0.784
H3 2.890 -1.044 -0.037
H4 1.647 -0.871 -1.302
C5 1.036 -0.069 0.637
H6 0.574 -1.003 0.972
H7 1.520 0.388 1.510
C8 -2.013 -0.725 -0.028
H9 -2.385 -1.091 -0.993
H10 -1.352 -1.487 0.395
H11 -2.876 -0.610 0.640
C12 -0.014 0.885 0.091
H13 0.351 1.894 -0.107
C14 -1.298 0.605 -0.205
H15 -1.916 1.403 -0.617

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 H11 C12 H13 C14 H15
C11.09551.09561.09461.55682.18102.18044.16314.58883.73375.11202.53452.89923.56064.4122
H21.09551.77931.77602.19233.08842.53574.83425.24044.58815.76492.76832.70093.93064.5898
H31.09561.77931.78162.19992.52662.51384.91345.36104.28705.82183.48863.88314.50365.4238
H41.09461.77601.78162.18532.51833.08353.87884.05023.50114.92962.78973.27873.47184.2819
C51.55682.19232.19992.18531.09501.09803.18963.92522.78853.94991.52052.20782.57133.5290
H62.18103.08842.52662.51831.09501.76682.78743.55292.06823.48822.16493.09922.73423.8097
H72.18042.53572.51383.08351.09801.76684.01154.86883.60744.59192.14852.49933.30624.1665
C84.16314.83424.91343.87883.18962.78744.01151.09701.09341.09692.56963.52951.52162.2112
H94.58885.24045.36104.05023.92523.55294.86881.09701.77461.77133.27094.14502.16352.5660
H103.73374.58814.28703.50112.78852.06823.60741.09341.77461.77522.74023.81882.17763.1139
H115.11205.76495.82184.92963.94993.48824.59191.09691.77131.77523.27524.15242.16432.5606
C122.53452.76833.48862.78971.52052.16492.14852.56963.27092.74023.27521.09101.34632.0939
H132.89922.70093.88313.27872.20783.09922.49933.52954.14503.81884.15241.09102.09472.3746
C143.56063.93064.50363.47182.57132.73423.30621.52162.16352.17762.16431.34632.09471.0900
H154.41224.58985.42384.28193.52903.80974.16652.21122.56603.11392.56062.09392.37461.0900

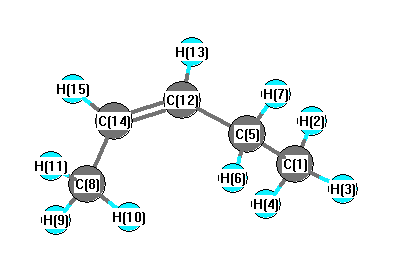
picture of 2-Pentene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 109.426 C1 C5 H7 109.208
C1 C5 C12 110.893 H2 C1 H3 108.594
H2 C1 H4 108.372 H2 C1 C5 110.286
H3 C1 H4 108.871 H3 C1 C5 110.878
H4 C1 C5 109.783 C5 C12 H13 114.425
C5 C12 C14 127.410 H6 C5 H7 107.348
H6 C5 C12 110.679 H7 C5 C12 109.207
C8 C14 C12 127.170 C8 C14 H15 114.697
H9 C8 H10 108.222 H9 C8 H11 107.679
H9 C8 C14 110.369 H10 C8 H11 108.289
H10 C8 C14 111.721 H11 C8 C14 110.433
C12 C14 H15 118.133 H13 C12 C14 118.126
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability