Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP=FULL/3-21G*
Geometric Data calculated at B2PLYP=FULL/3-21G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -551.073725 |
| Energy at 298.15K | -551.079280 |
| HF Energy | -550.904073 |
| Nuclear repulsion energy | 201.832403 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3273 |
3273 |
7.94 |
|
|
|
| 2 |
A |
3270 |
3270 |
5.38 |
|
|
|
| 3 |
A |
3206 |
3206 |
4.56 |
|
|
|
| 4 |
A |
3196 |
3196 |
5.01 |
|
|
|
| 5 |
A |
3186 |
3186 |
0.77 |
|
|
|
| 6 |
A |
3180 |
3180 |
1.34 |
|
|
|
| 7 |
A |
1666 |
1666 |
22.87 |
|
|
|
| 8 |
A |
1655 |
1655 |
20.17 |
|
|
|
| 9 |
A |
1490 |
1490 |
3.65 |
|
|
|
| 10 |
A |
1488 |
1488 |
7.34 |
|
|
|
| 11 |
A |
1362 |
1362 |
1.84 |
|
|
|
| 12 |
A |
1354 |
1354 |
7.71 |
|
|
|
| 13 |
A |
1118 |
1118 |
10.06 |
|
|
|
| 14 |
A |
1095 |
1095 |
10.31 |
|
|
|
| 15 |
A |
1035 |
1035 |
24.28 |
|
|
|
| 16 |
A |
1020 |
1020 |
38.07 |
|
|
|
| 17 |
A |
982 |
982 |
43.85 |
|
|
|
| 18 |
A |
949 |
949 |
53.18 |
|
|
|
| 19 |
A |
739 |
739 |
9.05 |
|
|
|
| 20 |
A |
690 |
690 |
1.96 |
|
|
|
| 21 |
A |
651 |
651 |
19.74 |
|
|
|
| 22 |
A |
627 |
627 |
14.54 |
|
|
|
| 23 |
A |
474 |
474 |
0.62 |
|
|
|
| 24 |
A |
387 |
387 |
0.62 |
|
|
|
| 25 |
A |
230 |
230 |
1.13 |
|
|
|
| 26 |
A |
149 |
149 |
4.17 |
|
|
|
| 27 |
A |
82 |
82 |
1.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19275.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 19275.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/3-21G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.008 |
-0.992 |
-0.003 |
| C2 |
1.082 |
0.359 |
0.388 |
| C3 |
-1.567 |
-0.188 |
-0.024 |
| C4 |
2.252 |
0.560 |
-0.224 |
| C5 |
-1.797 |
1.125 |
-0.129 |
| H6 |
0.757 |
0.981 |
1.217 |
| H7 |
-2.386 |
-0.895 |
0.066 |
| H8 |
2.928 |
1.337 |
0.110 |
| H9 |
2.571 |
-0.047 |
-1.063 |
| H10 |
-0.998 |
1.844 |
-0.250 |
| H11 |
-2.809 |
1.510 |
-0.106 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7695 | 1.7684 | 2.7368 | 2.7845 | 2.4376 | 2.3964 | 3.7371 | 2.9297 | 3.0190 | 3.7688 |
C2 | 1.7695 | | 2.7360 | 1.3359 | 3.0235 | 1.0855 | 3.7009 | 2.1082 | 2.1187 | 2.6334 | 4.0874 | C3 | 1.7684 | 2.7360 | | 3.8966 | 1.3368 | 2.8825 | 1.0848 | 4.7491 | 4.2688 | 2.1228 | 2.1051 | C4 | 2.7368 | 1.3359 | 3.8966 | | 4.0894 | 2.1185 | 4.8685 | 1.0835 | 1.0833 | 3.4940 | 5.1507 | C5 | 2.7845 | 3.0235 | 1.3368 | 4.0894 | | 2.8908 | 2.1121 | 4.7364 | 4.6181 | 1.0828 | 1.0830 | H6 | 2.4376 | 1.0855 | 2.8825 | 2.1185 | 2.8908 | | 3.8364 | 2.4631 | 3.0893 | 2.4444 | 3.8403 | H7 | 2.3964 | 3.7009 | 1.0848 | 4.8685 | 2.1121 | 3.8364 | | 5.7636 | 5.1534 | 3.0867 | 2.4473 | H8 | 3.7371 | 2.1082 | 4.7491 | 1.0835 | 4.7364 | 2.4631 | 5.7636 | | 1.8492 | 3.9747 | 5.7442 | H9 | 2.9297 | 2.1187 | 4.2688 | 1.0833 | 4.6181 | 3.0893 | 5.1534 | 1.8492 | | 4.1197 | 5.6821 | H10 | 3.0190 | 2.6334 | 2.1228 | 3.4940 | 1.0828 | 2.4444 | 3.0867 | 3.9747 | 4.1197 | | 1.8481 | H11 | 3.7688 | 4.0874 | 2.1051 | 5.1507 | 1.0830 | 3.8403 | 2.4473 | 5.7442 | 5.6821 | 1.8481 | |
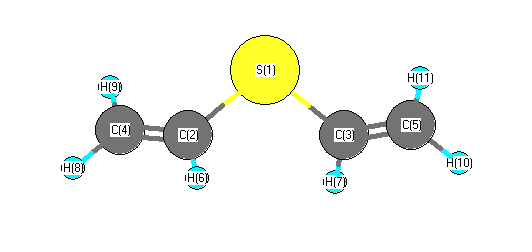 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
122.991 |
|
S1 |
C2 |
H6 |
115.139 |
| S1 |
C3 |
C5 |
126.904 |
|
S1 |
C3 |
H7 |
112.019 |
| C2 |
S1 |
C3 |
101.306 |
|
C2 |
C4 |
H8 |
120.885 |
| C2 |
C4 |
H9 |
121.936 |
|
C3 |
C5 |
H10 |
122.296 |
| C3 |
C5 |
H11 |
120.550 |
|
C4 |
C2 |
H6 |
121.730 |
| C5 |
C3 |
H7 |
121.076 |
|
H8 |
C4 |
H9 |
117.177 |
| H10 |
C5 |
H11 |
117.150 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
