return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H8 (1,4-Cyclohexadiene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1Ag
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-231.071680
Energy at 298.15K-231.080250
HF Energy-230.540244
Nuclear repulsion energy216.035148
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3195 3039 0.00      
2 Ag 3050 2901 0.00      
3 Ag 1720 1637 0.00      
4 Ag 1574 1497 0.00      
5 Ag 1294 1231 0.00      
6 Ag 836 795 0.00      
7 Ag 557 530 0.00      
8 Au 1286 1223 0.00      
9 Au 1012 962 0.00      
10 Au 377 358 0.00      
11 B1g 3166 3011 0.00      
12 B1g 1452 1381 0.00      
13 B1g 1395 1327 0.00      
14 B1g 1067 1015 0.00      
15 B1g 593 564 0.00      
16 B1u 3072 2923 38.22      
17 B1u 1045 994 17.84      
18 B1u 655 623 79.62      
19 B1u 65 62 0.48      
20 B2g 3073 2923 0.00      
21 B2g 1044 993 0.00      
22 B2g 996 947 0.00      
23 B2g 381 362 0.00      
24 B2u 3191 3036 67.67      
25 B2u 1684 1602 1.22      
26 B2u 1404 1335 1.72      
27 B2u 1244 1184 0.58      
28 B2u 929 884 0.43      
29 B3g 1290 1227 0.00      
30 B3g 742 706 0.00      
31 B3u 3167 3012 13.13      
32 B3u 3051 2903 47.71      
33 B3u 1583 1506 3.45      
34 B3u 1490 1417 0.06      
35 B3u 1004 955 0.53      
36 B3u 875 832 6.95      

Unscaled Zero Point Vibrational Energy (zpe) 27277.0 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 25948.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.16906 0.16010 0.08482

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.520 0.000 0.000
C2 -1.520 0.000 0.000
C3 -0.672 1.266 0.000
C4 0.672 1.266 0.000
C5 -0.672 -1.266 0.000
C6 0.672 -1.266 0.000
H7 1.212 -2.211 0.000
H8 -1.212 -2.211 0.000
H9 1.212 2.211 0.000
H10 -1.212 2.211 0.000
H11 2.180 0.000 0.881
H12 -2.180 0.000 0.881
H13 2.180 0.000 -0.881
H14 -2.180 0.000 -0.881

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C13.03932.53061.52342.53061.52342.23213.51432.23213.51431.10103.80291.10103.8029
C23.03931.52342.53061.52342.53063.51432.23213.51432.23213.80291.10103.80291.1010
C32.53061.52341.34352.53122.86573.95403.51812.10771.08883.24182.15693.24182.1569
C41.52342.53061.34352.86572.53123.51813.95401.08882.10772.15693.24182.15693.2418
C52.53061.52342.53122.86571.34352.10771.08883.95403.51813.24182.15693.24182.1569
C61.52342.53062.86572.53121.34351.08882.10773.51813.95402.15693.24182.15693.2418
H72.23213.51433.95403.51812.10771.08882.42434.42165.04262.56914.14362.56914.1436
H83.51432.23213.51813.95401.08882.10772.42435.04264.42164.14362.56914.14362.5691
H92.23213.51432.10771.08883.95403.51814.42165.04262.42432.56914.14362.56914.1436
H103.51432.23211.08882.10773.51813.95405.04264.42162.42434.14362.56914.14362.5691
H111.10103.80293.24182.15693.24182.15692.56914.14362.56914.14364.35951.76234.7022
H123.80291.10102.15693.24182.15693.24184.14362.56914.14362.56914.35954.70221.7623
H131.10103.80293.24182.15693.24182.15692.56914.14362.56914.14361.76234.70224.3595
H143.80291.10102.15693.24182.15693.24184.14362.56914.14362.56914.70221.76234.3595

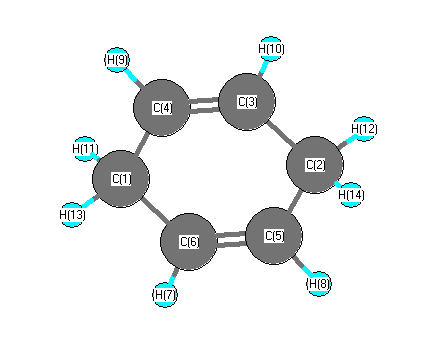
picture of 1,4-Cyclohexadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 C3 123.820 C1 C4 H9 116.423
C1 C6 C5 123.820 C1 C6 H7 116.423
C2 C3 C4 123.820 C2 C3 H10 116.423
C2 C5 C6 123.820 C2 C5 H8 116.423
C3 C2 C5 112.361 C3 C2 H12 109.493
C3 C2 H14 109.493 C3 C4 H9 119.758
C4 C1 C6 112.361 C4 C1 H11 109.493
C4 C1 H13 109.493 C4 C3 H10 119.758
C5 C2 H12 109.493 C5 C2 H14 109.493
C5 C6 H7 119.758 C6 C1 H11 109.493
C6 C1 H13 109.493 C6 C5 H8 119.758
H11 C1 H13 106.325 H12 C2 H14 106.325
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability