return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (2-Pentene, (E)-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-194.518066
Energy at 298.15K-194.528241
HF Energy-194.061929
Nuclear repulsion energy169.571286
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3187 3031 21.64      
2 A' 3157 3003 28.07      
3 A' 3145 2992 23.14      
4 A' 3141 2988 12.11      
5 A' 3086 2936 14.55      
6 A' 3072 2922 22.93      
7 A' 3068 2919 18.96      
8 A' 1718 1634 0.16      
9 A' 1609 1530 5.11      
10 A' 1599 1521 8.56      
11 A' 1584 1507 1.06      
12 A' 1493 1420 3.12      
13 A' 1490 1418 5.60      
14 A' 1434 1364 4.90      
15 A' 1393 1326 0.72      
16 A' 1372 1305 1.60      
17 A' 1172 1115 2.82      
18 A' 1096 1043 3.02      
19 A' 1063 1011 0.35      
20 A' 965 918 5.51      
21 A' 855 813 0.02      
22 A' 590 562 0.95      
23 A' 378 359 0.16      
24 A' 217 207 0.24      
25 A" 3164 3010 28.77      
26 A" 3133 2980 21.30      
27 A" 3098 2947 14.84      
28 A" 1602 1524 5.63      
29 A" 1586 1509 6.12      
30 A" 1362 1296 0.00      
31 A" 1174 1117 1.00      
32 A" 1131 1076 0.09      
33 A" 1031 981 33.09      
34 A" 869 827 4.47      
35 A" 732 697 0.35      
36 A" 309 294 1.76      
37 A" 220 210 1.48      
38 A" 192 183 1.36      
39 A" 112 107 0.03      

Unscaled Zero Point Vibrational Energy (zpe) 30800.8 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 29300.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.43555 0.08130 0.07125

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -0.723 -1.237 0.000
C2 -0.975 -0.178 0.000
H3 -0.280 1.802 0.000
C4 0.000 0.746 0.000
H5 -2.953 -0.267 0.886
H6 -2.953 -0.267 -0.886
H7 -2.610 1.236 0.000
C8 -2.460 0.150 0.000
H9 1.945 0.959 -0.881
H10 1.945 0.959 0.881
C11 1.500 0.475 0.000
H12 2.972 -1.138 0.000
H13 1.484 -1.526 0.889
H14 1.484 -1.526 -0.889
C15 1.883 -1.026 0.000

Atom - Atom Distances (Å)
  H1 C2 H3 C4 H5 H6 H7 C8 H9 H10 C11 H12 H13 H14 C15
H11.08873.07122.11092.58882.58883.11122.22323.56633.56632.80613.69672.39662.39662.6141
C21.08872.09831.34302.16992.16992.16211.52113.25503.25502.55944.06212.94132.94132.9806
H33.07122.09831.09233.49483.49482.39802.73502.53742.53742.21984.38383.86953.86953.5602
C42.11091.34301.09233.24563.24562.65582.53102.14612.14611.52423.51902.85532.85532.5856
H52.58882.16993.49483.24561.77231.77831.09685.35005.04994.60106.05464.61244.94224.9749
H62.58882.16993.49483.24561.77231.77831.09685.04995.35004.60106.05464.94224.61244.9749
H73.11122.16212.39802.65581.77831.77831.09614.64834.64834.18006.06635.01795.01795.0304
C82.22321.52112.73502.53101.09681.09681.09614.56474.56473.97325.58294.37644.37644.4992
H93.56633.25502.53742.14615.35005.04994.64834.56471.76151.09902.49513.08502.52692.1726
H103.56633.25502.53742.14615.04995.35004.64834.56471.76151.09902.49512.52693.08502.1726
C112.80612.55942.21981.52424.60104.60104.18003.97321.09901.09902.18392.18952.18951.5496
H123.69674.06214.38383.51906.05466.05466.06635.58292.49512.49512.18391.77661.77661.0954
H132.39662.94133.86952.85534.61244.94225.01794.37643.08502.52692.18951.77661.77801.0948
H142.39662.94133.86952.85534.94224.61245.01794.37642.52693.08502.18951.77661.77801.0948
C152.61412.98063.56022.58564.97494.97495.03044.49922.17262.17261.54961.09541.09481.0948

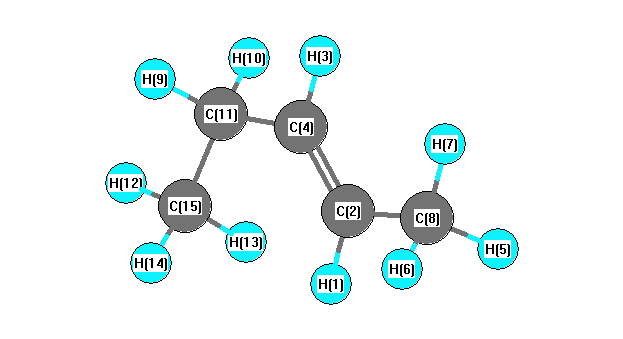
picture of 2-Pentene, (E)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C4 120.102 H1 C2 C8 115.837
C2 C4 H3 118.630 C2 C4 C11 126.298
C2 C8 H5 110.928 C2 C8 H6 110.928
C2 C8 H7 110.347 H3 C4 C11 115.072
C4 C2 C8 124.061 C4 C11 H9 108.706
C4 C11 H10 108.706 C4 C11 C15 114.529
H5 C8 H6 107.791 H5 C8 H7 108.375
H6 C8 H7 108.375 H9 C11 H10 106.521
H9 C11 C15 109.039 H10 C11 C15 109.039
C11 C15 H12 110.136 C11 C15 H13 110.609
C11 C15 H14 110.609 H12 C15 H13 108.418
H12 C15 H14 108.418 H13 C15 H14 108.584
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability