return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7NO (dimethylformamide)

using model chemistry: MP2=FULL/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/3-21G*
 hartrees
Energy at 0K-246.125267
Energy at 298.15K-246.133731
HF Energy-245.609856
Nuclear repulsion energy179.002426
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3185 3020 0.96      
2 A' 3151 2988 18.00      
3 A' 3069 2909 37.46      
4 A' 3059 2900 39.98      
5 A' 2994 2838 101.19      
6 A' 1693 1605 223.19      
7 A' 1627 1542 6.36      
8 A' 1609 1525 15.50      
9 A' 1528 1449 12.38      
10 A' 1499 1421 16.29      
11 A' 1471 1394 39.98      
12 A' 1421 1347 52.73      
13 A' 1275 1209 47.15      
14 A' 1110 1052 123.97      
15 A' 1106 1049 28.40      
16 A' 864 819 1.24      
17 A' 652 619 5.57      
18 A' 381 361 1.19      
19 A' 323 306 11.92      
20 A" 3127 2964 29.32      
21 A" 3114 2952 41.55      
22 A" 1604 1521 12.48      
23 A" 1580 1498 5.16      
24 A" 1218 1154 3.63      
25 A" 1173 1112 0.11      
26 A" 1060 1005 0.06      
27 A" 341 323 20.88      
28 A" 224 212 3.04      
29 A" 206 195 1.55      
30 A" 136 129 0.34      

Unscaled Zero Point Vibrational Energy (zpe) 22899.5 cm-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 21708.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/3-21G*
ABC
0.29064 0.13540 0.09574

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.722 -0.816 0.000
O2 0.229 -1.967 0.000
N3 0.000 0.360 0.000
C4 -1.476 0.299 0.000
C5 0.652 1.679 0.000
H6 1.807 -0.623 0.000
H7 -1.740 -0.760 0.000
H8 -1.883 0.787 0.893
H9 -1.883 0.787 -0.893
H10 1.738 1.539 0.000
H11 0.370 2.253 -0.891
H12 0.370 2.253 0.891

Atom - Atom Distances (Å)
  C1 O2 N3 C4 C5 H6 H7 H8 H9 H10 H11 H12
C11.25211.38042.46492.49641.10232.46303.18693.18692.56533.21563.2156
O21.25212.33892.83643.67092.07272.31003.58443.58443.81734.31594.3159
N31.38042.33891.47731.47122.05762.07012.12762.12762.10012.12472.1247
C42.46492.83641.47732.53633.41041.09231.09621.09623.44502.83202.8320
C52.49643.67091.47122.53632.57603.41702.83222.83221.09501.09691.0969
H61.10232.07272.05763.41042.57603.55034.05064.05062.16373.33683.3368
H72.46302.31002.07011.09233.41703.55031.79261.79264.17003.78563.7856
H83.18693.58442.12761.09622.83224.05061.79261.78643.80493.22662.6882
H93.18693.58442.12761.09622.83224.05061.79261.78643.80492.68823.2266
H102.56533.81732.10013.44501.09502.16374.17003.80493.80491.78201.7820
H113.21564.31592.12472.83201.09693.33683.78563.22662.68821.78201.7823
H123.21564.31592.12472.83201.09693.33683.78562.68823.22661.78201.7823

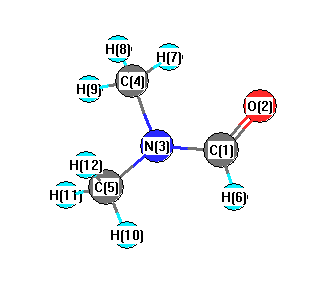
picture of dimethylformamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N3 C4 119.166 C1 N3 C5 122.158
O2 C1 N3 125.293 O2 C1 H6 123.252
N3 C1 H6 111.455 N3 C4 H7 106.377
N3 C4 H8 110.658 N3 C4 H9 110.658
N3 C5 H10 108.965 N3 C5 H11 110.805
N3 C5 H12 110.805 C4 N3 C5 118.676
H7 C4 H8 109.993 H7 C4 H9 109.993
H8 C4 H9 109.135 H10 C5 H11 108.778
H10 C5 H12 108.778 H11 C5 H12 108.661
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability