return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7NO (dimethylformamide)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-246.116567
Energy at 298.15K-246.125027
HF Energy-245.609795
Nuclear repulsion energy178.970503
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3185 3030 0.98      
2 A' 3151 2997 18.10      
3 A' 3068 2919 37.52      
4 A' 3059 2910 40.02      
5 A' 2993 2847 101.36      
6 A' 1692 1610 223.00      
7 A' 1627 1548 6.44      
8 A' 1609 1530 15.48      
9 A' 1527 1453 12.41      
10 A' 1499 1426 16.20      
11 A' 1470 1399 39.85      
12 A' 1421 1351 52.82      
13 A' 1274 1212 47.28      
14 A' 1109 1055 122.71      
15 A' 1106 1052 29.83      
16 A' 864 822 1.24      
17 A' 652 620 5.59      
18 A' 381 362 1.19      
19 A' 323 307 11.92      
20 A" 3126 2974 29.45      
21 A" 3114 2962 41.66      
22 A" 1604 1526 12.42      
23 A" 1580 1503 5.13      
24 A" 1217 1158 3.64      
25 A" 1173 1115 0.11      
26 A" 1060 1008 0.07      
27 A" 341 324 20.85      
28 A" 223 213 3.03      
29 A" 206 196 1.60      
30 A" 136 129 0.36      

Unscaled Zero Point Vibrational Energy (zpe) 22893.7 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 21778.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.29050 0.13536 0.09570

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.722 -0.816 0.000
O2 0.229 -1.967 0.000
N3 0.000 0.361 0.000
C4 -1.476 0.300 0.000
C5 0.652 1.680 0.000
H6 1.807 -0.623 0.000
H7 -1.741 -0.760 0.000
H8 -1.884 0.788 0.893
H9 -1.884 0.788 -0.893
H10 1.738 1.539 0.000
H11 0.370 2.254 -0.891
H12 0.370 2.254 0.891

Atom - Atom Distances (Å)
  C1 O2 N3 C4 C5 H6 H7 H8 H9 H10 H11 H12
C11.25221.38072.46552.49691.10242.46353.18743.18742.56563.21613.2161
O21.25222.33922.83703.67152.07302.31053.58503.58503.81774.31654.3165
N31.38072.33921.47771.47162.05782.07042.12802.12802.10042.12502.1250
C42.46552.83701.47772.53703.41101.09241.09641.09643.44572.83262.8326
C52.49693.67151.47162.53702.57633.41772.83282.83281.09511.09711.0971
H61.10242.07302.05783.41102.57633.55094.05124.05122.16363.33713.3371
H72.46352.31052.07041.09243.41773.55091.79291.79294.17063.78633.7863
H83.18743.58502.12801.09642.83284.05121.79291.78683.80563.22722.6888
H93.18743.58502.12801.09642.83284.05121.79291.78683.80562.68883.2272
H102.56563.81772.10043.44571.09512.16364.17063.80563.80561.78231.7823
H113.21614.31652.12502.83261.09713.33713.78633.22722.68881.78231.7826
H123.21614.31652.12502.83261.09713.33713.78632.68883.22721.78231.7826

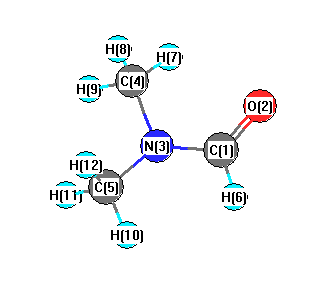
picture of dimethylformamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N3 C4 119.168 C1 N3 C5 122.155
O2 C1 N3 125.294 O2 C1 H6 123.263
N3 C1 H6 111.444 N3 C4 H7 106.368
N3 C4 H8 110.650 N3 C4 H9 110.650
N3 C5 H10 108.954 N3 C5 H11 110.799
N3 C5 H12 110.799 C4 N3 C5 118.678
H7 C4 H8 110.000 H7 C4 H9 110.000
H8 C4 H9 109.146 H10 C5 H11 108.785
H10 C5 H12 108.785 H11 C5 H12 108.672
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability