return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10O (Oxetane, 3,3-dimethyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-269.058635
Energy at 298.15K-269.070427
HF Energy-268.479872
Nuclear repulsion energy253.405504
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3162 3008 59.12      
2 A' 3155 3001 17.16      
3 A' 3144 2991 12.86      
4 A' 3087 2936 25.68      
5 A' 3071 2922 18.87      
6 A' 3069 2920 15.90      
7 A' 1617 1538 1.43      
8 A' 1601 1523 4.82      
9 A' 1588 1511 3.44      
10 A' 1496 1423 2.91      
11 A' 1476 1404 13.12      
12 A' 1378 1311 2.68      
13 A' 1327 1263 0.81      
14 A' 1297 1234 4.31      
15 A' 1138 1083 0.43      
16 A' 1040 989 2.31      
17 A' 1020 970 1.65      
18 A' 979 931 0.43      
19 A' 884 841 1.13      
20 A' 787 749 11.83      
21 A' 618 588 0.70      
22 A' 412 392 0.16      
23 A' 349 332 0.22      
24 A' 120 114 8.38      
25 A" 3158 3005 15.03      
26 A" 3146 2993 34.27      
27 A" 3142 2989 0.43      
28 A" 3079 2929 58.11      
29 A" 1609 1531 10.03      
30 A" 1591 1513 0.43      
31 A" 1589 1512 0.22      
32 A" 1309 1245 9.41      
33 A" 1276 1214 0.87      
34 A" 1134 1079 0.00      
35 A" 1107 1053 0.24      
36 A" 992 944 2.58      
37 A" 927 881 28.92      
38 A" 875 833 6.57      
39 A" 399 380 0.25      
40 A" 322 306 0.70      
41 A" 302 287 0.02      
42 A" 240 228 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 32004.5 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 30445.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.16899 0.10310 0.09365

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.940 1.623 0.000
H2 -0.765 2.241 0.889
H3 -0.765 2.241 -0.889
H4 -1.987 1.298 0.000
C5 1.486 0.816 0.000
H6 2.104 -0.088 0.000
H7 1.721 1.411 -0.891
H8 1.721 1.411 0.891
C9 0.000 0.414 0.000
C10 -0.263 -0.702 1.063
C11 -0.263 -0.702 -1.063
H12 0.456 -0.794 1.881
H13 0.456 -0.794 -1.881
H14 -1.293 -0.700 1.438
H15 -1.293 -0.700 -1.438
O16 -0.059 -1.778 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 C10 C11 H12 H13 H14 H15 O16
C11.09661.09661.09662.55643.49222.81462.81461.53122.64442.64443.36563.36562.75432.75433.5133
H21.09661.77721.78152.80783.80083.16802.62102.17072.99043.56683.41854.28613.03803.78684.1764
H31.09661.77721.78152.80783.80082.62103.16802.17073.56682.99044.28613.41853.78683.03804.1764
H41.09661.78151.78153.50664.32033.81603.81602.17532.84692.84693.72643.72642.55772.55773.6309
C52.55642.80782.80783.50661.09581.09671.09671.53932.54832.54832.68152.68153.47683.47683.0200
H63.49223.80083.80084.32031.09581.78551.78552.16342.66672.66672.59832.59833.73943.73942.7455
H72.81463.16802.62103.81601.09671.78551.78182.17963.49592.90373.76092.72814.35493.72043.7599
H82.81462.62103.16803.81601.09671.78551.78182.17962.90373.49592.72813.76093.72044.35493.7599
C91.53122.17072.17072.17531.53932.16342.17962.17961.56371.56372.28132.28132.23162.23162.1933
C102.64442.99043.56682.84692.54832.66673.49592.90371.56372.12691.09243.03201.09592.70501.5269
C112.64443.56682.99042.84692.54832.66672.90373.49591.56372.12693.03201.09242.70501.09591.5269
H123.36563.41854.28613.72642.68152.59833.76092.72812.28131.09243.03203.76121.80683.75232.1844
H133.36564.28613.41853.72642.68152.59832.72813.76092.28133.03201.09243.76123.75231.80682.1844
H142.75433.03803.78682.55773.47683.73944.35493.72042.23161.09592.70501.80683.75232.87552.1800
H152.75433.78683.03802.55773.47683.73943.72044.35492.23162.70501.09593.75231.80682.87552.1800
O163.51334.17644.17643.63093.02002.74553.75993.75992.19331.52691.52692.18442.18442.18002.1800

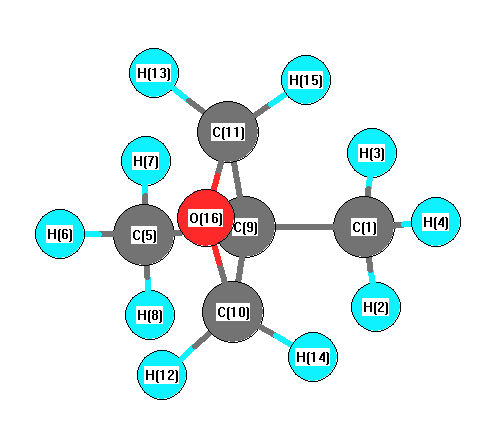
picture of Oxetane, 3,3-dimethyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C9 C5 112.724 C1 C9 C10 117.390
C1 C9 C11 117.390 H2 C1 H3 108.250
H2 C1 H4 108.640 H2 C1 C9 110.294
H3 C1 H4 108.640 H3 C1 C9 110.294
H4 C1 C9 110.658 C5 C9 C10 110.415
C5 C9 C11 110.415 H6 C5 H7 109.048
H6 C5 H8 109.048 H6 C5 C9 109.212
H7 C5 H8 108.651 H7 C5 C9 110.427
H8 C5 C9 110.427 C9 C10 H12 117.275
C9 C10 H14 112.896 C9 C10 O16 90.405
C9 C11 H13 117.275 C9 C11 H15 112.896
C9 C11 O16 90.405 C10 C9 C11 85.698
C10 O16 C11 88.289 H12 C10 H14 111.307
H12 C10 O16 111.940 H13 C11 H15 111.307
H13 C11 O16 111.940 H14 C10 O16 111.375
H15 C11 O16 111.375
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability