return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (1-Hexyne)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-232.228358
Energy at 298.15K-232.237711
HF Energy-231.679326
Nuclear repulsion energy208.542013
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3484 3314 43.64      
2 A' 3155 3002 29.27      
3 A' 3091 2940 36.85      
4 A' 3081 2931 16.90      
5 A' 3075 2925 8.92      
6 A' 3073 2923 6.18      
7 A' 2132 2029 0.07      
8 A' 1607 1529 5.17      
9 A' 1595 1517 2.55      
10 A' 1585 1508 0.27      
11 A' 1573 1497 1.29      
12 A' 1493 1421 3.80      
13 A' 1431 1361 0.52      
14 A' 1409 1340 3.43      
15 A' 1340 1275 0.86      
16 A' 1157 1100 6.78      
17 A' 1050 999 0.09      
18 A' 1002 953 0.76      
19 A' 941 895 0.46      
20 A' 895 852 4.23      
21 A' 617 587 54.47      
22 A' 497 472 1.88      
23 A' 388 369 0.17      
24 A' 259 247 0.77      
25 A' 114 108 0.13      
26 A" 3159 3005 57.76      
27 A" 3143 2990 13.42      
28 A" 3113 2961 0.02      
29 A" 3107 2956 5.01      
30 A" 1603 1525 7.07      
31 A" 1394 1326 0.01      
32 A" 1382 1315 1.03      
33 A" 1312 1248 0.15      
34 A" 1196 1137 0.05      
35 A" 992 944 0.53      
36 A" 833 792 0.61      
37 A" 757 720 3.73      
38 A" 615 585 52.80      
39 A" 303 288 2.96      
40 A" 252 240 0.18      
41 A" 119 113 0.00      
42 A" 89 85 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 31706.4 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 30162.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.48014 0.04328 0.04092

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.275 2.646 0.000
H2 2.266 3.114 0.000
H3 0.731 2.986 0.888
H4 0.731 2.986 -0.888
C5 1.391 1.100 0.000
H6 1.952 0.773 0.885
H7 1.952 0.773 -0.885
C8 0.000 0.420 0.000
H9 -0.564 0.738 0.886
H10 -0.564 0.738 -0.886
C11 0.111 -1.134 0.000
H12 0.678 -1.451 0.886
H13 0.678 -1.451 -0.886
C14 -2.321 -2.318 0.000
H15 -3.285 -2.772 0.000
C16 -1.219 -1.787 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 C11 H12 H13 C14 H15 C16
C11.09541.09561.09561.55032.17992.17992.56532.79452.79453.95554.23424.23426.13057.08135.0866
H21.09541.77781.77782.19512.52232.52233.51973.79963.79964.76304.91344.91347.10988.08976.0133
H31.09561.77781.77672.18632.52743.08762.81162.59443.14304.25984.43704.77866.18407.07555.2317
H41.09561.77781.77672.18633.08762.52742.81163.14302.59444.25984.77864.43706.18407.07555.2317
C51.55032.19512.18632.18631.09791.09791.54862.17682.17682.57522.79332.79335.04706.07093.8923
H62.17992.52232.52743.08761.09791.77052.17232.51613.07692.79452.56293.11535.34816.38524.1704
H72.17992.52233.08762.52741.09791.77052.17233.07692.51612.79453.11532.56295.34816.38524.1704
C82.56533.51972.81162.81161.54862.17232.17231.09691.09691.55832.17862.17863.59024.58012.5215
H92.79453.79962.59443.14302.17682.51613.07691.09691.77162.17782.51633.07753.63504.52802.7544
H102.79453.79963.14302.59442.17683.07692.51611.09691.77162.17783.07752.51633.63504.52802.7544
C113.95554.76304.25984.25982.57522.79452.79451.55832.17782.17781.09861.09862.70543.76991.4816
H124.23424.91344.43704.77862.79332.56293.11532.17862.51633.07751.09861.77193.24564.26992.1206
H134.23424.91344.77864.43702.79333.11532.56292.17863.07752.51631.09861.77193.24564.26992.1206
C146.13057.10986.18406.18405.04705.34815.34813.59023.63503.63502.70543.24563.24561.06451.2237
H157.08138.08977.07557.07556.07096.38526.38524.58014.52804.52803.76994.26994.26991.06452.2883
C165.08666.01335.23175.23173.89234.17044.17042.52152.75442.75441.48162.12062.12061.22372.2883

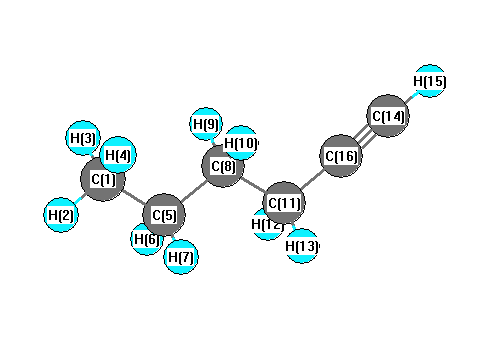
picture of 1-Hexyne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 109.617 C1 C5 H7 109.617
C1 C5 C8 111.745 H2 C1 H3 108.469
H2 C1 H4 108.469 H2 C1 C5 110.961
H3 C1 H4 108.356 H3 C1 C5 110.256
H4 C1 C5 110.256 C5 C8 H9 109.557
C5 C8 H10 109.557 C5 C8 C11 111.959
H6 C5 H7 107.474 H6 C5 C8 109.144
H7 C5 C8 109.144 C8 C11 H12 108.937
C8 C11 H13 108.937 C8 C11 C16 112.058
H9 C8 H10 107.721 H9 C8 C11 108.974
H10 C8 C11 108.974 C11 C16 C14 179.605
H12 C11 H13 107.494 H12 C11 C16 109.651
H13 C11 C16 109.651 H15 C14 C16 179.447
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability