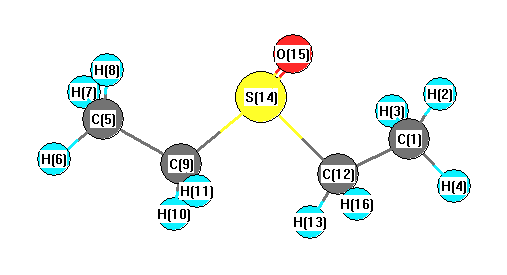
Vibrational Frequencies calculated at B3PW91/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3152 |
3024 |
15.69 |
|
|
|
| 2 |
A |
3146 |
3019 |
22.69 |
|
|
|
| 3 |
A |
3142 |
3014 |
4.31 |
|
|
|
| 4 |
A |
3135 |
3008 |
15.50 |
|
|
|
| 5 |
A |
3127 |
3000 |
3.15 |
|
|
|
| 6 |
A |
3123 |
2997 |
1.56 |
|
|
|
| 7 |
A |
3082 |
2957 |
13.07 |
|
|
|
| 8 |
A |
3081 |
2956 |
2.82 |
|
|
|
| 9 |
A |
3062 |
2938 |
14.23 |
|
|
|
| 10 |
A |
3055 |
2931 |
18.04 |
|
|
|
| 11 |
A |
1571 |
1507 |
10.86 |
|
|
|
| 12 |
A |
1562 |
1499 |
14.66 |
|
|
|
| 13 |
A |
1558 |
1494 |
8.59 |
|
|
|
| 14 |
A |
1555 |
1492 |
17.67 |
|
|
|
| 15 |
A |
1518 |
1456 |
9.54 |
|
|
|
| 16 |
A |
1507 |
1446 |
12.86 |
|
|
|
| 17 |
A |
1460 |
1401 |
14.78 |
|
|
|
| 18 |
A |
1453 |
1394 |
26.02 |
|
|
|
| 19 |
A |
1326 |
1272 |
1.35 |
|
|
|
| 20 |
A |
1310 |
1257 |
0.78 |
|
|
|
| 21 |
A |
1289 |
1236 |
1.07 |
|
|
|
| 22 |
A |
1281 |
1229 |
1.34 |
|
|
|
| 23 |
A |
1109 |
1064 |
14.00 |
|
|
|
| 24 |
A |
1094 |
1049 |
67.39 |
|
|
|
| 25 |
A |
1068 |
1024 |
12.56 |
|
|
|
| 26 |
A |
1059 |
1016 |
18.88 |
|
|
|
| 27 |
A |
1045 |
1003 |
5.68 |
|
|
|
| 28 |
A |
993 |
952 |
0.82 |
|
|
|
| 29 |
A |
989 |
949 |
2.10 |
|
|
|
| 30 |
A |
825 |
792 |
17.92 |
|
|
|
| 31 |
A |
788 |
756 |
9.74 |
|
|
|
| 32 |
A |
666 |
639 |
30.43 |
|
|
|
| 33 |
A |
585 |
561 |
10.09 |
|
|
|
| 34 |
A |
501 |
481 |
3.94 |
|
|
|
| 35 |
A |
362 |
347 |
0.76 |
|
|
|
| 36 |
A |
309 |
296 |
2.08 |
|
|
|
| 37 |
A |
271 |
260 |
6.03 |
|
|
|
| 38 |
A |
221 |
212 |
5.38 |
|
|
|
| 39 |
A |
203 |
195 |
3.81 |
|
|
|
| 40 |
A |
187 |
179 |
0.64 |
|
|
|
| 41 |
A |
126 |
121 |
1.12 |
|
|
|
| 42 |
A |
73 |
70 |
0.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30484.7 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 29247.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.653 |
|
|
|
| 2 |
H |
0.225 |
|
|
|
| 3 |
H |
0.213 |
|
|
|
| 4 |
H |
0.255 |
|
|
|
| 5 |
C |
-0.658 |
|
|
|
| 6 |
H |
0.231 |
|
|
|
| 7 |
H |
0.237 |
|
|
|
| 8 |
H |
0.221 |
|
|
|
| 9 |
C |
-0.688 |
|
|
|
| 10 |
H |
0.258 |
|
|
|
| 11 |
H |
0.254 |
|
|
|
| 12 |
C |
-0.689 |
|
|
|
| 13 |
H |
0.238 |
|
|
|
| 14 |
S |
0.844 |
|
|
|
| 15 |
O |
-0.540 |
|
|
|
| 16 |
H |
0.251 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.960 |
-2.986 |
-0.526 |
3.610 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.679 |
3.659 |
2.181 |
| y |
3.659 |
-49.197 |
-0.531 |
| z |
2.181 |
-0.531 |
-44.642 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.241 |
3.659 |
2.181 |
| y |
3.659 |
-5.037 |
-0.531 |
| z |
2.181 |
-0.531 |
1.796 |
|
| Polar |
|---|
| 3z2-r2 | 3.591 |
|---|
| x2-y2 | 5.518 |
|---|
| xy | 3.659 |
|---|
| xz | 2.181 |
|---|
| yz | -0.531 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
225.872 |
| (<r2>)1/2 |
15.029 |