Jump to
S1C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -192.435408 |
| Energy at 298.15K | -192.444411 |
| Nuclear repulsion energy | 128.811795 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3510 |
3339 |
0.47 |
|
|
|
| 2 |
A' |
3160 |
3006 |
26.83 |
|
|
|
| 3 |
A' |
3112 |
2961 |
15.74 |
|
|
|
| 4 |
A' |
3082 |
2932 |
18.25 |
|
|
|
| 5 |
A' |
3031 |
2883 |
41.49 |
|
|
|
| 6 |
A' |
1632 |
1553 |
3.09 |
|
|
|
| 7 |
A' |
1607 |
1529 |
4.84 |
|
|
|
| 8 |
A' |
1596 |
1518 |
2.22 |
|
|
|
| 9 |
A' |
1508 |
1435 |
7.78 |
|
|
|
| 10 |
A' |
1490 |
1417 |
2.29 |
|
|
|
| 11 |
A' |
1403 |
1334 |
2.31 |
|
|
|
| 12 |
A' |
1295 |
1232 |
48.90 |
|
|
|
| 13 |
A' |
1134 |
1079 |
7.99 |
|
|
|
| 14 |
A' |
1042 |
991 |
17.06 |
|
|
|
| 15 |
A' |
1008 |
959 |
40.80 |
|
|
|
| 16 |
A' |
898 |
854 |
11.00 |
|
|
|
| 17 |
A' |
452 |
430 |
11.29 |
|
|
|
| 18 |
A' |
273 |
260 |
7.22 |
|
|
|
| 19 |
A" |
3168 |
3014 |
51.08 |
|
|
|
| 20 |
A" |
3141 |
2988 |
2.90 |
|
|
|
| 21 |
A" |
3063 |
2914 |
58.20 |
|
|
|
| 22 |
A" |
1606 |
1528 |
7.53 |
|
|
|
| 23 |
A" |
1387 |
1320 |
0.89 |
|
|
|
| 24 |
A" |
1325 |
1260 |
0.00 |
|
|
|
| 25 |
A" |
1221 |
1161 |
0.71 |
|
|
|
| 26 |
A" |
960 |
914 |
3.41 |
|
|
|
| 27 |
A" |
809 |
769 |
0.95 |
|
|
|
| 28 |
A" |
298 |
284 |
136.68 |
|
|
|
| 29 |
A" |
246 |
234 |
6.39 |
|
|
|
| 30 |
A" |
134 |
128 |
5.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24295.1 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 23112.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.464 |
1.244 |
0.000 |
| C2 |
0.000 |
0.742 |
0.000 |
| C3 |
0.073 |
-0.791 |
0.000 |
| O4 |
1.508 |
-1.126 |
0.000 |
| H5 |
-1.499 |
2.338 |
0.000 |
| H6 |
-1.994 |
0.884 |
0.889 |
| H7 |
-1.994 |
0.884 |
-0.889 |
| H8 |
0.533 |
1.106 |
0.884 |
| H9 |
0.533 |
1.106 |
-0.884 |
| H10 |
-0.440 |
-1.184 |
0.890 |
| H11 |
-0.440 |
-1.184 |
-0.890 |
| H12 |
1.585 |
-2.115 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5478 | 2.5506 | 3.8008 | 1.0946 | 1.0960 | 1.0960 | 2.1881 | 2.1881 | 2.7816 | 2.7816 | 4.5367 |
C2 | 1.5478 | | 1.5344 | 2.3999 | 2.1899 | 2.1879 | 2.1879 | 1.0947 | 1.0947 | 2.1669 | 2.1669 | 3.2670 | C3 | 2.5506 | 1.5344 | | 1.4727 | 3.5020 | 2.8052 | 2.8052 | 2.1429 | 2.1429 | 1.1004 | 1.1004 | 2.0096 | O4 | 3.8008 | 2.3999 | 1.4727 | | 4.5866 | 4.1339 | 4.1339 | 2.5908 | 2.5908 | 2.1425 | 2.1425 | 0.9924 | H5 | 1.0946 | 2.1899 | 3.5020 | 4.5866 | | 1.7753 | 1.7753 | 2.5352 | 2.5352 | 3.7843 | 3.7843 | 5.4169 | H6 | 1.0960 | 2.1879 | 2.8052 | 4.1339 | 1.7753 | | 1.7774 | 2.5369 | 3.0949 | 2.5865 | 3.1390 | 4.7533 | H7 | 1.0960 | 2.1879 | 2.8052 | 4.1339 | 1.7753 | 1.7774 | | 3.0949 | 2.5369 | 3.1390 | 2.5865 | 4.7533 | H8 | 2.1881 | 1.0947 | 2.1429 | 2.5908 | 2.5352 | 2.5369 | 3.0949 | | 1.7682 | 2.4887 | 3.0563 | 3.5021 | H9 | 2.1881 | 1.0947 | 2.1429 | 2.5908 | 2.5352 | 3.0949 | 2.5369 | 1.7682 | | 3.0563 | 2.4887 | 3.5021 | H10 | 2.7816 | 2.1669 | 1.1004 | 2.1425 | 3.7843 | 2.5865 | 3.1390 | 2.4887 | 3.0563 | | 1.7799 | 2.4004 | H11 | 2.7816 | 2.1669 | 1.1004 | 2.1425 | 3.7843 | 3.1390 | 2.5865 | 3.0563 | 2.4887 | 1.7799 | | 2.4004 | H12 | 4.5367 | 3.2670 | 2.0096 | 0.9924 | 5.4169 | 4.7533 | 4.7533 | 3.5021 | 3.5021 | 2.4004 | 2.4004 | |
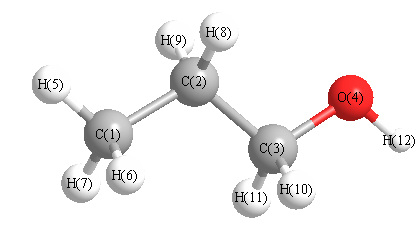 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.689 |
|
C1 |
C2 |
H8 |
110.625 |
| C1 |
C2 |
H9 |
110.625 |
|
C2 |
C1 |
H5 |
110.772 |
| C2 |
C1 |
H6 |
110.537 |
|
C2 |
C1 |
H7 |
110.537 |
| C2 |
C3 |
O4 |
105.873 |
|
C2 |
C3 |
H10 |
109.552 |
| C2 |
C3 |
H11 |
109.552 |
|
C3 |
C2 |
H8 |
108.017 |
| C3 |
C2 |
H9 |
108.017 |
|
C3 |
O4 |
H12 |
107.621 |
| O4 |
C3 |
H10 |
111.937 |
|
O4 |
C3 |
H11 |
111.937 |
| H5 |
C1 |
H6 |
108.274 |
|
H5 |
C1 |
H7 |
108.274 |
| H6 |
C1 |
H7 |
108.362 |
|
H8 |
C2 |
H9 |
107.726 |
| H10 |
C3 |
H11 |
107.959 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -192.436821 |
| Energy at 298.15K | |
| HF Energy | -192.041426 |
| Nuclear repulsion energy | 131.728400 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3516 |
3345 |
0.73 |
131.89 |
0.32 |
0.48 |
| 2 |
A |
3198 |
3042 |
13.37 |
27.52 |
0.71 |
0.83 |
| 3 |
A |
3158 |
3004 |
45.03 |
36.60 |
0.60 |
0.75 |
| 4 |
A |
3142 |
2989 |
10.00 |
77.38 |
0.73 |
0.84 |
| 5 |
A |
3099 |
2948 |
30.79 |
86.14 |
0.10 |
0.18 |
| 6 |
A |
3087 |
2937 |
23.14 |
76.93 |
0.06 |
0.11 |
| 7 |
A |
3070 |
2921 |
52.14 |
70.42 |
0.75 |
0.86 |
| 8 |
A |
3035 |
2888 |
45.91 |
79.51 |
0.16 |
0.27 |
| 9 |
A |
1627 |
1547 |
2.83 |
8.57 |
0.70 |
0.83 |
| 10 |
A |
1618 |
1539 |
5.80 |
15.36 |
0.75 |
0.86 |
| 11 |
A |
1598 |
1520 |
7.45 |
26.62 |
0.75 |
0.86 |
| 12 |
A |
1579 |
1502 |
2.96 |
16.92 |
0.75 |
0.85 |
| 13 |
A |
1502 |
1429 |
8.57 |
1.66 |
0.68 |
0.81 |
| 14 |
A |
1475 |
1403 |
4.83 |
3.15 |
0.69 |
0.82 |
| 15 |
A |
1443 |
1372 |
0.21 |
4.31 |
0.75 |
0.85 |
| 16 |
A |
1383 |
1316 |
8.32 |
18.47 |
0.73 |
0.84 |
| 17 |
A |
1315 |
1251 |
8.73 |
8.80 |
0.74 |
0.85 |
| 18 |
A |
1286 |
1223 |
27.78 |
15.67 |
0.74 |
0.85 |
| 19 |
A |
1193 |
1135 |
6.14 |
4.43 |
0.65 |
0.79 |
| 20 |
A |
1150 |
1094 |
6.44 |
2.66 |
0.63 |
0.78 |
| 21 |
A |
1054 |
1003 |
22.92 |
3.15 |
0.65 |
0.79 |
| 22 |
A |
989 |
941 |
34.98 |
1.48 |
0.43 |
0.60 |
| 23 |
A |
966 |
919 |
17.59 |
4.74 |
0.67 |
0.80 |
| 24 |
A |
869 |
827 |
2.99 |
10.40 |
0.17 |
0.29 |
| 25 |
A |
803 |
764 |
1.52 |
1.74 |
0.31 |
0.47 |
| 26 |
A |
489 |
465 |
8.72 |
0.45 |
0.59 |
0.75 |
| 27 |
A |
339 |
322 |
16.17 |
0.88 |
0.71 |
0.83 |
| 28 |
A |
280 |
267 |
125.82 |
8.16 |
0.74 |
0.85 |
| 29 |
A |
254 |
241 |
3.37 |
0.12 |
0.74 |
0.85 |
| 30 |
A |
164 |
156 |
8.00 |
0.12 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 24339.7 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 23154.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.497 |
-0.548 |
0.141 |
| C2 |
-0.648 |
0.670 |
-0.299 |
| C3 |
0.766 |
0.592 |
0.295 |
| O4 |
1.343 |
-0.663 |
-0.224 |
| H5 |
-2.477 |
-0.543 |
-0.346 |
| H6 |
-1.648 |
-0.534 |
1.226 |
| H7 |
-0.959 |
-1.459 |
-0.128 |
| H8 |
-0.554 |
0.680 |
-1.390 |
| H9 |
-1.120 |
1.608 |
0.016 |
| H10 |
0.703 |
0.587 |
1.393 |
| H11 |
1.358 |
1.467 |
-0.013 |
| H12 |
2.225 |
-0.793 |
0.212 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5473 | 2.5384 | 2.8655 | 1.0948 | 1.0964 | 1.0916 | 2.1769 | 2.1923 | 2.7738 | 3.4970 | 3.7302 |
C2 | 1.5473 | | 1.5360 | 2.3972 | 2.1948 | 2.1847 | 2.1574 | 1.0952 | 1.0966 | 2.1671 | 2.1777 | 3.2642 | C3 | 2.5384 | 1.5360 | | 1.4754 | 3.4956 | 2.8216 | 2.7128 | 2.1424 | 2.1600 | 1.1001 | 1.0995 | 2.0133 | O4 | 2.8655 | 2.3972 | 1.4754 | | 3.8243 | 3.3262 | 2.4374 | 2.6009 | 3.3586 | 2.1410 | 2.1396 | 0.9921 | H5 | 1.0948 | 2.1948 | 3.4956 | 3.8243 | | 1.7783 | 1.7868 | 2.5067 | 2.5696 | 3.7968 | 4.3426 | 4.7417 | H6 | 1.0964 | 2.1847 | 2.8216 | 3.3262 | 1.7783 | | 1.7787 | 3.0847 | 2.5165 | 2.6092 | 3.8169 | 4.0112 | H7 | 1.0916 | 2.1574 | 2.7128 | 2.4374 | 1.7868 | 1.7787 | | 2.5164 | 3.0746 | 3.0424 | 3.7331 | 3.2700 | H8 | 2.1769 | 1.0952 | 2.1424 | 2.6009 | 2.5067 | 3.0847 | 2.5164 | | 1.7768 | 3.0555 | 2.4844 | 3.5301 | H9 | 2.1923 | 1.0966 | 2.1600 | 3.3586 | 2.5696 | 2.5165 | 3.0746 | 1.7768 | | 2.5027 | 2.4817 | 4.1218 | H10 | 2.7738 | 2.1671 | 1.1001 | 2.1410 | 3.7968 | 2.6092 | 3.0424 | 3.0555 | 2.5027 | | 1.7830 | 2.3690 | H11 | 3.4970 | 2.1777 | 1.0995 | 2.1396 | 4.3426 | 3.8169 | 3.7331 | 2.4844 | 2.4817 | 1.7830 | | 2.4303 | H12 | 3.7302 | 3.2642 | 2.0133 | 0.9921 | 4.7417 | 4.0112 | 3.2700 | 3.5301 | 4.1218 | 2.3690 | 2.4303 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
110.830 |
|
C1 |
C2 |
H8 |
109.748 |
| C1 |
C2 |
H9 |
110.875 |
|
C2 |
C1 |
H5 |
111.185 |
| C2 |
C1 |
H6 |
110.287 |
|
C2 |
C1 |
H7 |
108.445 |
| C2 |
C3 |
O4 |
105.491 |
|
C2 |
C3 |
H10 |
109.478 |
| C2 |
C3 |
H11 |
110.349 |
|
C3 |
C2 |
H8 |
107.851 |
| C3 |
C2 |
H9 |
109.132 |
|
C3 |
O4 |
H12 |
107.746 |
| O4 |
C3 |
H10 |
111.635 |
|
O4 |
C3 |
H11 |
111.558 |
| H5 |
C1 |
H6 |
108.499 |
|
H5 |
C1 |
H7 |
109.621 |
| H6 |
C1 |
H7 |
108.770 |
|
H8 |
C2 |
H9 |
108.319 |
| H10 |
C3 |
H11 |
108.310 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
