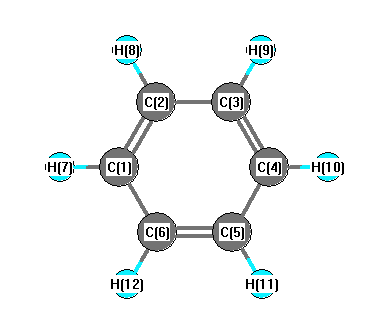
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3223 |
3066 |
0.00 |
302.47 |
0.10 |
0.19 |
| 2 |
A1g |
993 |
944 |
0.00 |
49.06 |
0.12 |
0.21 |
| 3 |
A2g |
1435 |
1366 |
0.00 |
0.00 |
0.00 |
0.00 |
| 4 |
A2u |
684 |
651 |
122.50 |
0.00 |
0.00 |
0.00 |
| 5 |
B1u |
3183 |
3028 |
0.00 |
0.00 |
0.75 |
0.86 |
| 6 |
B1u |
1068 |
1016 |
0.00 |
0.00 |
0.00 |
0.00 |
| 7 |
B2g |
933 |
888 |
0.00 |
0.00 |
0.75 |
0.86 |
| 8 |
B2g |
678 |
645 |
0.00 |
0.00 |
0.00 |
0.00 |
| 9 |
B2u |
1319 |
1255 |
0.00 |
0.00 |
0.00 |
0.00 |
| 10 |
B2u |
1237 |
1177 |
0.00 |
0.00 |
0.00 |
0.00 |
| 11 |
E1g |
852 |
811 |
0.00 |
7.32 |
0.75 |
0.86 |
| 11 |
E1g |
852 |
811 |
0.00 |
7.32 |
0.75 |
0.86 |
| 12 |
E1u |
3210 |
3054 |
30.93 |
0.00 |
0.00 |
0.00 |
| 12 |
E1u |
3210 |
3054 |
30.93 |
0.00 |
0.00 |
0.00 |
| 13 |
E1u |
1533 |
1458 |
11.39 |
0.00 |
0.00 |
0.00 |
| 13 |
E1u |
1533 |
1458 |
11.39 |
0.00 |
0.00 |
0.00 |
| 14 |
E1u |
1056 |
1005 |
3.04 |
0.00 |
0.00 |
0.00 |
| 14 |
E1u |
1056 |
1005 |
3.04 |
0.00 |
0.00 |
0.00 |
| 15 |
E2g |
3192 |
3037 |
0.00 |
113.20 |
0.75 |
0.86 |
| 15 |
E2g |
3192 |
3037 |
0.00 |
113.20 |
0.75 |
0.86 |
| 16 |
E2g |
1599 |
1521 |
0.00 |
12.25 |
0.75 |
0.86 |
| 16 |
E2g |
1599 |
1521 |
0.00 |
12.25 |
0.75 |
0.86 |
| 17 |
E2g |
1247 |
1186 |
0.00 |
7.58 |
0.75 |
0.86 |
| 17 |
E2g |
1247 |
1186 |
0.00 |
7.58 |
0.75 |
0.86 |
| 18 |
E2g |
646 |
614 |
0.00 |
3.38 |
0.75 |
0.86 |
| 18 |
E2g |
646 |
614 |
0.00 |
3.38 |
0.75 |
0.86 |
| 19 |
E2u |
931 |
886 |
0.00 |
0.00 |
0.00 |
0.00 |
| 19 |
E2u |
931 |
886 |
0.00 |
0.00 |
0.00 |
0.00 |
| 20 |
E2u |
412 |
392 |
0.00 |
0.00 |
0.00 |
0.00 |
| 20 |
E2u |
412 |
392 |
0.00 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 22054.9 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 20980.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.