return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (1,4-Hexadiene, (E)-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-232.231925
Energy at 298.15K-232.241606
HF Energy-231.691295
Nuclear repulsion energy213.598009
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3265 3106 16.02      
2 A 3184 3029 9.45      
3 A 3166 3012 16.32      
4 A 3164 3010 44.59      
5 A 3155 3001 1.78      
6 A 3143 2990 5.68      
7 A 3133 2981 19.21      
8 A 3093 2942 13.58      
9 A 3072 2922 18.95      
10 A 3056 2907 18.58      
11 A 1712 1629 0.39      
12 A 1691 1608 5.24      
13 A 1598 1520 7.26      
14 A 1586 1509 5.96      
15 A 1571 1494 8.72      
16 A 1521 1447 1.36      
17 A 1493 1420 4.18      
18 A 1408 1340 1.73      
19 A 1388 1321 1.15      
20 A 1383 1315 1.86      
21 A 1369 1302 0.93      
22 A 1283 1221 0.72      
23 A 1165 1108 4.32      
24 A 1144 1088 3.63      
25 A 1131 1076 0.11      
26 A 1059 1007 15.59      
27 A 1045 995 2.96      
28 A 1025 975 36.60      
29 A 1009 960 8.31      
30 A 963 916 34.01      
31 A 907 863 2.16      
32 A 901 857 1.00      
33 A 789 750 0.27      
34 A 622 592 4.99      
35 A 559 532 8.21      
36 A 471 448 0.26      
37 A 329 313 0.83      
38 A 289 275 1.27      
39 A 199 189 1.34      
40 A 155 148 0.42      
41 A 128 122 0.03      
42 A 73 70 0.13      

Unscaled Zero Point Vibrational Energy (zpe) 31698.0 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 30154.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.26297 0.05812 0.05082

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.747 0.464 0.039
H2 2.942 1.344 -0.585
H3 2.688 0.786 1.085
H4 3.600 -0.217 -0.066
C5 1.453 -0.219 -0.376
H6 1.407 -0.578 -1.405
C7 0.397 -0.406 0.433
H8 0.434 -0.041 1.461
C9 -0.897 -1.082 0.016
H10 -0.757 -1.564 -0.963
H11 -1.140 -1.880 0.733
C12 -2.084 -0.117 -0.064
H13 -3.063 -0.586 -0.155
C14 -1.987 1.221 -0.050
H15 -2.864 1.857 -0.120
H16 -1.024 1.719 0.026

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 C7 H8 C9 H10 H11 C12 H13 C14 H15 H16
C11.09681.09601.09681.52042.22862.53602.76173.95874.17094.59174.86675.90754.79455.78343.9745
H21.09681.77881.77232.16882.59313.25163.52084.58144.72085.36625.25986.32254.95885.84684.0302
H31.09601.77881.77852.16103.11512.66332.43074.18244.64684.67874.99116.04184.83045.78173.9718
H41.09681.77231.77852.16902.59503.24703.51984.58074.64835.08695.68506.67445.76896.78905.0141
C51.52042.16882.16102.16901.09111.34342.10872.53462.65363.27343.55254.53713.74354.79743.1709
H62.22862.59313.11512.59501.09112.10463.07442.75372.41893.57153.76784.64184.07295.08123.6375
C72.53603.25162.66333.24701.34342.10461.09131.51952.15112.15122.54703.51512.92614.00762.5882
H82.76173.52082.43073.51982.10873.07441.09132.22423.10152.52762.94513.89133.12054.12102.6992
C93.95874.58144.18244.58072.53462.75371.51952.22421.10041.09971.53162.22872.54883.53902.8038
H104.17094.72084.64684.64832.65362.41892.15113.10151.10041.76772.15962.63253.17884.10533.4391
H114.59175.36624.67875.08693.27343.57152.15122.52761.09971.76772.15212.48183.30804.20233.6688
C124.86675.25984.99115.68503.55253.76782.54702.94511.53162.15962.15211.08971.34192.12322.1215
H135.90756.32256.04186.67444.53714.64183.51513.89132.22872.63252.48181.08972.10632.45123.0825
C144.79454.95884.83045.76893.74354.07292.92613.12052.54883.17883.30801.34192.10631.08581.0859
H155.78345.84685.78176.78904.79745.08124.00764.12103.53904.10534.20232.12322.45121.08581.8507
H163.97454.03023.97185.01413.17093.63752.58822.69922.80383.43913.66882.12153.08251.08591.8507

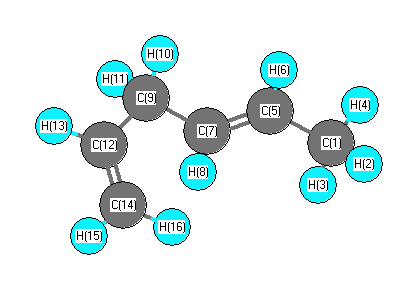
picture of 1,4-Hexadiene, (E)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 116.197 C1 C5 C7 124.517
H2 C1 H3 108.429 H2 C1 H4 107.798
H2 C1 C5 110.892 H3 C1 H4 108.402
H3 C1 C5 110.315 H4 C1 C5 110.910
C5 C7 H8 119.652 C5 C7 C9 124.471
H6 C5 C7 119.285 C7 C9 H10 109.340
C7 C9 H11 109.388 C7 C9 C12 113.185
H8 C7 C9 115.868 C9 C12 H13 115.438
C9 C12 C14 124.868 H10 C9 H11 106.929
H10 C9 C12 109.175 H11 C9 C12 108.630
C12 C14 H15 121.638 C12 C14 H16 121.463
H13 C12 C14 119.685 H15 C14 H16 116.898
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability