Jump to
S1C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -78.982055 |
| Energy at 298.15K | -78.988022 |
| HF Energy | -78.793451 |
| Nuclear repulsion energy | 41.923450 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3091 |
2940 |
0.00 |
164.82 |
0.02 |
0.04 |
| 2 |
A1g |
1499 |
1426 |
0.00 |
8.70 |
0.72 |
0.84 |
| 3 |
A1g |
976 |
928 |
0.00 |
8.89 |
0.28 |
0.43 |
| 4 |
A1u |
323 |
307 |
0.00 |
0.00 |
0.00 |
0.00 |
| 5 |
A2u |
3092 |
2942 |
38.25 |
0.00 |
0.02 |
0.04 |
| 6 |
A2u |
1503 |
1430 |
4.09 |
0.00 |
0.00 |
0.00 |
| 7 |
Eg |
3149 |
2996 |
0.00 |
107.96 |
0.75 |
0.86 |
| 7 |
Eg |
3149 |
2996 |
0.00 |
107.96 |
0.75 |
0.86 |
| 8 |
Eg |
1609 |
1530 |
0.00 |
45.38 |
0.75 |
0.86 |
| 8 |
Eg |
1609 |
1530 |
0.00 |
45.38 |
0.75 |
0.86 |
| 9 |
Eg |
1296 |
1233 |
0.00 |
4.91 |
0.75 |
0.86 |
| 9 |
Eg |
1296 |
1233 |
0.00 |
4.91 |
0.75 |
0.86 |
| 10 |
Eu |
3172 |
3017 |
53.66 |
0.00 |
0.00 |
0.00 |
| 10 |
Eu |
3172 |
3017 |
53.66 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1611 |
1532 |
8.42 |
0.00 |
0.75 |
0.86 |
| 11 |
Eu |
1611 |
1532 |
8.42 |
0.00 |
0.75 |
0.86 |
| 12 |
Eu |
875 |
832 |
6.16 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
875 |
832 |
6.16 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 16952.5 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 16126.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.776 |
| C2 |
0.000 |
0.000 |
-0.776 |
| H3 |
0.000 |
1.024 |
1.163 |
| H4 |
-0.887 |
-0.512 |
1.163 |
| H5 |
0.887 |
-0.512 |
1.163 |
| H6 |
0.000 |
-1.024 |
-1.163 |
| H7 |
-0.887 |
0.512 |
-1.163 |
| H8 |
0.887 |
0.512 |
-1.163 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5515 | 1.0947 | 1.0947 | 1.0947 | 2.1923 | 2.1923 | 2.1923 |
C2 | 1.5515 | | 2.1923 | 2.1923 | 2.1923 | 1.0947 | 1.0947 | 1.0947 | H3 | 1.0947 | 2.1923 | | 1.7736 | 1.7736 | 3.0987 | 2.5409 | 2.5409 | H4 | 1.0947 | 2.1923 | 1.7736 | | 1.7736 | 2.5409 | 2.5409 | 3.0987 | H5 | 1.0947 | 2.1923 | 1.7736 | 1.7736 | | 2.5409 | 3.0987 | 2.5409 | H6 | 2.1923 | 1.0947 | 3.0987 | 2.5409 | 2.5409 | | 1.7736 | 1.7736 | H7 | 2.1923 | 1.0947 | 2.5409 | 2.5409 | 3.0987 | 1.7736 | | 1.7736 | H8 | 2.1923 | 1.0947 | 2.5409 | 3.0987 | 2.5409 | 1.7736 | 1.7736 | |
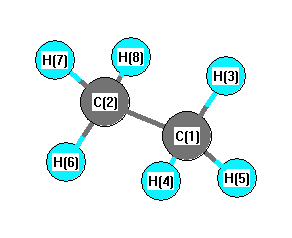 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
110.701 |
|
C1 |
C2 |
H7 |
110.701 |
| C1 |
C2 |
H8 |
110.701 |
|
C2 |
C1 |
H3 |
110.701 |
| C2 |
C1 |
H4 |
110.701 |
|
C2 |
C1 |
H5 |
110.701 |
| H3 |
C1 |
H4 |
108.214 |
|
H3 |
C1 |
H5 |
108.214 |
| H4 |
C1 |
H5 |
108.214 |
|
H6 |
C2 |
H7 |
108.214 |
| H6 |
C2 |
H8 |
108.214 |
|
H7 |
C2 |
H8 |
108.214 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -78.977259 |
| Energy at 298.15K | |
| HF Energy | -78.789042 |
| Nuclear repulsion energy | 41.744729 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3103 |
2952 |
0.00 |
|
|
|
| 2 |
A1' |
1520 |
1446 |
0.00 |
|
|
|
| 3 |
A1' |
962 |
915 |
0.00 |
|
|
|
| 4 |
A1" |
313i |
298i |
0.00 |
|
|
|
| 5 |
A2" |
3097 |
2946 |
40.90 |
|
|
|
| 6 |
A2" |
1499 |
1426 |
3.62 |
|
|
|
| 7 |
E' |
3177 |
3022 |
46.85 |
|
|
|
| 7 |
E' |
3177 |
3022 |
46.85 |
|
|
|
| 8 |
E' |
1618 |
1539 |
10.06 |
|
|
|
| 8 |
E' |
1618 |
1539 |
10.06 |
|
|
|
| 9 |
E' |
958 |
911 |
5.85 |
|
|
|
| 9 |
E' |
958 |
911 |
5.85 |
|
|
|
| 10 |
E" |
3155 |
3002 |
0.00 |
|
|
|
| 10 |
E" |
3155 |
3002 |
0.00 |
|
|
|
| 11 |
E" |
1605 |
1527 |
0.00 |
|
|
|
| 11 |
E" |
1605 |
1527 |
0.00 |
|
|
|
| 12 |
E" |
1238 |
1177 |
0.00 |
|
|
|
| 12 |
E" |
1238 |
1177 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16684.3 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 15871.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.783 |
| C2 |
0.000 |
0.000 |
-0.783 |
| H3 |
0.000 |
1.020 |
1.178 |
| H4 |
-0.884 |
-0.510 |
1.178 |
| H5 |
0.884 |
-0.510 |
1.178 |
| H6 |
0.000 |
1.020 |
-1.178 |
| H7 |
0.884 |
-0.510 |
-1.178 |
| H8 |
-0.884 |
-0.510 |
-1.178 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5664 | 1.0939 | 1.0939 | 1.0939 | 2.2103 | 2.2103 | 2.2103 |
C2 | 1.5664 | | 2.2103 | 2.2103 | 2.2103 | 1.0939 | 1.0939 | 1.0939 | H3 | 1.0939 | 2.2103 | | 1.7673 | 1.7673 | 2.3550 | 2.9444 | 2.9444 | H4 | 1.0939 | 2.2103 | 1.7673 | | 1.7673 | 2.9444 | 2.9444 | 2.3550 | H5 | 1.0939 | 2.2103 | 1.7673 | 1.7673 | | 2.9444 | 2.3550 | 2.9444 | H6 | 2.2103 | 1.0939 | 2.3550 | 2.9444 | 2.9444 | | 1.7673 | 1.7673 | H7 | 2.2103 | 1.0939 | 2.9444 | 2.9444 | 2.3550 | 1.7673 | | 1.7673 | H8 | 2.2103 | 1.0939 | 2.9444 | 2.3550 | 2.9444 | 1.7673 | 1.7673 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.128 |
|
C1 |
C2 |
H7 |
111.128 |
| C1 |
C2 |
H8 |
111.128 |
|
C2 |
C1 |
H3 |
111.128 |
| C2 |
C1 |
H4 |
111.128 |
|
C2 |
C1 |
H5 |
111.128 |
| H3 |
C1 |
H4 |
107.765 |
|
H3 |
C1 |
H5 |
107.765 |
| H4 |
C1 |
H5 |
107.765 |
|
H6 |
C2 |
H7 |
107.765 |
| H6 |
C2 |
H8 |
107.765 |
|
H7 |
C2 |
H8 |
107.765 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
