return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H8 (Propane)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-117.892411
Energy at 298.15K-117.900883
HF Energy-117.612537
Nuclear repulsion energy81.920067
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3155 3001 41.82 66.41 0.65 0.79
2 A1 3089 2938 20.55 69.59 0.23 0.37
3 A1 3080 2930 9.19 170.51 0.00 0.01
4 A1 1612 1533 4.43 18.66 0.74 0.85
5 A1 1593 1516 0.38 36.08 0.75 0.86
6 A1 1502 1429 3.99 3.02 0.63 0.77
7 A1 1231 1172 1.92 5.44 0.61 0.76
8 A1 866 823 0.84 10.93 0.24 0.39
9 A1 383 365 0.21 0.26 0.10 0.18
10 A2 3148 2995 0.00 3.95 0.75 0.86
11 A2 1595 1518 0.00 43.85 0.75 0.86
12 A2 1387 1320 0.00 15.59 0.75 0.86
13 A2 974 927 0.00 0.04 0.75 0.86
14 A2 238 226 0.00 0.00 0.75 0.86
15 B1 3161 3007 77.96 18.58 0.75 0.86
16 B1 3121 2969 1.07 127.76 0.75 0.86
17 B1 1611 1532 12.34 0.16 0.75 0.86
18 B1 1287 1224 0.44 1.77 0.75 0.86
19 B1 785 746 4.69 0.14 0.75 0.86
20 B1 284 270 0.00 0.02 0.75 0.86
21 B2 3152 2998 22.06 42.94 0.75 0.86
22 B2 3079 2929 25.12 4.23 0.75 0.86
23 B2 1601 1523 2.46 0.07 0.75 0.86
24 B2 1485 1413 3.74 5.00 0.75 0.86
25 B2 1429 1359 2.32 2.58 0.75 0.86
26 B2 1057 1005 2.03 4.70 0.75 0.86
27 B2 976 928 2.81 1.35 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 23439.6 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 22298.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.95918 0.27779 0.24518

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.603
C2 0.000 1.282 -0.267
C3 0.000 -1.282 -0.267
H4 0.883 0.000 1.253
H5 -0.883 0.000 1.253
H6 0.000 2.183 0.357
H7 0.000 -2.183 0.357
H8 0.888 1.305 -0.909
H9 -0.888 1.305 -0.909
H10 -0.888 -1.305 -0.909
H11 0.888 -1.305 -0.909

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 H6 H7 H8 H9 H10 H11
C11.54971.54971.09661.09662.19692.19692.18572.18572.18572.1857
C21.54972.56462.17612.17611.09553.52091.09581.09582.80962.8096
C31.54972.56462.17612.17613.52091.09552.80962.80961.09581.0958
H41.09662.17612.17611.76652.51982.51982.52523.08443.08442.5252
H51.09662.17612.17611.76652.51982.51983.08442.52522.52523.0844
H62.19691.09553.52092.51982.51984.36601.77781.77783.81513.8151
H72.19693.52091.09552.51982.51984.36603.81513.81511.77781.7778
H82.18571.09582.80962.52523.08441.77783.81511.77603.15682.6098
H92.18571.09582.80963.08442.52521.77783.81511.77602.60983.1568
H102.18572.80961.09583.08442.52523.81511.77783.15682.60981.7760
H112.18572.80961.09582.52523.08443.81511.77782.60983.15681.7760

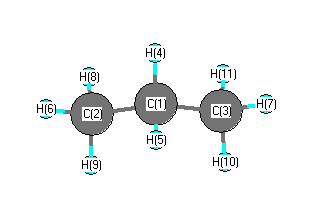
picture of Propane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H6 111.141 C1 C2 H8 110.237
C1 C2 H9 110.237 C1 C3 H7 111.141
C1 C3 H10 110.237 C1 C3 H11 110.237
C2 C1 C3 111.676 C2 C1 H4 109.440
C2 C1 H5 109.440 C3 C1 H4 109.440
C3 C1 H5 109.440 H4 C1 H5 107.304
H6 C2 H8 108.444 H6 C2 H9 108.444
H7 C3 H10 108.444 H7 C3 H11 108.444
H8 C2 H9 108.258 H10 C3 H11 108.258
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability