Jump to
S1C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -474.763672 |
| Energy at 298.15K | -474.770017 |
| HF Energy | -474.456338 |
| Nuclear repulsion energy | 106.773267 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3168 |
3013 |
17.71 |
|
|
|
| 2 |
A' |
3115 |
2963 |
16.37 |
|
|
|
| 3 |
A' |
3093 |
2942 |
12.43 |
|
|
|
| 4 |
A' |
2782 |
2647 |
10.66 |
|
|
|
| 5 |
A' |
1605 |
1527 |
3.54 |
|
|
|
| 6 |
A' |
1588 |
1510 |
5.56 |
|
|
|
| 7 |
A' |
1500 |
1427 |
6.22 |
|
|
|
| 8 |
A' |
1384 |
1316 |
30.73 |
|
|
|
| 9 |
A' |
1154 |
1098 |
2.88 |
|
|
|
| 10 |
A' |
1006 |
957 |
4.16 |
|
|
|
| 11 |
A' |
889 |
846 |
1.88 |
|
|
|
| 12 |
A' |
684 |
651 |
1.01 |
|
|
|
| 13 |
A' |
322 |
306 |
2.83 |
|
|
|
| 14 |
A" |
3181 |
3026 |
24.41 |
|
|
|
| 15 |
A" |
3159 |
3005 |
0.04 |
|
|
|
| 16 |
A" |
1597 |
1519 |
8.24 |
|
|
|
| 17 |
A" |
1347 |
1282 |
0.63 |
|
|
|
| 18 |
A" |
1119 |
1064 |
0.12 |
|
|
|
| 19 |
A" |
838 |
797 |
4.93 |
|
|
|
| 20 |
A" |
264 |
251 |
1.74 |
|
|
|
| 21 |
A" |
194 |
184 |
23.80 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16992.3 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 16164.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.540 |
0.674 |
0.000 |
| C2 |
0.000 |
0.837 |
0.000 |
| S3 |
-0.769 |
-0.833 |
0.000 |
| H4 |
2.016 |
1.659 |
0.000 |
| H5 |
1.866 |
0.126 |
0.889 |
| H6 |
1.866 |
0.126 |
-0.889 |
| H7 |
-0.317 |
1.386 |
0.890 |
| H8 |
-0.317 |
1.386 |
-0.890 |
| H9 |
-2.044 |
-0.423 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5486 | 2.7572 | 1.0948 | 1.0942 | 1.0942 | 2.1792 | 2.1792 | 3.7480 |
C2 | 1.5486 | | 1.8384 | 2.1776 | 2.1861 | 2.1861 | 1.0930 | 1.0930 | 2.4013 | S3 | 2.7572 | 1.8384 | | 3.7377 | 2.9420 | 2.9420 | 2.4332 | 2.4332 | 1.3387 | H4 | 1.0948 | 2.1776 | 3.7377 | | 1.7791 | 1.7791 | 2.5123 | 2.5123 | 4.5632 | H5 | 1.0942 | 2.1861 | 2.9420 | 1.7791 | | 1.7788 | 2.5208 | 3.0857 | 4.0473 | H6 | 1.0942 | 2.1861 | 2.9420 | 1.7791 | 1.7788 | | 3.0857 | 2.5208 | 4.0473 | H7 | 2.1792 | 1.0930 | 2.4332 | 2.5123 | 2.5208 | 3.0857 | | 1.7805 | 2.6550 | H8 | 2.1792 | 1.0930 | 2.4332 | 2.5123 | 3.0857 | 2.5208 | 1.7805 | | 2.6550 | H9 | 3.7480 | 2.4013 | 1.3387 | 4.5632 | 4.0473 | 4.0473 | 2.6550 | 2.6550 | |
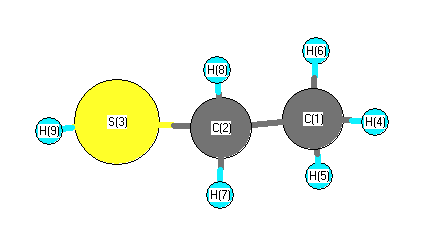 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
108.681 |
|
C1 |
C2 |
H7 |
109.967 |
| C1 |
C2 |
H8 |
109.967 |
|
C2 |
C1 |
H4 |
109.730 |
| C2 |
C1 |
H5 |
110.438 |
|
C2 |
C1 |
H6 |
110.438 |
| C2 |
S3 |
H9 |
96.932 |
|
S3 |
C2 |
H7 |
109.569 |
| S3 |
C2 |
H8 |
109.569 |
|
H4 |
C1 |
H5 |
108.726 |
| H4 |
C1 |
H6 |
108.726 |
|
H5 |
C1 |
H6 |
108.739 |
| H7 |
C2 |
H8 |
109.076 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -474.764567 |
| Energy at 298.15K | -474.770995 |
| HF Energy | -474.456832 |
| Nuclear repulsion energy | 106.566662 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3181 |
3026 |
21.24 |
|
|
|
| 2 |
A |
3161 |
3007 |
9.82 |
|
|
|
| 3 |
A |
3152 |
2998 |
13.41 |
|
|
|
| 4 |
A |
3114 |
2962 |
10.79 |
|
|
|
| 5 |
A |
3086 |
2936 |
15.46 |
|
|
|
| 6 |
A |
2780 |
2645 |
9.38 |
|
|
|
| 7 |
A |
1601 |
1523 |
3.13 |
|
|
|
| 8 |
A |
1595 |
1517 |
9.48 |
|
|
|
| 9 |
A |
1576 |
1499 |
3.82 |
|
|
|
| 10 |
A |
1498 |
1425 |
5.07 |
|
|
|
| 11 |
A |
1385 |
1317 |
14.97 |
|
|
|
| 12 |
A |
1357 |
1290 |
2.99 |
|
|
|
| 13 |
A |
1187 |
1130 |
10.14 |
|
|
|
| 14 |
A |
1113 |
1059 |
2.23 |
|
|
|
| 15 |
A |
984 |
936 |
4.46 |
|
|
|
| 16 |
A |
929 |
884 |
10.59 |
|
|
|
| 17 |
A |
778 |
741 |
2.59 |
|
|
|
| 18 |
A |
671 |
639 |
3.78 |
|
|
|
| 19 |
A |
345 |
328 |
1.74 |
|
|
|
| 20 |
A |
274 |
261 |
3.91 |
|
|
|
| 21 |
A |
236 |
225 |
21.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17001.9 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 16173.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.647 |
-0.359 |
-0.052 |
| C2 |
0.494 |
0.661 |
0.088 |
| S3 |
-1.168 |
-0.104 |
-0.079 |
| H4 |
2.613 |
0.154 |
0.011 |
| H5 |
1.598 |
-1.106 |
0.747 |
| H6 |
1.579 |
-0.875 |
-1.014 |
| H7 |
0.556 |
1.191 |
1.042 |
| H8 |
0.547 |
1.404 |
-0.713 |
| H9 |
-1.057 |
-0.918 |
0.978 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5457 | 2.8266 | 1.0961 | 1.0951 | 1.0934 | 2.1888 | 2.1808 | 2.9468 |
C2 | 1.5457 | | 1.8371 | 2.1803 | 2.1850 | 2.1798 | 1.0933 | 1.0944 | 2.3850 | S3 | 2.8266 | 1.8371 | | 3.7912 | 3.0557 | 3.0025 | 2.4304 | 2.3695 | 1.3388 | H4 | 1.0961 | 2.1803 | 3.7912 | | 1.7777 | 1.7834 | 2.5239 | 2.5216 | 3.9437 | H5 | 1.0951 | 2.1850 | 3.0557 | 1.7777 | | 1.7759 | 2.5395 | 3.0883 | 2.6714 | H6 | 1.0934 | 2.1798 | 3.0025 | 1.7834 | 1.7759 | | 3.0893 | 2.5200 | 3.3041 | H7 | 2.1888 | 1.0933 | 2.4304 | 2.5239 | 2.5395 | 3.0893 | | 1.7687 | 2.6556 | H8 | 2.1808 | 1.0944 | 2.3695 | 2.5216 | 3.0883 | 2.5200 | 1.7687 | | 3.2895 | H9 | 2.9468 | 2.3850 | 1.3388 | 3.9437 | 2.6714 | 3.3041 | 2.6556 | 3.2895 | |
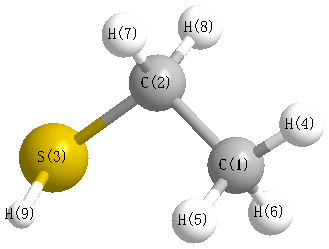 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
113.075 |
|
C1 |
C2 |
H7 |
110.916 |
| C1 |
C2 |
H8 |
110.216 |
|
C2 |
C1 |
H4 |
110.083 |
| C2 |
C1 |
H5 |
110.505 |
|
C2 |
C1 |
H6 |
110.191 |
| C2 |
S3 |
H9 |
96.080 |
|
S3 |
C2 |
H7 |
109.436 |
| S3 |
C2 |
H8 |
105.020 |
|
H4 |
C1 |
H5 |
108.452 |
| H4 |
C1 |
H6 |
109.081 |
|
H5 |
C1 |
H6 |
108.483 |
| H7 |
C2 |
H8 |
107.901 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
