Jump to
S1C2
Energy calculated at MP2=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -168.309388 |
| Energy at 298.15K | -168.313328 |
| HF Energy | -167.981488 |
| Nuclear repulsion energy | 69.941365 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3703 |
3510 |
33.12 |
|
|
|
| 2 |
A' |
3563 |
3378 |
40.38 |
|
|
|
| 3 |
A' |
3000 |
2844 |
115.64 |
|
|
|
| 4 |
A' |
1738 |
1647 |
237.19 |
|
|
|
| 5 |
A' |
1697 |
1609 |
19.55 |
|
|
|
| 6 |
A' |
1476 |
1399 |
10.01 |
|
|
|
| 7 |
A' |
1272 |
1206 |
108.92 |
|
|
|
| 8 |
A' |
1062 |
1007 |
11.02 |
|
|
|
| 9 |
A' |
572 |
542 |
8.94 |
|
|
|
| 10 |
A" |
1089 |
1032 |
0.07 |
|
|
|
| 11 |
A" |
650 |
616 |
98.80 |
|
|
|
| 12 |
A" |
426 |
404 |
300.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10123.7 cm
-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 9597.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.423 |
0.000 |
| O2 |
1.232 |
0.225 |
0.000 |
| N3 |
-0.967 |
-0.558 |
0.000 |
| H4 |
-0.451 |
1.428 |
0.000 |
| H5 |
-0.686 |
-1.531 |
0.000 |
| H6 |
-1.952 |
-0.331 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2479 | 1.3774 | 1.1015 | 2.0712 | 2.0923 |
O2 | 1.2479 | | 2.3344 | 2.0688 | 2.6010 | 3.2320 | N3 | 1.3774 | 2.3344 | | 2.0516 | 1.0128 | 1.0105 | H4 | 1.1015 | 2.0688 | 2.0516 | | 2.9683 | 2.3120 | H5 | 2.0712 | 2.6010 | 1.0128 | 2.9683 | | 1.7438 | H6 | 2.0923 | 3.2320 | 1.0105 | 2.3120 | 1.7438 | |
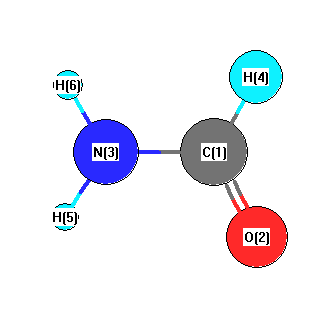 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.329 |
|
C1 |
N3 |
H6 |
121.624 |
| O2 |
C1 |
N3 |
125.470 |
|
O2 |
C1 |
H4 |
123.302 |
| N3 |
C1 |
H4 |
111.227 |
|
H5 |
N3 |
H6 |
119.048 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -168.309388 |
| Energy at 298.15K | -168.313328 |
| HF Energy | -167.981485 |
| Nuclear repulsion energy | 69.944098 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3703 |
3510 |
33.22 |
|
|
|
| 2 |
A |
3563 |
3377 |
40.35 |
|
|
|
| 3 |
A |
3000 |
2844 |
115.63 |
|
|
|
| 4 |
A |
1738 |
1647 |
236.96 |
|
|
|
| 5 |
A |
1697 |
1609 |
19.76 |
|
|
|
| 6 |
A |
1475 |
1399 |
10.09 |
|
|
|
| 7 |
A |
1273 |
1206 |
108.75 |
|
|
|
| 8 |
A |
1089 |
1032 |
0.07 |
|
|
|
| 9 |
A |
1063 |
1007 |
10.89 |
|
|
|
| 10 |
A |
651 |
617 |
99.13 |
|
|
|
| 11 |
A |
572 |
542 |
8.97 |
|
|
|
| 12 |
A |
426 |
404 |
299.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10124.0 cm
-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 9597.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/3-21G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.155 |
0.393 |
-0.000 |
| O2 |
1.229 |
-0.244 |
0.000 |
| N3 |
-1.104 |
-0.163 |
-0.000 |
| H4 |
0.106 |
1.494 |
-0.000 |
| H5 |
-1.201 |
-1.171 |
0.000 |
| H6 |
-1.937 |
0.410 |
0.001 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2480 | 1.3771 | 1.1015 | 2.0706 | 2.0922 |
O2 | 1.2480 | | 2.3341 | 2.0689 | 2.6001 | 3.2320 | N3 | 1.3771 | 2.3341 | | 2.0516 | 1.0129 | 1.0105 | H4 | 1.1015 | 2.0689 | 2.0516 | | 2.9681 | 2.3124 | H5 | 2.0706 | 2.6001 | 1.0129 | 2.9681 | | 1.7440 | H6 | 2.0922 | 3.2320 | 1.0105 | 2.3124 | 1.7440 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.289 |
|
C1 |
N3 |
H6 |
121.639 |
| O2 |
C1 |
N3 |
125.461 |
|
O2 |
C1 |
H4 |
123.292 |
| N3 |
C1 |
H4 |
111.247 |
|
H5 |
N3 |
H6 |
119.072 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
