return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHBrCH3 (i-propyl bromide)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-2677.718252
Energy at 298.15K-2677.728852
HF Energy-2677.307411
Nuclear repulsion energy249.860365
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3185 3030 19.70      
2 A' 3163 3009 22.04      
3 A' 3145 2992 0.49      
4 A' 3084 2934 17.34      
5 A' 1604 1526 9.21      
6 A' 1594 1516 8.26      
7 A' 1503 1430 5.82      
8 A' 1327 1263 45.18      
9 A' 1243 1182 21.80      
10 A' 1116 1062 6.24      
11 A' 885 841 5.77      
12 A' 583 555 11.78      
13 A' 416 395 0.95      
14 A' 308 293 1.30      
15 A' 280 266 0.75      
16 A" 3179 3024 8.13      
17 A" 3156 3002 2.34      
18 A" 3081 2931 7.49      
19 A" 1587 1510 0.90      
20 A" 1583 1506 3.36      
21 A" 1486 1414 10.93      
22 A" 1417 1348 1.38      
23 A" 1159 1102 1.24      
24 A" 1009 960 2.47      
25 A" 986 938 0.14      
26 A" 295 280 0.64      
27 A" 270 257 0.04      

Unscaled Zero Point Vibrational Energy (zpe) 21321.1 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 20282.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.26330 0.09633 0.07576

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.471 -0.901 0.000
Br2 -0.068 1.009 0.000
H3 1.563 -0.880 0.000
C4 -0.068 -1.558 1.282
C5 -0.068 -1.558 -1.282
H6 -1.161 -1.509 1.291
H7 0.243 -2.610 1.308
H8 0.317 -1.049 2.170
H9 -1.161 -1.509 -1.291
H10 0.243 -2.610 -1.308
H11 0.317 -1.049 -2.170

Atom - Atom Distances (Å)
  C1 Br2 H3 C4 C5 H6 H7 H8 H9 H10 H11
C11.98481.09221.53831.53832.16782.16412.18062.16782.16412.1806
Br21.98482.49552.86972.86973.03343.86083.01583.03343.86083.0158
H31.09222.49552.18272.18273.07942.53912.50813.07942.53912.5081
C41.53832.86972.18272.56441.09411.09721.09332.79642.81303.5108
C51.53832.86972.18272.56442.79642.81303.51081.09411.09721.0933
H62.16783.03343.07941.09412.79641.78451.77972.58263.15303.7916
H72.16413.86082.53911.09722.81301.78451.78433.15302.61633.8130
H82.18063.01582.50811.09333.51081.77971.78433.79163.81304.3402
H92.16783.03343.07942.79641.09412.58263.15303.79161.78451.7797
H102.16413.86082.53912.81301.09723.15302.61633.81301.78451.7843
H112.18063.01582.50813.51081.09333.79163.81304.34021.77971.7843

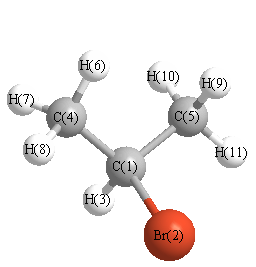
picture of i-propyl bromide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H6 109.730 C1 C4 H7 109.257
C1 C4 H8 110.782 C1 C5 H9 109.730
C1 C5 H10 109.257 C1 C5 H11 110.782
Br2 C1 H3 104.629 Br2 C1 C4 108.420
Br2 C1 C5 108.420 H3 C1 C4 111.018
H3 C1 C5 111.018 C4 C1 C5 112.931
H6 C4 H7 109.054 H6 C4 H8 108.898
H7 C4 H8 109.093 H9 C5 H10 109.054
H9 C5 H11 108.898 H10 C5 H11 109.093
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability