return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHClCH3 (Propane, 2-chloro-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-574.851696
Energy at 298.15K-574.859515
HF Energy-574.440156
Nuclear repulsion energy164.590882
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3187 3031 23.09 34.83 0.46 0.63
2 A' 3168 3014 21.51 78.88 0.74 0.85
3 A' 3149 2996 0.31 118.14 0.53 0.69
4 A' 3089 2939 12.00 170.05 0.02 0.04
5 A' 1605 1527 10.63 2.45 0.68 0.81
6 A' 1594 1516 7.35 32.73 0.75 0.86
7 A' 1504 1431 6.98 3.25 0.51 0.67
8 A' 1358 1292 32.91 7.26 0.72 0.84
9 A' 1247 1186 15.01 1.85 0.73 0.85
10 A' 1144 1089 13.01 6.65 0.58 0.73
11 A' 888 845 9.37 9.15 0.63 0.77
12 A' 627 596 25.24 16.90 0.28 0.43
13 A' 433 412 1.71 1.52 0.17 0.28
14 A' 349 332 1.79 1.27 0.52 0.69
15 A' 284 270 0.12 0.03 0.62 0.77
16 A" 3180 3025 8.60 29.75 0.75 0.86
17 A" 3161 3007 2.67 12.96 0.75 0.86
18 A" 3086 2936 7.65 1.58 0.75 0.86
19 A" 1588 1511 0.97 28.27 0.75 0.86
20 A" 1582 1505 3.18 9.79 0.75 0.86
21 A" 1488 1416 11.03 6.04 0.75 0.86
22 A" 1420 1351 1.71 5.73 0.75 0.86
23 A" 1165 1108 3.52 4.10 0.75 0.86
24 A" 1012 963 2.32 0.38 0.75 0.86
25 A" 985 937 0.21 2.96 0.75 0.86
26 A" 334 318 1.53 1.57 0.75 0.86
27 A" 264 251 0.03 0.05 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 21445.8 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 20401.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.26510 0.14886 0.10490

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Cl1 -0.910 1.014 0.000
C2 0.584 -0.052 0.000
C3 0.584 -0.905 1.279
C4 0.584 -0.905 -1.279
H5 1.427 0.641 0.000
H6 1.491 -1.521 1.302
H7 1.491 -1.521 -1.302
H8 0.562 -0.269 2.169
H9 0.562 -0.269 -2.169
H10 -0.293 -1.559 1.288
H11 -0.293 -1.559 -1.288

Atom - Atom Distances (Å)
  Cl1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11
Cl11.83512.74762.74762.36643.72573.72572.91842.91842.94242.9424
C21.83511.53751.53751.09162.16182.16182.17992.17992.16762.1676
C32.74761.53752.55882.17671.09642.80411.09343.50641.09412.7910
C42.74761.53752.55882.17672.80411.09643.50641.09342.79101.0941
H52.36641.09162.17672.17672.52452.52452.50632.50633.07553.0755
H63.72572.16181.09642.80412.52452.60321.78343.80431.78423.1450
H73.72572.16182.80411.09642.52452.60323.80431.78343.14501.7842
H82.91842.17991.09343.50642.50631.78343.80434.33781.78043.7876
H92.91842.17993.50641.09342.50633.80431.78344.33783.78761.7804
H102.94242.16761.09412.79103.07551.78423.14501.78043.78762.5765
H112.94242.16762.79101.09413.07553.14501.78423.78761.78042.5765

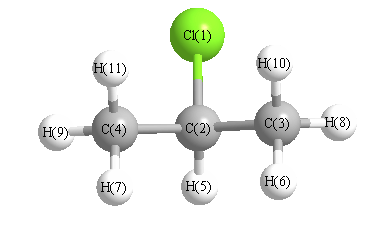
picture of Propane, 2-chloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Cl1 C2 C3 108.793 Cl1 C2 C4 108.793
Cl1 C3 H5 56.006 C2 C3 H6 109.177
C2 C3 H8 110.767 C2 C3 H10 109.766
C2 C4 H7 109.177 C2 C4 H9 110.767
C2 C4 H11 109.766 C3 C2 C4 112.633
C3 C2 H5 110.626 C4 C2 H5 110.626
H6 C3 H8 109.061 H6 C3 H10 109.088
H7 C4 H9 109.061 H7 C4 H11 109.088
H8 C3 H10 108.954 H9 C4 H11 108.954
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability