Jump to
S1C2
Vibrational Frequencies calculated at MP2/3-21G*
Geometric Data calculated at MP2/3-21G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -231.362519 |
| Energy at 298.15K | -231.373832 |
| HF Energy | -230.871291 |
| Nuclear repulsion energy | 197.357092 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3480 |
3311 |
2.48 |
|
|
|
| 2 |
A |
3177 |
3023 |
42.07 |
|
|
|
| 3 |
A |
3176 |
3021 |
27.92 |
|
|
|
| 4 |
A |
3170 |
3016 |
0.80 |
|
|
|
| 5 |
A |
3170 |
3016 |
1.41 |
|
|
|
| 6 |
A |
3149 |
2996 |
41.63 |
|
|
|
| 7 |
A |
3143 |
2990 |
3.41 |
|
|
|
| 8 |
A |
3092 |
2942 |
7.27 |
|
|
|
| 9 |
A |
3076 |
2926 |
22.78 |
|
|
|
| 10 |
A |
3072 |
2922 |
10.28 |
|
|
|
| 11 |
A |
1615 |
1537 |
11.39 |
|
|
|
| 12 |
A |
1602 |
1524 |
7.43 |
|
|
|
| 13 |
A |
1599 |
1521 |
3.50 |
|
|
|
| 14 |
A |
1589 |
1511 |
0.19 |
|
|
|
| 15 |
A |
1586 |
1509 |
0.01 |
|
|
|
| 16 |
A |
1578 |
1501 |
0.05 |
|
|
|
| 17 |
A |
1500 |
1427 |
2.54 |
|
|
|
| 18 |
A |
1474 |
1402 |
15.39 |
|
|
|
| 19 |
A |
1474 |
1402 |
12.76 |
|
|
|
| 20 |
A |
1421 |
1352 |
24.72 |
|
|
|
| 21 |
A |
1309 |
1245 |
27.74 |
|
|
|
| 22 |
A |
1299 |
1236 |
30.12 |
|
|
|
| 23 |
A |
1201 |
1142 |
52.95 |
|
|
|
| 24 |
A |
1102 |
1049 |
1.81 |
|
|
|
| 25 |
A |
1082 |
1030 |
13.12 |
|
|
|
| 26 |
A |
1021 |
972 |
0.01 |
|
|
|
| 27 |
A |
953 |
907 |
0.52 |
|
|
|
| 28 |
A |
940 |
894 |
0.45 |
|
|
|
| 29 |
A |
932 |
886 |
30.49 |
|
|
|
| 30 |
A |
730 |
695 |
2.19 |
|
|
|
| 31 |
A |
448 |
426 |
10.28 |
|
|
|
| 32 |
A |
444 |
423 |
9.85 |
|
|
|
| 33 |
A |
403 |
383 |
0.60 |
|
|
|
| 34 |
A |
352 |
335 |
116.03 |
|
|
|
| 35 |
A |
346 |
329 |
2.90 |
|
|
|
| 36 |
A |
340 |
324 |
14.28 |
|
|
|
| 37 |
A |
282 |
268 |
0.05 |
|
|
|
| 38 |
A |
274 |
260 |
0.34 |
|
|
|
| 39 |
A |
231 |
220 |
1.99 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30415.8 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 28934.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.001 |
0.000 |
-0.003 |
| H2 |
-1.767 |
-1.266 |
-0.145 |
| H3 |
-0.221 |
-2.151 |
-0.070 |
| H4 |
-0.715 |
-1.329 |
-1.579 |
| C5 |
-0.723 |
-1.272 |
-0.486 |
| H6 |
1.968 |
-0.890 |
-0.029 |
| H7 |
1.968 |
0.888 |
-0.029 |
| H8 |
1.555 |
-0.001 |
-1.526 |
| C9 |
1.475 |
-0.001 |
-0.434 |
| H10 |
-0.220 |
2.151 |
-0.070 |
| H11 |
-1.766 |
1.268 |
-0.145 |
| H12 |
-0.714 |
1.330 |
-1.579 |
| C13 |
-0.722 |
1.273 |
-0.486 |
| H14 |
-0.865 |
0.000 |
1.805 |
| O15 |
0.074 |
0.000 |
1.478 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
H4 |
C5 |
H6 |
H7 |
H8 |
C9 |
H10 |
H11 |
H12 |
C13 |
H14 |
O15 |
| C1 | | 2.1794 | 2.1636 | 2.1832 | 1.5418 | 2.1585 | 2.1584 | 2.1760 | 1.5359 | 2.1636 | 2.1794 | 2.1832 | 1.5418 | 2.0044 | 1.4820 |
H2 | 2.1794 | | 1.7823 | 1.7796 | 1.0977 | 3.7552 | 4.3135 | 3.8139 | 3.4924 | 3.7522 | 2.5341 | 3.1475 | 2.7666 | 2.4934 | 2.7613 | H3 | 2.1636 | 1.7823 | | 1.7878 | 1.0943 | 2.5267 | 3.7460 | 3.1463 | 2.7632 | 4.3022 | 3.7522 | 3.8259 | 3.4851 | 2.9255 | 2.6664 | H4 | 2.1832 | 1.7796 | 1.7878 | | 1.0948 | 3.1298 | 3.8109 | 2.6315 | 2.8064 | 3.8259 | 3.1475 | 2.6594 | 2.8225 | 3.6389 | 3.4256 | C5 | 1.5418 | 1.0977 | 1.0943 | 1.0948 | | 2.7563 | 3.4816 | 2.8091 | 2.5405 | 3.4851 | 2.7666 | 2.8225 | 2.5449 | 2.6242 | 2.4719 | H6 | 2.1585 | 3.7552 | 2.5267 | 3.1298 | 2.7563 | | 1.7777 | 1.7891 | 1.0941 | 3.7460 | 4.3135 | 3.8109 | 3.4815 | 3.4902 | 2.5781 | H7 | 2.1584 | 4.3135 | 3.7460 | 3.8109 | 3.4816 | 1.7777 | | 1.7891 | 1.0941 | 2.5267 | 3.7552 | 3.1298 | 2.7563 | 3.4901 | 2.5781 | H8 | 2.1760 | 3.8139 | 3.1463 | 2.6315 | 2.8091 | 1.7891 | 1.7891 | | 1.0945 | 3.1463 | 3.8139 | 2.6315 | 2.8090 | 4.1173 | 3.3487 | C9 | 1.5359 | 3.4924 | 2.7632 | 2.8064 | 2.5405 | 1.0941 | 1.0941 | 1.0945 | | 2.7632 | 3.4924 | 2.8064 | 2.5405 | 3.2391 | 2.3703 | H10 | 2.1636 | 3.7522 | 4.3022 | 3.8259 | 3.4851 | 3.7460 | 2.5267 | 3.1463 | 2.7632 | | 1.7823 | 1.7878 | 1.0943 | 2.9255 | 2.6664 | H11 | 2.1794 | 2.5341 | 3.7522 | 3.1475 | 2.7666 | 4.3135 | 3.7552 | 3.8139 | 3.4924 | 1.7823 | | 1.7796 | 1.0977 | 2.4934 | 2.7613 | H12 | 2.1832 | 3.1475 | 3.8259 | 2.6594 | 2.8225 | 3.8109 | 3.1298 | 2.6315 | 2.8064 | 1.7878 | 1.7796 | | 1.0948 | 3.6389 | 3.4256 | C13 | 1.5418 | 2.7666 | 3.4851 | 2.8225 | 2.5449 | 3.4815 | 2.7563 | 2.8090 | 2.5405 | 1.0943 | 1.0977 | 1.0948 | | 2.6242 | 2.4718 | H14 | 2.0044 | 2.4934 | 2.9255 | 3.6389 | 2.6242 | 3.4902 | 3.4901 | 4.1173 | 3.2391 | 2.9255 | 2.4934 | 3.6389 | 2.6242 | | 0.9950 | O15 | 1.4820 | 2.7613 | 2.6664 | 3.4256 | 2.4719 | 2.5781 | 2.5781 | 3.3487 | 2.3703 | 2.6664 | 2.7613 | 3.4256 | 2.4718 | 0.9950 | |
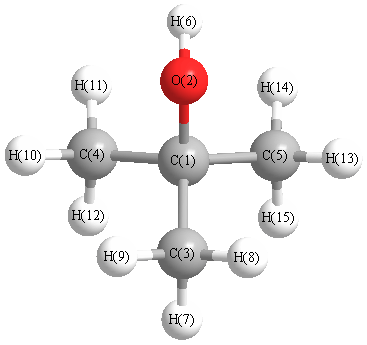 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C5 |
H2 |
110.177 |
|
C1 |
C5 |
H3 |
109.142 |
| C1 |
C5 |
H4 |
110.654 |
|
C1 |
C9 |
H6 |
109.156 |
| C1 |
C9 |
H7 |
109.156 |
|
C1 |
C9 |
H8 |
110.512 |
| C1 |
C13 |
H10 |
109.142 |
|
C1 |
C13 |
H11 |
110.181 |
| C1 |
C13 |
H12 |
110.656 |
|
C1 |
O15 |
H14 |
106.370 |
| H2 |
C5 |
H3 |
108.804 |
|
H2 |
C5 |
H4 |
108.523 |
| H3 |
C5 |
H4 |
109.512 |
|
C5 |
C1 |
C9 |
111.266 |
| C5 |
C1 |
C13 |
111.237 |
|
C5 |
C1 |
O15 |
109.646 |
| H6 |
C9 |
H7 |
108.663 |
|
H6 |
C9 |
H8 |
109.661 |
| H7 |
C9 |
H8 |
109.663 |
|
C9 |
C1 |
C13 |
111.266 |
| C9 |
C1 |
O15 |
103.498 |
|
H10 |
C13 |
H11 |
108.800 |
| H10 |
C13 |
H12 |
109.510 |
|
H11 |
C13 |
H12 |
108.523 |
| C13 |
C1 |
O15 |
109.647 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
