return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(CH3)3SH (2-Propanethiol, 2-methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-552.596221
Energy at 298.15K-552.607246
HF Energy-552.099639
Nuclear repulsion energy243.395434
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3166 3012 20.85      
2 A' 3154 3000 20.31      
3 A' 3143 2990 33.72      
4 A' 3077 2927 11.94      
5 A' 3071 2921 21.76      
6 A' 2769 2635 11.68      
7 A' 1609 1531 9.91      
8 A' 1597 1519 10.98      
9 A' 1586 1509 0.43      
10 A' 1507 1434 1.56      
11 A' 1481 1409 14.12      
12 A' 1295 1231 0.86      
13 A' 1268 1206 37.21      
14 A' 1119 1064 1.53      
15 A' 977 929 4.08      
16 A' 909 865 3.05      
17 A' 826 786 1.63      
18 A' 598 568 3.78      
19 A' 406 386 1.03      
20 A' 378 360 0.25      
21 A' 314 299 0.30      
22 A' 297 283 1.04      
23 A" 3164 3010 20.44      
24 A" 3159 3005 3.92      
25 A" 3138 2985 0.36      
26 A" 3068 2918 11.37      
27 A" 1598 1521 9.09      
28 A" 1584 1507 0.44      
29 A" 1578 1501 0.02      
30 A" 1482 1410 11.10      
31 A" 1282 1220 4.58      
32 A" 1105 1051 0.01      
33 A" 1035 984 0.02      
34 A" 958 911 0.63      
35 A" 412 392 0.47      
36 A" 318 303 2.58      
37 A" 297 283 0.08      
38 A" 277 264 7.27      
39 A" 240 228 15.03      

Unscaled Zero Point Vibrational Energy (zpe) 29619.3 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 28176.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.14748 0.09856 0.09770

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.341 -0.006 0.000
S2 -1.508 0.085 0.000
C3 0.839 1.461 0.000
C4 0.839 -0.734 1.270
C5 0.839 -0.734 -1.270
H6 -1.713 -1.239 0.000
H7 1.935 1.465 0.000
H8 0.482 1.988 -0.892
H9 0.482 1.988 0.892
H10 1.936 -0.747 1.277
H11 1.936 -0.747 -1.277
H12 0.479 -0.221 2.167
H13 0.478 -1.769 1.287
H14 0.479 -0.221 -2.167
H15 0.478 -1.769 -1.287

Atom - Atom Distances (Å)
  C1 S2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.85111.54951.54601.54602.39572.16932.18912.18912.17322.17322.18252.18722.18252.1872
S21.85112.72032.79112.79111.34013.70922.89422.89423.76603.76602.95633.00652.95633.0065
C31.54952.72032.53612.53613.71531.09641.09561.09562.77632.77632.76723.49602.76723.4960
C41.54602.79112.53612.53972.89482.76603.49432.77111.09762.77301.09451.09643.49392.7821
C51.54602.79112.53612.53972.89482.76602.77113.49432.77301.09763.49392.78211.09451.0964
H62.39571.34013.71532.89482.89484.54124.00364.00363.89753.89753.24662.59593.24662.5959
H72.16933.70921.09642.76602.76604.54121.78321.78322.55362.55363.10833.77373.10833.7737
H82.18912.89421.09563.49432.77114.00361.78321.78413.78073.12073.77374.34332.55083.7778
H92.18912.89421.09562.77113.49434.00361.78321.78413.12073.78072.55083.77783.77374.3433
H102.17323.76602.77631.09762.77303.89752.55363.78073.12072.55341.78671.78123.77643.1218
H112.17323.76602.77632.77301.09763.89752.55363.12073.78072.55343.77643.12181.78671.7812
H122.18252.95632.76721.09453.49393.24663.10833.77372.55081.78673.77641.78084.33493.7855
H132.18723.00653.49601.09642.78212.59593.77374.34333.77781.78123.12181.78083.78552.5742
H142.18252.95632.76723.49391.09453.24663.10832.55083.77373.77641.78674.33493.78551.7808
H152.18723.00653.49602.78211.09642.59593.77373.77784.34333.12181.78123.78552.57421.7808

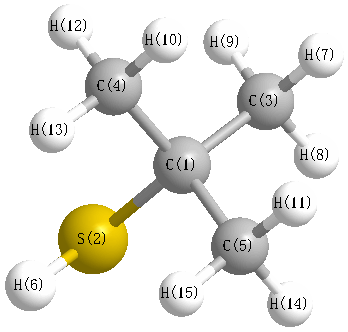
picture of 2-Propanethiol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 H6 95.984 C1 C3 H7 108.944
C1 C3 H8 110.533 C1 C3 H9 110.533
C1 C4 H10 109.412 C1 C4 H12 110.317
C1 C4 H13 110.578 C1 C5 H11 109.412
C1 C5 H14 110.317 C1 C5 H15 110.578
S2 C1 C3 105.910 S2 C1 C4 110.170
S2 C1 C5 110.170 C3 C1 C4 110.025
C3 C1 C5 110.025 C4 C1 C5 110.447
H7 C3 H8 108.885 H7 C3 H9 108.885
H8 C3 H9 109.023 H10 C4 H12 109.189
H10 C4 H13 108.560 H11 C5 H14 109.189
H11 C5 H15 108.560 H12 C4 H13 108.748
H14 C5 H15 108.748
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability