return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H14 (Butane, 2,2-dimethyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-234.631561
Energy at 298.15K-234.646936
HF Energy-234.070879
Nuclear repulsion energy264.335615
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3157 3004 40.65      
2 A' 3149 2996 46.24      
3 A' 3142 2989 58.04      
4 A' 3135 2983 0.67      
5 A' 3088 2938 14.55      
6 A' 3070 2920 6.30      
7 A' 3061 2912 26.44      
8 A' 3056 2907 18.65      
9 A' 1615 1536 12.90      
10 A' 1610 1532 6.98      
11 A' 1603 1525 5.31      
12 A' 1589 1512 0.44      
13 A' 1576 1500 0.14      
14 A' 1504 1431 2.61      
15 A' 1487 1415 10.37      
16 A' 1473 1401 6.36      
17 A' 1423 1353 0.13      
18 A' 1325 1261 4.07      
19 A' 1285 1222 5.71      
20 A' 1144 1088 3.57      
21 A' 1066 1014 4.17      
22 A' 1026 976 1.18      
23 A' 955 909 1.29      
24 A' 883 840 0.20      
25 A' 702 668 0.41      
26 A' 494 470 0.74      
27 A' 414 394 0.00      
28 A' 369 352 0.04      
29 A' 287 273 0.04      
30 A' 267 254 0.05      
31 A" 3171 3017 26.60      
32 A" 3148 2995 20.83      
33 A" 3139 2986 27.23      
34 A" 3135 2982 1.39      
35 A" 3098 2947 13.47      
36 A" 3065 2916 25.09      
37 A" 1613 1535 0.47      
38 A" 1601 1523 11.22      
39 A" 1587 1510 0.18      
40 A" 1583 1506 0.10      
41 A" 1474 1402 10.02      
42 A" 1397 1329 0.98      
43 A" 1301 1237 1.82      
44 A" 1162 1106 0.47      
45 A" 1059 1008 0.87      
46 A" 1020 970 0.01      
47 A" 958 911 0.07      
48 A" 820 781 3.39      
49 A" 425 404 0.00      
50 A" 350 333 0.02      
51 A" 302 287 0.00      
52 A" 271 258 0.00      
53 A" 227 216 0.02      
54 A" 89 85 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 42475.6 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 40407.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.14143 0.08210 0.08192

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.032 -2.195 0.000
C2 0.843 -0.873 0.000
C3 0.000 0.435 0.000
C4 0.981 1.637 0.000
C5 -0.888 0.518 1.266
C6 -0.888 0.518 -1.266
H7 0.424 2.583 0.000
H8 0.716 -3.052 0.000
H9 -0.269 0.445 2.169
H10 -0.269 0.445 -2.169
H11 -1.630 -0.289 1.281
H12 -1.630 -0.289 -1.281
H13 -1.422 1.476 1.285
H14 -1.422 1.476 -1.285
H15 -0.602 -2.268 0.889
H16 -0.602 -2.268 -0.889
H17 1.495 -0.853 0.885
H18 1.495 -0.853 -0.885
H19 1.620 1.607 0.891
H20 1.620 1.607 -0.891

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20
C11.55092.63103.94843.13213.13214.79401.09623.43053.43052.83502.83504.15274.15271.09441.09442.17372.17374.21564.2156
C21.55091.55652.51422.55612.55613.48102.18262.77122.77122.84522.84523.50703.50702.19632.19631.09971.09972.74742.7474
C32.63101.55651.55151.54891.54892.18863.56022.18602.18602.19602.19602.18102.18102.90892.90892.16322.16322.18882.1888
C43.94842.51421.55152.52042.52041.09724.69682.77332.77333.48843.48842.73012.73014.30694.30692.69262.69261.09721.0972
C53.13212.55611.54892.52042.53242.75514.11321.09743.49161.09542.77321.09722.77732.82553.53342.77583.49112.76033.4834
C63.13212.55611.54892.52042.53242.75514.11323.49161.09742.77321.09542.77731.09723.53342.82553.49112.77583.48342.7603
H74.79403.48102.18861.09722.75512.75515.64223.12353.12353.75583.75582.50702.50705.03715.03713.70603.70601.78251.7825
H81.09622.18263.56024.69684.11324.11325.64224.23154.23153.84423.84425.16975.16971.77271.77272.49502.49504.82864.8286
H93.43052.77122.18602.77331.09743.49163.12354.23154.33861.78253.78121.78143.78493.01854.10162.53923.75862.55963.7798
H103.43052.77122.18602.77333.49161.09743.12354.23154.33863.78121.78253.78491.78144.10163.01853.75862.53923.77982.5596
H112.83502.84522.19603.48841.09542.77323.75583.84421.78253.78122.56301.77723.12192.26403.11153.19983.84383.78234.3445
H122.83502.84522.19603.48842.77321.09543.75583.84423.78121.78252.56303.12191.77723.11152.26403.84383.19984.34453.7823
H134.15273.50702.18102.73011.09722.77732.50705.16971.78143.78491.77723.12192.57043.85324.40633.75434.31793.07033.7429
H144.15273.50702.18102.73012.77731.09722.50705.16973.78491.78143.12191.77722.57044.40633.85324.31793.75433.74293.0703
H151.09442.19632.90894.30692.82553.53345.03711.77273.01854.10162.26403.11153.85324.40631.77752.53003.08984.46684.8084
H161.09442.19632.90894.30693.53342.82555.03711.77274.10163.01853.11152.26404.40633.85321.77753.08982.53004.80844.4668
H172.17371.09972.16322.69262.77583.49113.70602.49502.53923.75863.19983.84383.75434.31792.53003.08981.77002.46303.0368
H182.17371.09972.16322.69263.49112.77583.70602.49503.75862.53923.84383.19984.31793.75433.08982.53001.77003.03682.4630
H194.21562.74742.18881.09722.76033.48341.78254.82862.55963.77983.78234.34453.07033.74294.46684.80842.46303.03681.7829
H204.21562.74742.18881.09723.48342.76031.78254.82863.77982.55964.34453.78233.74293.07034.80844.46683.03682.46301.7829

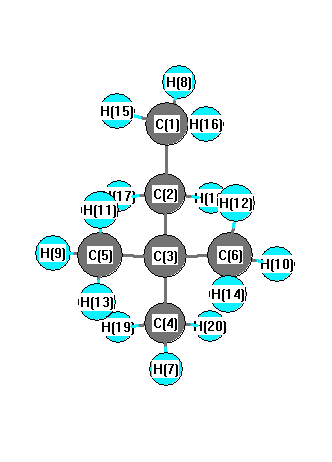
picture of Butane, 2,2-dimethyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 115.708 C1 C2 H17 108.998
C1 C2 H18 108.998 C2 C1 H8 109.892
C2 C1 H15 111.079 C2 C1 H16 111.079
C2 C3 C4 107.990 C2 C3 C5 110.797
C2 C3 C6 110.797 C3 C2 H17 107.817
C3 C2 H18 107.817 C3 C4 H7 110.264
C3 C4 H19 110.280 C3 C4 H20 110.280
C3 C5 H9 110.221 C3 C5 H11 111.128
C3 C5 H13 109.840 C3 C6 H10 110.221
C3 C6 H12 111.128 C3 C6 H14 109.840
C4 C3 C5 108.766 C4 C3 C6 108.766
C5 C3 C6 109.663 H7 C4 H19 108.646
H7 C4 H20 108.646 H8 C1 H15 108.043
H8 C1 H16 108.043 H9 C5 H11 108.760
H9 C5 H13 108.532 H10 C6 H12 108.760
H10 C6 H14 108.532 H11 C5 H13 108.294
H12 C6 H14 108.294 H15 C1 H16 108.602
H17 C2 H18 107.183 H19 C4 H20 108.677
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability